Jump to
S1C2
Energy calculated at CCD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -151.508870 |
| Energy at 298.15K | |
| HF Energy | -151.096345 |
| Nuclear repulsion energy | 52.313837 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σ |
3498 |
3347 |
163.29 |
|
|
|
| 2 |
Σ |
2283 |
2184 |
765.51 |
|
|
|
| 3 |
Σ |
1391 |
1331 |
31.39 |
|
|
|
| 4 |
Π |
604 |
578 |
0.46 |
|
|
|
| 4 |
Π |
565 |
541 |
11.05 |
|
|
|
| 5 |
Π |
405 |
388 |
29.27 |
|
|
|
| 5 |
Π |
422i |
403i |
144.77 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4162.8 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 3982.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/cc-pVDZ
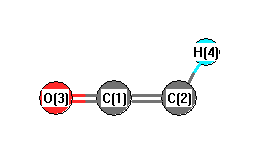
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.029 |
| C2 |
0.000 |
0.000 |
-1.250 |
| O3 |
0.000 |
0.000 |
1.206 |
| H4 |
0.000 |
0.000 |
-2.324 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
| C1 | | 1.2786 | 1.1776 | 2.3529 |
C2 | 1.2786 | | 2.4562 | 1.0743 | O3 | 1.1776 | 2.4562 | | 3.5306 | H4 | 2.3529 | 1.0743 | 3.5306 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H4 |
180.000 |
|
C2 |
C1 |
O3 |
180.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -151.513762 |
| Energy at 298.15K | |
| HF Energy | -151.096042 |
| Nuclear repulsion energy | 52.024055 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3328 |
3184 |
28.59 |
|
|
|
| 2 |
A' |
2239 |
2142 |
709.19 |
|
|
|
| 3 |
A' |
1256 |
1202 |
0.84 |
|
|
|
| 4 |
A' |
640 |
613 |
165.65 |
|
|
|
| 5 |
A' |
574 |
549 |
76.69 |
|
|
|
| 6 |
A" |
514 |
492 |
6.82 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4275.8 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 4090.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.058 |
0.000 |
| C2 |
1.007 |
-0.807 |
0.000 |
| O3 |
-1.014 |
0.633 |
0.000 |
| H4 |
2.072 |
-0.570 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
| C1 | | 1.3281 | 1.1657 | 2.1649 |
C2 | 1.3281 | | 2.4821 | 1.0904 | O3 | 1.1657 | 2.4821 | | 3.3122 | H4 | 2.1649 | 1.0904 | 3.3122 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H4 |
126.780 |
|
C2 |
C1 |
O3 |
168.842 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
