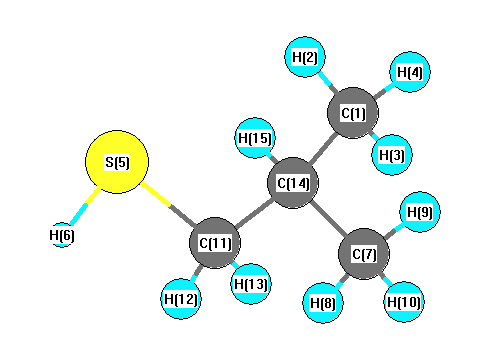
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3059 |
3041 |
22.11 |
|
|
|
| 2 |
A |
3052 |
3034 |
25.26 |
|
|
|
| 3 |
A |
3044 |
3027 |
47.39 |
|
|
|
| 4 |
A |
3039 |
3022 |
10.99 |
|
|
|
| 5 |
A |
3032 |
3015 |
6.13 |
|
|
|
| 6 |
A |
2972 |
2954 |
35.64 |
|
|
|
| 7 |
A |
2964 |
2947 |
19.60 |
|
|
|
| 8 |
A |
2961 |
2944 |
20.65 |
|
|
|
| 9 |
A |
2947 |
2930 |
6.47 |
|
|
|
| 10 |
A |
2616 |
2601 |
7.86 |
|
|
|
| 11 |
A |
1431 |
1423 |
5.12 |
|
|
|
| 12 |
A |
1425 |
1416 |
11.58 |
|
|
|
| 13 |
A |
1417 |
1409 |
2.46 |
|
|
|
| 14 |
A |
1405 |
1397 |
0.64 |
|
|
|
| 15 |
A |
1397 |
1389 |
2.63 |
|
|
|
| 16 |
A |
1351 |
1343 |
5.77 |
|
|
|
| 17 |
A |
1335 |
1327 |
6.90 |
|
|
|
| 18 |
A |
1312 |
1304 |
2.00 |
|
|
|
| 19 |
A |
1295 |
1287 |
0.63 |
|
|
|
| 20 |
A |
1213 |
1206 |
19.12 |
|
|
|
| 21 |
A |
1183 |
1177 |
6.82 |
|
|
|
| 22 |
A |
1156 |
1149 |
5.13 |
|
|
|
| 23 |
A |
1111 |
1105 |
3.48 |
|
|
|
| 24 |
A |
1039 |
1032 |
3.02 |
|
|
|
| 25 |
A |
949 |
944 |
1.99 |
|
|
|
| 26 |
A |
936 |
930 |
1.19 |
|
|
|
| 27 |
A |
897 |
891 |
1.69 |
|
|
|
| 28 |
A |
876 |
870 |
1.08 |
|
|
|
| 29 |
A |
843 |
838 |
2.81 |
|
|
|
| 30 |
A |
783 |
778 |
3.12 |
|
|
|
| 31 |
A |
710 |
706 |
1.90 |
|
|
|
| 32 |
A |
411 |
409 |
0.15 |
|
|
|
| 33 |
A |
379 |
377 |
0.95 |
|
|
|
| 34 |
A |
326 |
324 |
0.10 |
|
|
|
| 35 |
A |
246 |
245 |
0.14 |
|
|
|
| 36 |
A |
218 |
217 |
0.09 |
|
|
|
| 37 |
A |
197 |
196 |
1.86 |
|
|
|
| 38 |
A |
152 |
151 |
9.04 |
|
|
|
| 39 |
A |
74 |
73 |
9.57 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 27874.6 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 27713.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.011 |
|
|
|
| 2 |
H |
0.042 |
|
|
|
| 3 |
H |
0.028 |
|
|
|
| 4 |
H |
0.029 |
|
|
|
| 5 |
S |
-0.116 |
|
|
|
| 6 |
H |
0.084 |
|
|
|
| 7 |
C |
-0.015 |
|
|
|
| 8 |
H |
0.027 |
|
|
|
| 9 |
H |
0.030 |
|
|
|
| 10 |
H |
0.029 |
|
|
|
| 11 |
C |
-0.099 |
|
|
|
| 12 |
H |
0.055 |
|
|
|
| 13 |
H |
0.055 |
|
|
|
| 14 |
C |
-0.158 |
|
|
|
| 15 |
H |
0.020 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.933 |
-1.006 |
0.594 |
1.495 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.703 |
2.225 |
-1.825 |
| y |
2.225 |
-40.357 |
-1.114 |
| z |
-1.825 |
-1.114 |
-41.383 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.167 |
2.225 |
-1.825 |
| y |
2.225 |
0.686 |
-1.114 |
| z |
-1.825 |
-1.114 |
-0.853 |
|
| Polar |
|---|
| 3z2-r2 | -1.706 |
|---|
| x2-y2 | -0.346 |
|---|
| xy | 2.225 |
|---|
| xz | -1.825 |
|---|
| yz | -1.114 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.655 |
-0.007 |
-0.122 |
| y |
-0.007 |
9.059 |
-0.402 |
| z |
-0.122 |
-0.402 |
7.767 |
<r2> (average value of r
2) Å
2
| <r2> |
203.714 |
| (<r2>)1/2 |
14.273 |