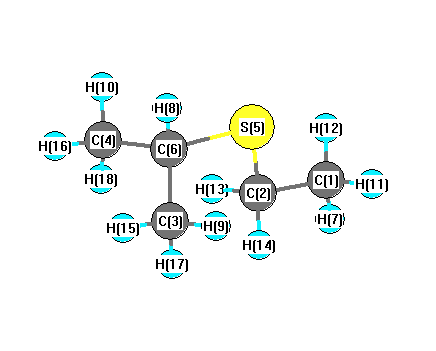
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3071 |
3053 |
19.05 |
|
|
|
| 2 |
A |
3057 |
3039 |
16.95 |
|
|
|
| 3 |
A |
3041 |
3023 |
52.12 |
|
|
|
| 4 |
A |
2976 |
2959 |
3.47 |
|
|
|
| 5 |
A |
2975 |
2958 |
21.86 |
|
|
|
| 6 |
A |
2974 |
2956 |
43.60 |
|
|
|
| 7 |
A |
2967 |
2950 |
21.19 |
|
|
|
| 8 |
A |
1429 |
1421 |
3.13 |
|
|
|
| 9 |
A |
1427 |
1418 |
4.79 |
|
|
|
| 10 |
A |
1420 |
1412 |
11.69 |
|
|
|
| 11 |
A |
1410 |
1402 |
0.87 |
|
|
|
| 12 |
A |
1346 |
1338 |
2.81 |
|
|
|
| 13 |
A |
1341 |
1333 |
1.08 |
|
|
|
| 14 |
A |
1228 |
1221 |
19.67 |
|
|
|
| 15 |
A |
1222 |
1215 |
21.66 |
|
|
|
| 16 |
A |
1135 |
1128 |
10.07 |
|
|
|
| 17 |
A |
1058 |
1052 |
17.57 |
|
|
|
| 18 |
A |
1015 |
1009 |
5.02 |
|
|
|
| 19 |
A |
970 |
964 |
4.79 |
|
|
|
| 20 |
A |
880 |
875 |
1.27 |
|
|
|
| 21 |
A |
671 |
667 |
0.63 |
|
|
|
| 22 |
A |
594 |
591 |
2.00 |
|
|
|
| 23 |
A |
453 |
451 |
1.87 |
|
|
|
| 24 |
A |
355 |
353 |
0.06 |
|
|
|
| 25 |
A |
278 |
277 |
0.83 |
|
|
|
| 26 |
A |
248 |
247 |
0.19 |
|
|
|
| 27 |
A |
150 |
149 |
0.24 |
|
|
|
| 28 |
A |
3069 |
3051 |
9.49 |
|
|
|
| 29 |
A |
3064 |
3046 |
24.89 |
|
|
|
| 30 |
A |
3037 |
3019 |
0.49 |
|
|
|
| 31 |
A |
3024 |
3006 |
8.14 |
|
|
|
| 32 |
A |
2963 |
2946 |
18.57 |
|
|
|
| 33 |
A |
1414 |
1406 |
1.57 |
|
|
|
| 34 |
A |
1410 |
1402 |
7.35 |
|
|
|
| 35 |
A |
1401 |
1393 |
0.67 |
|
|
|
| 36 |
A |
1329 |
1321 |
7.77 |
|
|
|
| 37 |
A |
1278 |
1271 |
1.59 |
|
|
|
| 38 |
A |
1208 |
1201 |
0.25 |
|
|
|
| 39 |
A |
1107 |
1100 |
1.82 |
|
|
|
| 40 |
A |
991 |
985 |
0.03 |
|
|
|
| 41 |
A |
919 |
914 |
0.23 |
|
|
|
| 42 |
A |
894 |
889 |
1.01 |
|
|
|
| 43 |
A |
768 |
763 |
4.07 |
|
|
|
| 44 |
A |
311 |
309 |
1.91 |
|
|
|
| 45 |
A |
243 |
242 |
0.04 |
|
|
|
| 46 |
A |
226 |
225 |
0.00 |
|
|
|
| 47 |
A |
64 |
63 |
1.14 |
|
|
|
| 48 |
A |
44 |
43 |
0.58 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 34226.8 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 34028.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.052 |
|
|
|
| 2 |
C |
-0.169 |
|
|
|
| 3 |
C |
0.000 |
|
|
|
| 4 |
C |
0.000 |
|
|
|
| 5 |
S |
-0.054 |
|
|
|
| 6 |
C |
-0.235 |
|
|
|
| 7 |
H |
0.037 |
|
|
|
| 8 |
H |
0.048 |
|
|
|
| 9 |
H |
0.039 |
|
|
|
| 10 |
H |
0.039 |
|
|
|
| 11 |
H |
0.047 |
|
|
|
| 12 |
H |
0.047 |
|
|
|
| 13 |
H |
0.061 |
|
|
|
| 14 |
H |
0.061 |
|
|
|
| 15 |
H |
0.032 |
|
|
|
| 16 |
H |
0.032 |
|
|
|
| 17 |
H |
0.035 |
|
|
|
| 18 |
H |
0.035 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.120 |
-1.495 |
0.000 |
1.500 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-43.206 |
0.547 |
0.000 |
| y |
0.547 |
-48.956 |
0.000 |
| z |
0.000 |
0.000 |
-48.123 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.334 |
0.547 |
0.000 |
| y |
0.547 |
-3.292 |
0.000 |
| z |
0.000 |
0.000 |
-2.042 |
|
| Polar |
|---|
| 3z2-r2 | -4.084 |
|---|
| x2-y2 | 5.750 |
|---|
| xy | 0.547 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.707 |
1.544 |
0.000 |
| y |
1.544 |
10.941 |
0.000 |
| z |
0.000 |
0.000 |
9.914 |
<r2> (average value of r
2) Å
2
| <r2> |
269.740 |
| (<r2>)1/2 |
16.424 |