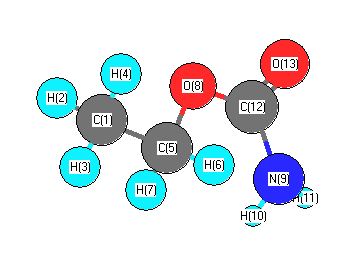
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3589 |
3568 |
34.15 |
|
|
|
| 2 |
A |
3466 |
3446 |
9.95 |
|
|
|
| 3 |
A |
3079 |
3061 |
15.03 |
|
|
|
| 4 |
A |
3075 |
3057 |
14.83 |
|
|
|
| 5 |
A |
2987 |
2969 |
14.34 |
|
|
|
| 6 |
A |
2976 |
2959 |
27.37 |
|
|
|
| 7 |
A |
2905 |
2888 |
47.40 |
|
|
|
| 8 |
A |
1809 |
1798 |
417.43 |
|
|
|
| 9 |
A |
1541 |
1532 |
120.67 |
|
|
|
| 10 |
A |
1444 |
1435 |
0.26 |
|
|
|
| 11 |
A |
1425 |
1417 |
3.32 |
|
|
|
| 12 |
A |
1404 |
1396 |
6.51 |
|
|
|
| 13 |
A |
1364 |
1356 |
10.41 |
|
|
|
| 14 |
A |
1335 |
1327 |
43.62 |
|
|
|
| 15 |
A |
1286 |
1278 |
277.18 |
|
|
|
| 16 |
A |
1245 |
1238 |
2.50 |
|
|
|
| 17 |
A |
1122 |
1115 |
4.83 |
|
|
|
| 18 |
A |
1111 |
1105 |
11.71 |
|
|
|
| 19 |
A |
1069 |
1063 |
51.06 |
|
|
|
| 20 |
A |
1048 |
1042 |
102.05 |
|
|
|
| 21 |
A |
962 |
957 |
28.24 |
|
|
|
| 22 |
A |
832 |
827 |
13.93 |
|
|
|
| 23 |
A |
806 |
802 |
1.51 |
|
|
|
| 24 |
A |
738 |
734 |
25.63 |
|
|
|
| 25 |
A |
559 |
556 |
51.50 |
|
|
|
| 26 |
A |
536 |
532 |
26.22 |
|
|
|
| 27 |
A |
527 |
524 |
46.59 |
|
|
|
| 28 |
A |
407 |
405 |
73.80 |
|
|
|
| 29 |
A |
367 |
364 |
4.11 |
|
|
|
| 30 |
A |
254 |
253 |
0.55 |
|
|
|
| 31 |
A |
209 |
208 |
2.29 |
|
|
|
| 32 |
A |
115 |
115 |
1.11 |
|
|
|
| 33 |
A |
87 |
87 |
1.22 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22837.6 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 22705.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.049 |
|
|
|
| 2 |
H |
0.052 |
|
|
|
| 3 |
H |
0.036 |
|
|
|
| 4 |
H |
0.054 |
|
|
|
| 5 |
C |
0.020 |
|
|
|
| 6 |
H |
0.040 |
|
|
|
| 7 |
H |
0.031 |
|
|
|
| 8 |
O |
-0.221 |
|
|
|
| 9 |
N |
-0.114 |
|
|
|
| 10 |
H |
0.095 |
|
|
|
| 11 |
H |
0.107 |
|
|
|
| 12 |
C |
0.183 |
|
|
|
| 13 |
O |
-0.234 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.102 |
3.963 |
-0.679 |
4.537 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.267 |
-5.555 |
1.090 |
| y |
-5.555 |
-35.649 |
-1.559 |
| z |
1.090 |
-1.559 |
-35.891 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.503 |
-5.555 |
1.090 |
| y |
-5.555 |
-0.071 |
-1.559 |
| z |
1.090 |
-1.559 |
-0.433 |
|
| Polar |
|---|
| 3z2-r2 | -0.866 |
|---|
| x2-y2 | 0.383 |
|---|
| xy | -5.555 |
|---|
| xz | 1.090 |
|---|
| yz | -1.559 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.325 |
0.215 |
-0.087 |
| y |
0.215 |
7.249 |
-0.116 |
| z |
-0.087 |
-0.116 |
5.063 |
<r2> (average value of r
2) Å
2
| <r2> |
193.730 |
| (<r2>)1/2 |
13.919 |