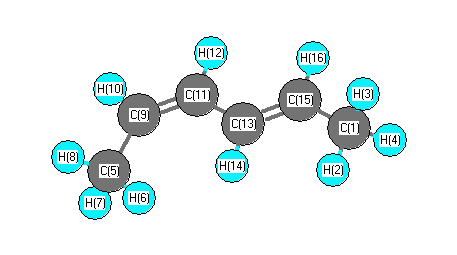
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3089 |
3071 |
49.42 |
|
|
|
| 2 |
A' |
3075 |
3057 |
21.16 |
|
|
|
| 3 |
A' |
3070 |
3052 |
5.79 |
|
|
|
| 4 |
A' |
3064 |
3046 |
20.41 |
|
|
|
| 5 |
A' |
3058 |
3040 |
0.34 |
|
|
|
| 6 |
A' |
3047 |
3030 |
1.95 |
|
|
|
| 7 |
A' |
2955 |
2938 |
31.69 |
|
|
|
| 8 |
A' |
2949 |
2932 |
44.86 |
|
|
|
| 9 |
A' |
1677 |
1667 |
3.63 |
|
|
|
| 10 |
A' |
1641 |
1631 |
7.63 |
|
|
|
| 11 |
A' |
1417 |
1409 |
9.82 |
|
|
|
| 12 |
A' |
1406 |
1398 |
19.07 |
|
|
|
| 13 |
A' |
1380 |
1372 |
3.39 |
|
|
|
| 14 |
A' |
1342 |
1334 |
2.71 |
|
|
|
| 15 |
A' |
1335 |
1327 |
3.34 |
|
|
|
| 16 |
A' |
1276 |
1269 |
1.71 |
|
|
|
| 17 |
A' |
1246 |
1239 |
1.34 |
|
|
|
| 18 |
A' |
1221 |
1214 |
1.58 |
|
|
|
| 19 |
A' |
1149 |
1142 |
0.18 |
|
|
|
| 20 |
A' |
1087 |
1080 |
0.09 |
|
|
|
| 21 |
A' |
1018 |
1012 |
0.58 |
|
|
|
| 22 |
A' |
914 |
909 |
26.67 |
|
|
|
| 23 |
A' |
904 |
898 |
20.13 |
|
|
|
| 24 |
A' |
584 |
581 |
5.79 |
|
|
|
| 25 |
A' |
454 |
452 |
0.19 |
|
|
|
| 26 |
A' |
307 |
306 |
2.27 |
|
|
|
| 27 |
A' |
144 |
143 |
0.49 |
|
|
|
| 28 |
A" |
3008 |
2990 |
15.74 |
|
|
|
| 29 |
A" |
3004 |
2987 |
15.92 |
|
|
|
| 30 |
A" |
1398 |
1390 |
7.06 |
|
|
|
| 31 |
A" |
1396 |
1388 |
5.36 |
|
|
|
| 32 |
A" |
1006 |
1000 |
0.20 |
|
|
|
| 33 |
A" |
1001 |
995 |
2.09 |
|
|
|
| 34 |
A" |
986 |
980 |
12.13 |
|
|
|
| 35 |
A" |
936 |
930 |
10.51 |
|
|
|
| 36 |
A" |
815 |
811 |
10.42 |
|
|
|
| 37 |
A" |
704 |
700 |
11.38 |
|
|
|
| 38 |
A" |
431 |
429 |
4.16 |
|
|
|
| 39 |
A" |
217 |
216 |
0.08 |
|
|
|
| 40 |
A" |
195 |
194 |
0.65 |
|
|
|
| 41 |
A" |
137 |
137 |
0.01 |
|
|
|
| 42 |
A" |
111 |
111 |
1.53 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30077.2 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 29902.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.012 |
|
|
|
| 2 |
H |
0.029 |
|
|
|
| 3 |
H |
0.048 |
|
|
|
| 4 |
H |
0.048 |
|
|
|
| 5 |
C |
-0.016 |
|
|
|
| 6 |
H |
0.035 |
|
|
|
| 7 |
H |
0.049 |
|
|
|
| 8 |
H |
0.049 |
|
|
|
| 9 |
C |
-0.100 |
|
|
|
| 10 |
H |
-0.008 |
|
|
|
| 11 |
C |
0.019 |
|
|
|
| 12 |
H |
-0.027 |
|
|
|
| 13 |
C |
0.029 |
|
|
|
| 14 |
H |
-0.029 |
|
|
|
| 15 |
C |
-0.096 |
|
|
|
| 16 |
H |
-0.018 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.303 |
0.157 |
0.000 |
0.341 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.728 |
-0.703 |
0.000 |
| y |
-0.703 |
-34.054 |
0.000 |
| z |
0.000 |
0.000 |
-40.889 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.744 |
-0.703 |
0.000 |
| y |
-0.703 |
4.755 |
0.000 |
| z |
0.000 |
0.000 |
-5.498 |
|
| Polar |
|---|
| 3z2-r2 | -10.997 |
|---|
| x2-y2 | -2.674 |
|---|
| xy | -0.703 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.752 |
-1.142 |
0.000 |
| y |
-1.142 |
19.177 |
0.000 |
| z |
0.000 |
0.000 |
5.939 |
<r2> (average value of r
2) Å
2
| <r2> |
263.225 |
| (<r2>)1/2 |
16.224 |