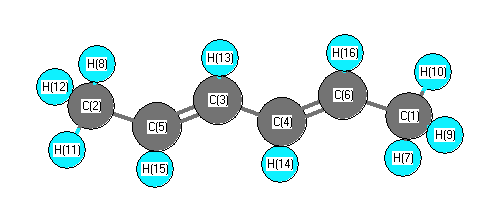
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3065 |
3047 |
0.00 |
|
|
|
| 2 |
Ag |
3051 |
3034 |
0.00 |
|
|
|
| 3 |
Ag |
3047 |
3030 |
0.00 |
|
|
|
| 4 |
Ag |
2950 |
2932 |
0.00 |
|
|
|
| 5 |
Ag |
1682 |
1672 |
0.00 |
|
|
|
| 6 |
Ag |
1414 |
1406 |
0.00 |
|
|
|
| 7 |
Ag |
1345 |
1337 |
0.00 |
|
|
|
| 8 |
Ag |
1294 |
1287 |
0.00 |
|
|
|
| 9 |
Ag |
1268 |
1260 |
0.00 |
|
|
|
| 10 |
Ag |
1169 |
1163 |
0.00 |
|
|
|
| 11 |
Ag |
1104 |
1098 |
0.00 |
|
|
|
| 12 |
Ag |
901 |
896 |
0.00 |
|
|
|
| 13 |
Ag |
426 |
424 |
0.00 |
|
|
|
| 14 |
Ag |
325 |
324 |
0.00 |
|
|
|
| 15 |
Au |
3005 |
2987 |
33.68 |
|
|
|
| 16 |
Au |
1395 |
1387 |
12.68 |
|
|
|
| 17 |
Au |
1004 |
998 |
1.42 |
|
|
|
| 18 |
Au |
995 |
989 |
43.82 |
|
|
|
| 19 |
Au |
744 |
740 |
5.80 |
|
|
|
| 20 |
Au |
255 |
253 |
1.53 |
|
|
|
| 21 |
Au |
170 |
169 |
0.00 |
|
|
|
| 22 |
Au |
105 |
105 |
1.98 |
|
|
|
| 23 |
Bg |
3005 |
2987 |
0.00 |
|
|
|
| 24 |
Bg |
1396 |
1388 |
0.00 |
|
|
|
| 25 |
Bg |
1007 |
1001 |
0.00 |
|
|
|
| 26 |
Bg |
931 |
926 |
0.00 |
|
|
|
| 27 |
Bg |
830 |
825 |
0.00 |
|
|
|
| 28 |
Bg |
333 |
331 |
0.00 |
|
|
|
| 29 |
Bg |
198 |
197 |
0.00 |
|
|
|
| 30 |
Bu |
3066 |
3049 |
101.72 |
|
|
|
| 31 |
Bu |
3057 |
3040 |
5.81 |
|
|
|
| 32 |
Bu |
3047 |
3029 |
4.31 |
|
|
|
| 33 |
Bu |
2949 |
2932 |
92.15 |
|
|
|
| 34 |
Bu |
1652 |
1642 |
8.79 |
|
|
|
| 35 |
Bu |
1407 |
1398 |
24.45 |
|
|
|
| 36 |
Bu |
1339 |
1331 |
5.78 |
|
|
|
| 37 |
Bu |
1274 |
1267 |
2.74 |
|
|
|
| 38 |
Bu |
1202 |
1195 |
2.86 |
|
|
|
| 39 |
Bu |
1065 |
1059 |
0.20 |
|
|
|
| 40 |
Bu |
916 |
910 |
45.15 |
|
|
|
| 41 |
Bu |
527 |
524 |
0.04 |
|
|
|
| 42 |
Bu |
137 |
136 |
1.43 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30025.0 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 29850.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.012 |
|
|
|
| 2 |
C |
-0.012 |
|
|
|
| 3 |
C |
0.037 |
|
|
|
| 4 |
C |
0.037 |
|
|
|
| 5 |
C |
-0.101 |
|
|
|
| 6 |
C |
-0.101 |
|
|
|
| 7 |
H |
0.028 |
|
|
|
| 8 |
H |
0.028 |
|
|
|
| 9 |
H |
0.048 |
|
|
|
| 10 |
H |
0.048 |
|
|
|
| 11 |
H |
0.048 |
|
|
|
| 12 |
H |
0.048 |
|
|
|
| 13 |
H |
-0.030 |
|
|
|
| 14 |
H |
-0.030 |
|
|
|
| 15 |
H |
-0.018 |
|
|
|
| 16 |
H |
-0.018 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.599 |
-0.364 |
0.000 |
| y |
-0.364 |
-33.822 |
0.000 |
| z |
0.000 |
0.000 |
-40.902 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.763 |
-0.364 |
0.000 |
| y |
-0.364 |
4.928 |
0.000 |
| z |
0.000 |
0.000 |
-5.692 |
|
| Polar |
|---|
| 3z2-r2 | -11.384 |
|---|
| x2-y2 | -2.777 |
|---|
| xy | -0.364 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.441 |
-0.837 |
0.000 |
| y |
-0.837 |
20.321 |
0.000 |
| z |
0.000 |
0.000 |
5.966 |
<r2> (average value of r
2) Å
2
| <r2> |
293.955 |
| (<r2>)1/2 |
17.145 |