Jump to
S1C2
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Geometric Data calculated at QCISD/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -268.582244 |
| Energy at 298.15K | -268.588882 |
| HF Energy | -267.686813 |
| Nuclear repulsion energy | 221.025273 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3313 |
3178 |
1.25 |
|
|
|
| 2 |
A' |
3287 |
3154 |
0.95 |
|
|
|
| 3 |
A' |
3275 |
3142 |
4.40 |
|
|
|
| 4 |
A' |
3168 |
3039 |
11.74 |
|
|
|
| 5 |
A' |
3064 |
2940 |
27.93 |
|
|
|
| 6 |
A' |
1682 |
1614 |
12.88 |
|
|
|
| 7 |
A' |
1588 |
1524 |
19.27 |
|
|
|
| 8 |
A' |
1496 |
1435 |
4.07 |
|
|
|
| 9 |
A' |
1435 |
1376 |
6.00 |
|
|
|
| 10 |
A' |
1418 |
1360 |
1.58 |
|
|
|
| 11 |
A' |
1280 |
1228 |
21.54 |
|
|
|
| 12 |
A' |
1254 |
1203 |
0.78 |
|
|
|
| 13 |
A' |
1187 |
1139 |
19.87 |
|
|
|
| 14 |
A' |
1120 |
1074 |
6.88 |
|
|
|
| 15 |
A' |
1033 |
991 |
19.30 |
|
|
|
| 16 |
A' |
994 |
953 |
2.38 |
|
|
|
| 17 |
A' |
944 |
905 |
18.75 |
|
|
|
| 18 |
A' |
896 |
860 |
2.56 |
|
|
|
| 19 |
A' |
662 |
635 |
1.05 |
|
|
|
| 20 |
A' |
332 |
318 |
1.62 |
|
|
|
| 21 |
A" |
3137 |
3010 |
15.27 |
|
|
|
| 22 |
A" |
1474 |
1414 |
5.96 |
|
|
|
| 23 |
A" |
1067 |
1024 |
0.96 |
|
|
|
| 24 |
A" |
854 |
819 |
0.01 |
|
|
|
| 25 |
A" |
808 |
775 |
15.36 |
|
|
|
| 26 |
A" |
753 |
722 |
47.70 |
|
|
|
| 27 |
A" |
631 |
605 |
3.70 |
|
|
|
| 28 |
A" |
601 |
577 |
4.51 |
|
|
|
| 29 |
A" |
243 |
234 |
5.27 |
|
|
|
| 30 |
A" |
131 |
125 |
0.17 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21561.5 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 20686.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
1.045 |
-0.242 |
0.000 |
| C2 |
0.000 |
0.646 |
0.000 |
| C3 |
0.500 |
-1.499 |
0.000 |
| C4 |
-1.194 |
-0.025 |
0.000 |
| C5 |
-0.866 |
-1.439 |
0.000 |
| C6 |
0.361 |
2.099 |
0.000 |
| H7 |
1.211 |
-2.324 |
0.000 |
| H8 |
-2.187 |
0.427 |
0.000 |
| H9 |
-1.554 |
-2.285 |
0.000 |
| H10 |
-0.556 |
2.710 |
0.000 |
| H11 |
0.957 |
2.359 |
0.893 |
| H12 |
0.957 |
2.359 |
-0.893 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| O1 | | 1.3720 | 1.3703 | 2.2503 | 2.2549 | 2.4391 | 2.0887 | 3.3006 | 3.3057 | 3.3589 | 2.7518 | 2.7518 |
C2 | 1.3720 | | 2.2034 | 1.3704 | 2.2579 | 1.4968 | 3.2081 | 2.1975 | 3.3177 | 2.1375 | 2.1559 | 2.1559 | C3 | 1.3703 | 2.2034 | | 2.2461 | 1.3674 | 3.6011 | 1.0888 | 3.3063 | 2.1990 | 4.3403 | 3.9869 | 3.9869 | C4 | 2.2503 | 1.3704 | 2.2461 | | 1.4512 | 2.6330 | 3.3273 | 1.0906 | 2.2880 | 2.8092 | 3.3338 | 3.3338 | C5 | 2.2549 | 2.2579 | 1.3674 | 1.4512 | | 3.7446 | 2.2577 | 2.2863 | 1.0904 | 4.1607 | 4.3066 | 4.3066 | C6 | 2.4391 | 1.4968 | 3.6011 | 2.6330 | 3.7446 | | 4.5042 | 3.0472 | 4.7839 | 1.1023 | 1.1049 | 1.1049 | H7 | 2.0887 | 3.2081 | 1.0888 | 3.3273 | 2.2577 | 4.5042 | | 4.3721 | 2.7648 | 5.3357 | 4.7746 | 4.7746 | H8 | 3.3006 | 2.1975 | 3.3063 | 1.0906 | 2.2863 | 3.0472 | 4.3721 | | 2.7853 | 2.8054 | 3.7966 | 3.7966 | H9 | 3.3057 | 3.3177 | 2.1990 | 2.2880 | 1.0904 | 4.7839 | 2.7648 | 2.7853 | | 5.0940 | 5.3545 | 5.3545 | H10 | 3.3589 | 2.1375 | 4.3403 | 2.8092 | 4.1607 | 1.1023 | 5.3357 | 2.8054 | 5.0940 | | 1.7921 | 1.7921 | H11 | 2.7518 | 2.1559 | 3.9869 | 3.3338 | 4.3066 | 1.1049 | 4.7746 | 3.7966 | 5.3545 | 1.7921 | | 1.7862 | H12 | 2.7518 | 2.1559 | 3.9869 | 3.3338 | 4.3066 | 1.1049 | 4.7746 | 3.7966 | 5.3545 | 1.7921 | 1.7862 | |
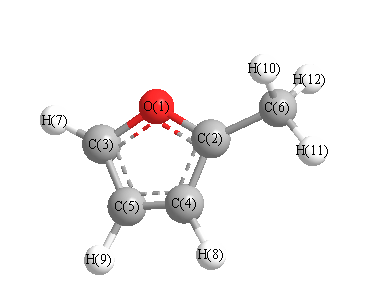 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C4 |
110.280 |
|
O1 |
C2 |
C6 |
116.402 |
| O1 |
C3 |
C5 |
110.901 |
|
O1 |
C3 |
H7 |
115.812 |
| C2 |
O1 |
C3 |
106.925 |
|
C2 |
C4 |
C5 |
106.266 |
| C2 |
C4 |
H8 |
126.118 |
|
C2 |
C6 |
H10 |
109.721 |
| C2 |
C6 |
H11 |
111.023 |
|
C2 |
C6 |
H12 |
111.023 |
| C3 |
C5 |
C6 |
73.412 |
|
C3 |
C5 |
H9 |
126.563 |
| C4 |
C2 |
C6 |
133.317 |
|
C4 |
C5 |
H9 |
127.809 |
| C5 |
C3 |
H7 |
133.287 |
|
C5 |
C4 |
H8 |
127.616 |
| H10 |
C6 |
H11 |
108.569 |
|
H10 |
C6 |
H12 |
108.569 |
| H11 |
C6 |
H12 |
107.858 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
