Jump to
S1C2
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
Geometric Data calculated at PBEPBE/cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at PBEPBE/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -269.028033 |
| Energy at 298.15K | -269.034492 |
| HF Energy | -269.028033 |
| Nuclear repulsion energy | 220.833178 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3220 |
3201 |
0.56 |
|
|
|
| 2 |
A' |
3196 |
3177 |
0.99 |
|
|
|
| 3 |
A' |
3184 |
3165 |
4.40 |
|
|
|
| 4 |
A' |
3084 |
3066 |
7.07 |
|
|
|
| 5 |
A' |
2968 |
2951 |
33.57 |
|
|
|
| 6 |
A' |
1605 |
1596 |
14.34 |
|
|
|
| 7 |
A' |
1510 |
1501 |
15.79 |
|
|
|
| 8 |
A' |
1417 |
1409 |
4.82 |
|
|
|
| 9 |
A' |
1368 |
1360 |
4.80 |
|
|
|
| 10 |
A' |
1347 |
1339 |
1.44 |
|
|
|
| 11 |
A' |
1225 |
1218 |
10.33 |
|
|
|
| 12 |
A' |
1197 |
1190 |
5.66 |
|
|
|
| 13 |
A' |
1140 |
1134 |
14.43 |
|
|
|
| 14 |
A' |
1082 |
1076 |
3.21 |
|
|
|
| 15 |
A' |
998 |
992 |
26.83 |
|
|
|
| 16 |
A' |
948 |
943 |
2.48 |
|
|
|
| 17 |
A' |
909 |
903 |
11.63 |
|
|
|
| 18 |
A' |
869 |
863 |
2.43 |
|
|
|
| 19 |
A' |
648 |
644 |
0.63 |
|
|
|
| 20 |
A' |
328 |
326 |
1.86 |
|
|
|
| 21 |
A" |
3028 |
3011 |
12.31 |
|
|
|
| 22 |
A" |
1392 |
1384 |
7.39 |
|
|
|
| 23 |
A" |
1010 |
1004 |
2.57 |
|
|
|
| 24 |
A" |
832 |
827 |
0.11 |
|
|
|
| 25 |
A" |
771 |
766 |
18.31 |
|
|
|
| 26 |
A" |
699 |
695 |
40.10 |
|
|
|
| 27 |
A" |
620 |
616 |
0.71 |
|
|
|
| 28 |
A" |
598 |
594 |
4.37 |
|
|
|
| 29 |
A" |
234 |
232 |
4.67 |
|
|
|
| 30 |
A" |
113 |
112 |
0.13 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20767.6 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 20647.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
1.048 |
-0.242 |
0.000 |
| C2 |
0.000 |
0.652 |
0.000 |
| C3 |
0.505 |
-1.502 |
0.000 |
| C4 |
-1.193 |
-0.034 |
0.000 |
| C5 |
-0.865 |
-1.435 |
0.000 |
| C6 |
0.354 |
2.098 |
0.000 |
| H7 |
1.225 |
-2.324 |
0.000 |
| H8 |
-2.192 |
0.413 |
0.000 |
| H9 |
-1.558 |
-2.282 |
0.000 |
| H10 |
-0.567 |
2.710 |
0.000 |
| H11 |
0.953 |
2.373 |
0.893 |
| H12 |
0.953 |
2.373 |
-0.893 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| O1 | | 1.3774 | 1.3728 | 2.2508 | 2.2549 | 2.4399 | 2.0904 | 3.3057 | 3.3101 | 3.3644 | 2.7650 | 2.7650 |
C2 | 1.3774 | | 2.2128 | 1.3767 | 2.2594 | 1.4882 | 3.2189 | 2.2052 | 3.3223 | 2.1345 | 2.1607 | 2.1607 | C3 | 1.3728 | 2.2128 | | 2.2444 | 1.3716 | 3.6031 | 1.0931 | 3.3079 | 2.2055 | 4.3465 | 4.0025 | 4.0025 | C4 | 2.2508 | 1.3767 | 2.2444 | | 1.4384 | 2.6344 | 3.3305 | 1.0947 | 2.2769 | 2.8152 | 3.3469 | 3.3469 | C5 | 2.2549 | 2.2594 | 1.3716 | 1.4384 | | 3.7371 | 2.2716 | 2.2752 | 1.0944 | 4.1558 | 4.3137 | 4.3137 | C6 | 2.4399 | 1.4882 | 3.6031 | 2.6344 | 3.7371 | | 4.5071 | 3.0530 | 4.7789 | 1.1058 | 1.1103 | 1.1103 | H7 | 2.0904 | 3.2189 | 1.0931 | 3.3305 | 2.2716 | 4.5071 | | 4.3786 | 2.7838 | 5.3439 | 4.7898 | 4.7898 | H8 | 3.3057 | 2.2052 | 3.3079 | 1.0947 | 2.2752 | 3.0530 | 4.3786 | | 2.7685 | 2.8140 | 3.8122 | 3.8122 | H9 | 3.3101 | 3.3223 | 2.2055 | 2.2769 | 1.0944 | 4.7789 | 2.7838 | 2.7685 | | 5.0895 | 5.3644 | 5.3644 | H10 | 3.3644 | 2.1345 | 4.3465 | 2.8152 | 4.1558 | 1.1058 | 5.3439 | 2.8140 | 5.0895 | | 1.7946 | 1.7946 | H11 | 2.7650 | 2.1607 | 4.0025 | 3.3469 | 4.3137 | 1.1103 | 4.7898 | 3.8122 | 5.3644 | 1.7946 | | 1.7868 | H12 | 2.7650 | 2.1607 | 4.0025 | 3.3469 | 4.3137 | 1.1103 | 4.7898 | 3.8122 | 5.3644 | 1.7946 | 1.7868 | |
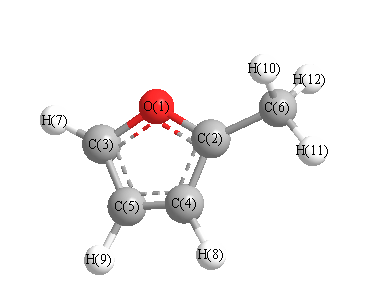 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C4 |
109.623 |
|
O1 |
C2 |
C6 |
116.693 |
| O1 |
C3 |
C5 |
110.501 |
|
O1 |
C3 |
H7 |
115.463 |
| C2 |
O1 |
C3 |
107.144 |
|
C2 |
C4 |
C5 |
106.737 |
| C2 |
C4 |
H8 |
125.946 |
|
C2 |
C6 |
H10 |
109.867 |
| C2 |
C6 |
H11 |
111.690 |
|
C2 |
C6 |
H12 |
111.690 |
| C3 |
C5 |
C6 |
73.770 |
|
C3 |
C5 |
H9 |
126.492 |
| C4 |
C2 |
C6 |
133.684 |
|
C4 |
C5 |
H9 |
127.514 |
| C5 |
C3 |
H7 |
134.036 |
|
C5 |
C4 |
H8 |
127.317 |
| H10 |
C6 |
H11 |
108.147 |
|
H10 |
C6 |
H12 |
108.147 |
| H11 |
C6 |
H12 |
107.151 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.154 |
|
|
|
| 2 |
C |
0.014 |
|
|
|
| 3 |
C |
0.088 |
|
|
|
| 4 |
C |
-0.047 |
|
|
|
| 5 |
C |
-0.006 |
|
|
|
| 6 |
C |
0.003 |
|
|
|
| 7 |
H |
-0.004 |
|
|
|
| 8 |
H |
-0.031 |
|
|
|
| 9 |
H |
-0.018 |
|
|
|
| 10 |
H |
0.044 |
|
|
|
| 11 |
H |
0.055 |
|
|
|
| 12 |
H |
0.055 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.269 |
0.425 |
0.000 |
0.503 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-34.051 |
-0.454 |
0.000 |
| y |
-0.454 |
-30.398 |
0.000 |
| z |
0.000 |
0.000 |
-37.683 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.011 |
-0.454 |
0.000 |
| y |
-0.454 |
5.469 |
0.000 |
| z |
0.000 |
0.000 |
-5.459 |
|
| Polar |
|---|
| 3z2-r2 | -10.917 |
|---|
| x2-y2 | -3.653 |
|---|
| xy | -0.454 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.663 |
0.377 |
0.000 |
| y |
0.377 |
10.898 |
0.000 |
| z |
0.000 |
0.000 |
4.440 |
<r2> (average value of r
2) Å
2
| <r2> |
142.548 |
| (<r2>)1/2 |
11.939 |
