return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H4F2 (1,4-difluorobenzene)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes D2H 1Ag
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-429.608310
Energy at 298.15K-429.613125
Nuclear repulsion energy339.086627
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 Ag 3252 3120 0.00      
2 Ag 1704 1635 0.00      
3 Ag 1314 1261 0.00      
4 Ag 1158 1111 0.00      
5 Ag 872 837 0.00      
6 Ag 452 434 0.00      
7 Au 965 926 0.00      
8 Au 429 412 0.00      
9 B1g 831 798 0.00      
10 B1u 3235 3104 4.91      
11 B1u 1574 1510 247.42      
12 B1u 1257 1206 146.12      
13 B1u 1027 986 3.43      
14 B1u 749 718 50.76      
15 B2g 932 894 0.00      
16 B2g 682 654 0.00      
17 B2g 385 369 0.00      
18 B2u 3251 3119 1.34      
19 B2u 1457 1398 0.58      
20 B2u 1295 1242 2.66      
21 B2u 1098 1054 13.83      
22 B2u 346 332 4.60      
23 B3g 3237 3106 0.00      
24 B3g 1685 1617 0.00      
25 B3g 1296 1243 0.00      
26 B3g 644 618 0.00      
27 B3g 441 423 0.00      
28 B3u 861 826 78.76      
29 B3u 526 505 11.59      
30 B3u 162 155 1.58      

Unscaled Zero Point Vibrational Energy (zpe) 18557.8 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 17804.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.18655 0.04681 0.03742

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is D2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.384
C2 0.000 0.000 -1.384
C3 0.000 1.221 0.702
C4 0.000 -1.221 0.702
C5 0.000 -1.221 -0.702
C6 0.000 1.221 -0.702
F7 0.000 0.000 2.734
F8 0.000 0.000 -2.734
H9 0.000 2.157 1.270
H10 0.000 -2.157 1.270
H11 0.000 -2.157 -1.270
H12 0.000 2.157 -1.270

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 F7 F8 H9 H10 H11 H12
C12.76711.39851.39852.41722.41721.35044.11752.15992.15993.41923.4192
C22.76712.41722.41721.39851.39854.11751.35043.41923.41922.15992.1599
C31.39852.41722.44282.81791.40472.37053.64691.09393.42563.91172.1825
C41.39852.41722.44281.40472.81792.37053.64693.42561.09392.18253.9117
C52.41721.39852.81791.40472.44283.64692.37053.91172.18251.09393.4256
C62.41721.39851.40472.81792.44283.64692.37052.18253.91173.42561.0939
F71.35044.11752.37052.37053.64693.64695.46792.60702.60704.54754.5475
F84.11751.35043.64693.64692.37052.37055.46794.54754.54752.60702.6070
H92.15993.41921.09393.42563.91172.18252.60704.54754.31375.00552.5391
H102.15993.41923.42561.09392.18253.91172.60704.54754.31372.53915.0055
H113.41922.15993.91172.18251.09393.42564.54752.60705.00552.53914.3137
H123.41922.15992.18253.91173.42561.09394.54752.60702.53915.00554.3137

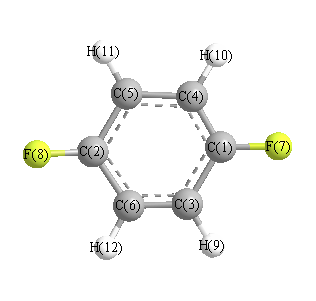
picture of 1,4-difluorobenzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C3 C6 119.148 C1 C3 H9 119.623
C1 C4 C5 119.148 C1 C4 H10 119.623
C2 C5 C4 119.148 C2 C5 H11 119.623
C2 C6 C3 119.148 C2 C6 H12 119.623
C3 C1 C4 121.703 C3 C1 F7 119.148
C3 C6 H12 121.229 C4 C1 F7 119.148
C4 C5 H11 121.229 C5 C2 C6 121.703
C5 C2 F8 119.148 C5 C4 H10 121.229
C6 C2 F8 119.148 C6 C3 H9 121.229
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability