Jump to
S1C2
Energy calculated at QCISD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -577.824355 |
| Energy at 298.15K | -577.832051 |
| Nuclear repulsion energy | 157.537885 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3146 |
3018 |
27.59 |
|
|
|
| 2 |
A' |
3100 |
2974 |
26.25 |
|
|
|
| 3 |
A' |
3077 |
2952 |
11.60 |
|
|
|
| 4 |
A' |
3055 |
2931 |
25.06 |
|
|
|
| 5 |
A' |
1507 |
1446 |
4.00 |
|
|
|
| 6 |
A' |
1494 |
1433 |
0.18 |
|
|
|
| 7 |
A' |
1491 |
1430 |
1.44 |
|
|
|
| 8 |
A' |
1422 |
1364 |
0.16 |
|
|
|
| 9 |
A' |
1388 |
1332 |
6.99 |
|
|
|
| 10 |
A' |
1288 |
1236 |
26.51 |
|
|
|
| 11 |
A' |
1134 |
1088 |
0.19 |
|
|
|
| 12 |
A' |
1073 |
1030 |
0.88 |
|
|
|
| 13 |
A' |
925 |
887 |
10.35 |
|
|
|
| 14 |
A' |
764 |
733 |
29.95 |
|
|
|
| 15 |
A' |
369 |
354 |
2.28 |
|
|
|
| 16 |
A' |
238 |
228 |
1.73 |
|
|
|
| 17 |
A" |
3166 |
3037 |
27.75 |
|
|
|
| 18 |
A" |
3138 |
3011 |
30.22 |
|
|
|
| 19 |
A" |
3117 |
2991 |
0.09 |
|
|
|
| 20 |
A" |
1497 |
1436 |
6.39 |
|
|
|
| 21 |
A" |
1323 |
1269 |
0.00 |
|
|
|
| 22 |
A" |
1253 |
1202 |
0.35 |
|
|
|
| 23 |
A" |
1101 |
1056 |
0.88 |
|
|
|
| 24 |
A" |
873 |
837 |
0.09 |
|
|
|
| 25 |
A" |
755 |
725 |
1.96 |
|
|
|
| 26 |
A" |
244 |
234 |
0.02 |
|
|
|
| 27 |
A" |
124 |
119 |
1.29 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21029.8 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 20176.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.592 |
0.000 |
| C2 |
0.902 |
-0.640 |
0.000 |
| C3 |
2.386 |
-0.245 |
0.000 |
| Cl4 |
-1.747 |
0.134 |
0.000 |
| H5 |
0.166 |
1.213 |
0.896 |
| H6 |
0.166 |
1.213 |
-0.896 |
| H7 |
0.669 |
-1.256 |
-0.888 |
| H8 |
0.669 |
-1.256 |
0.888 |
| H9 |
3.030 |
-1.142 |
0.000 |
| H10 |
2.639 |
0.354 |
-0.894 |
| H11 |
2.639 |
0.354 |
0.894 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5270 | 2.5281 | 1.8062 | 1.1029 | 1.1029 | 2.1570 | 2.1570 | 3.4907 | 2.7966 | 2.7966 |
C2 | 1.5270 | | 1.5356 | 2.7599 | 2.1859 | 2.1859 | 1.1056 | 1.1056 | 2.1863 | 2.1927 | 2.1927 | C3 | 2.5281 | 1.5356 | | 4.1502 | 2.8023 | 2.8023 | 2.1812 | 2.1812 | 1.1042 | 1.1060 | 1.1060 | Cl4 | 1.8062 | 2.7599 | 4.1502 | | 2.3724 | 2.3724 | 2.9261 | 2.9261 | 4.9445 | 4.4820 | 4.4820 | H5 | 1.1029 | 2.1859 | 2.8023 | 2.3724 | | 1.7926 | 3.0876 | 2.5195 | 3.8139 | 3.1713 | 2.6174 | H6 | 1.1029 | 2.1859 | 2.8023 | 2.3724 | 1.7926 | | 2.5195 | 3.0876 | 3.8139 | 2.6174 | 3.1713 | H7 | 2.1570 | 1.1056 | 2.1812 | 2.9261 | 3.0876 | 2.5195 | | 1.7768 | 2.5247 | 2.5445 | 3.1068 | H8 | 2.1570 | 1.1056 | 2.1812 | 2.9261 | 2.5195 | 3.0876 | 1.7768 | | 2.5247 | 3.1068 | 2.5445 | H9 | 3.4907 | 2.1863 | 1.1042 | 4.9445 | 3.8139 | 3.8139 | 2.5247 | 2.5247 | | 1.7863 | 1.7863 | H10 | 2.7966 | 2.1927 | 1.1060 | 4.4820 | 3.1713 | 2.6174 | 2.5445 | 3.1068 | 1.7863 | | 1.7885 | H11 | 2.7966 | 2.1927 | 1.1060 | 4.4820 | 2.6174 | 3.1713 | 3.1068 | 2.5445 | 1.7863 | 1.7885 | |
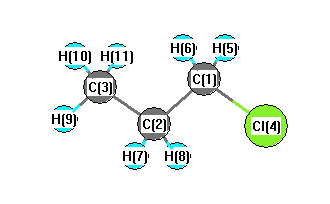 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
111.269 |
|
C1 |
C2 |
H7 |
108.984 |
| C1 |
C2 |
H8 |
108.984 |
|
C2 |
C1 |
Cl4 |
111.514 |
| C2 |
C1 |
H5 |
111.416 |
|
C2 |
C1 |
H6 |
111.416 |
| C2 |
C3 |
H9 |
110.767 |
|
C2 |
C3 |
H10 |
111.169 |
| C2 |
C3 |
H11 |
111.169 |
|
C3 |
C2 |
H7 |
110.279 |
| C3 |
C2 |
H8 |
110.279 |
|
Cl4 |
C1 |
H5 |
106.771 |
| Cl4 |
C1 |
H6 |
106.771 |
|
H5 |
C1 |
H6 |
108.720 |
| H7 |
C2 |
H8 |
106.934 |
|
H9 |
C3 |
H10 |
107.843 |
| H9 |
C3 |
H11 |
107.843 |
|
H10 |
C3 |
H11 |
107.903 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -577.824443 |
| Energy at 298.15K | -577.832251 |
| Nuclear repulsion energy | 161.564406 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3167 |
3038 |
16.62 |
|
|
|
| 2 |
A |
3159 |
3031 |
20.70 |
|
|
|
| 3 |
A |
3140 |
3013 |
35.14 |
|
|
|
| 4 |
A |
3111 |
2985 |
23.55 |
|
|
|
| 5 |
A |
3100 |
2974 |
9.56 |
|
|
|
| 6 |
A |
3062 |
2937 |
16.26 |
|
|
|
| 7 |
A |
3054 |
2930 |
30.02 |
|
|
|
| 8 |
A |
1506 |
1445 |
3.98 |
|
|
|
| 9 |
A |
1499 |
1438 |
5.94 |
|
|
|
| 10 |
A |
1484 |
1424 |
2.34 |
|
|
|
| 11 |
A |
1479 |
1419 |
2.38 |
|
|
|
| 12 |
A |
1427 |
1369 |
3.94 |
|
|
|
| 13 |
A |
1389 |
1333 |
0.72 |
|
|
|
| 14 |
A |
1342 |
1287 |
30.25 |
|
|
|
| 15 |
A |
1292 |
1239 |
4.01 |
|
|
|
| 16 |
A |
1243 |
1192 |
0.57 |
|
|
|
| 17 |
A |
1126 |
1080 |
1.21 |
|
|
|
| 18 |
A |
1096 |
1051 |
0.77 |
|
|
|
| 19 |
A |
1067 |
1024 |
1.92 |
|
|
|
| 20 |
A |
916 |
878 |
7.08 |
|
|
|
| 21 |
A |
878 |
842 |
2.45 |
|
|
|
| 22 |
A |
801 |
769 |
13.47 |
|
|
|
| 23 |
A |
682 |
654 |
15.73 |
|
|
|
| 24 |
A |
426 |
409 |
1.13 |
|
|
|
| 25 |
A |
301 |
289 |
0.46 |
|
|
|
| 26 |
A |
224 |
215 |
1.08 |
|
|
|
| 27 |
A |
137 |
131 |
0.75 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21053.2 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 20198.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.186 |
0.886 |
0.312 |
| C2 |
-1.148 |
0.566 |
-0.362 |
| C3 |
-1.806 |
-0.722 |
0.146 |
| Cl4 |
1.452 |
-0.349 |
-0.069 |
| H5 |
0.586 |
1.854 |
-0.029 |
| H6 |
0.088 |
0.907 |
1.410 |
| H7 |
-1.818 |
1.429 |
-0.172 |
| H8 |
-0.994 |
0.514 |
-1.456 |
| H9 |
-2.780 |
-0.885 |
-0.349 |
| H10 |
-1.166 |
-1.599 |
-0.054 |
| H11 |
-1.983 |
-0.669 |
1.237 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5283 | 2.5653 | 1.8093 | 1.1017 | 1.1032 | 2.1320 | 2.1575 | 3.5168 | 2.8526 | 2.8250 |
C2 | 1.5283 | | 1.5331 | 2.7719 | 2.1859 | 2.1873 | 1.1086 | 1.1061 | 2.1837 | 2.1871 | 2.1864 | C3 | 2.5653 | 1.5331 | | 3.2858 | 3.5201 | 2.7997 | 2.1746 | 2.1804 | 1.1044 | 1.1036 | 1.1064 | Cl4 | 1.8093 | 2.7719 | 3.2858 | | 2.3676 | 2.3721 | 3.7236 | 2.9413 | 4.2741 | 2.9008 | 3.6885 | H5 | 1.1017 | 2.1859 | 3.5201 | 2.3676 | | 1.7938 | 2.4461 | 2.5157 | 4.3516 | 3.8725 | 3.8177 | H6 | 1.1032 | 2.1873 | 2.7997 | 2.3721 | 1.7938 | | 2.5318 | 3.0886 | 3.8117 | 3.1618 | 2.6085 | H7 | 2.1320 | 1.1086 | 2.1746 | 3.7236 | 2.4461 | 2.5318 | | 1.7784 | 2.5123 | 3.0996 | 2.5333 | H8 | 2.1575 | 1.1061 | 2.1804 | 2.9413 | 2.5157 | 3.0886 | 1.7784 | | 2.5242 | 2.5418 | 3.1034 | H9 | 3.5168 | 2.1837 | 1.1044 | 4.2741 | 4.3516 | 3.8117 | 2.5123 | 2.5242 | | 1.7886 | 1.7877 | H10 | 2.8526 | 2.1871 | 1.1036 | 2.9008 | 3.8725 | 3.1618 | 3.0996 | 2.5418 | 1.7886 | | 1.7881 | H11 | 2.8250 | 2.1864 | 1.1064 | 3.6885 | 3.8177 | 2.6085 | 2.5333 | 3.1034 | 1.7877 | 1.7881 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
113.846 |
|
C1 |
C2 |
H7 |
106.823 |
| C1 |
C2 |
H8 |
108.912 |
|
C2 |
C1 |
Cl4 |
112.030 |
| C2 |
C1 |
H5 |
111.399 |
|
C2 |
C1 |
H6 |
111.420 |
| C2 |
C3 |
H9 |
110.720 |
|
C2 |
C3 |
H10 |
111.043 |
| C2 |
C3 |
H11 |
110.821 |
|
C3 |
C2 |
H7 |
109.768 |
| C3 |
C2 |
H8 |
110.362 |
|
Cl4 |
C1 |
H5 |
106.292 |
| Cl4 |
C1 |
H6 |
106.542 |
|
H5 |
C1 |
H6 |
108.893 |
| H7 |
C2 |
H8 |
106.842 |
|
H9 |
C3 |
H10 |
108.195 |
| H9 |
C3 |
H11 |
107.924 |
|
H10 |
C3 |
H11 |
108.019 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
