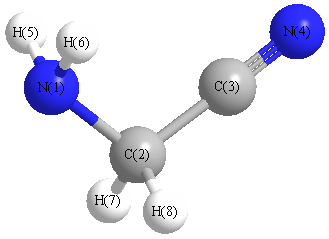
Vibrational Frequencies calculated at BLYP/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3338 |
3344 |
0.09 |
|
|
|
| 2 |
A' |
2965 |
2970 |
14.32 |
|
|
|
| 3 |
A' |
2238 |
2242 |
2.30 |
|
|
|
| 4 |
A' |
1613 |
1616 |
13.92 |
|
|
|
| 5 |
A' |
1397 |
1399 |
5.27 |
|
|
|
| 6 |
A' |
1307 |
1309 |
10.32 |
|
|
|
| 7 |
A' |
1076 |
1078 |
18.44 |
|
|
|
| 8 |
A' |
899 |
900 |
106.22 |
|
|
|
| 9 |
A' |
829 |
830 |
43.85 |
|
|
|
| 10 |
A' |
547 |
548 |
9.36 |
|
|
|
| 11 |
A' |
205 |
206 |
7.80 |
|
|
|
| 12 |
A" |
3410 |
3415 |
0.25 |
|
|
|
| 13 |
A" |
3002 |
3007 |
6.13 |
|
|
|
| 14 |
A" |
1343 |
1345 |
0.24 |
|
|
|
| 15 |
A" |
1155 |
1157 |
0.04 |
|
|
|
| 16 |
A" |
867 |
868 |
0.00 |
|
|
|
| 17 |
A" |
393 |
393 |
16.49 |
|
|
|
| 18 |
A" |
266 |
267 |
36.59 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13424.0 cm
-1
Scaled (by 1.0016) Zero Point Vibrational Energy (zpe) 13445.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.129 |
|
|
|
| 2 |
C |
0.094 |
|
|
|
| 3 |
C |
-0.152 |
|
|
|
| 4 |
N |
-0.085 |
|
|
|
| 5 |
H |
0.079 |
|
|
|
| 6 |
H |
0.079 |
|
|
|
| 7 |
H |
0.057 |
|
|
|
| 8 |
H |
0.057 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.581 |
1.894 |
0.000 |
2.467 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.917 |
6.295 |
0.000 |
| y |
6.295 |
-28.807 |
0.000 |
| z |
0.000 |
0.000 |
-21.789 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.619 |
6.295 |
0.000 |
| y |
6.295 |
-4.954 |
0.000 |
| z |
0.000 |
0.000 |
5.573 |
|
| Polar |
|---|
| 3z2-r2 | 11.146 |
|---|
| x2-y2 | 2.890 |
|---|
| xy | 6.295 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.154 |
-1.005 |
0.000 |
| y |
-1.005 |
5.488 |
0.000 |
| z |
0.000 |
0.000 |
3.808 |
<r2> (average value of r
2) Å
2
| <r2> |
83.813 |
| (<r2>)1/2 |
9.155 |