return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3OC2H5 (Ethane, methoxy-)

using model chemistry: QCISD(T)/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD(T)/cc-pVDZ
 hartrees
Energy at 0K-193.810982
Energy at 298.15K-193.819950
HF Energy-193.116852
Nuclear repulsion energy132.956697
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3141 2994        
2 A' 3138 2991        
3 A' 3052 2909        
4 A' 2983 2843        
5 A' 2964 2825        
6 A' 1526 1454        
7 A' 1498 1427        
8 A' 1491 1421        
9 A' 1471 1402        
10 A' 1429 1362        
11 A' 1385 1320        
12 A' 1239 1181        
13 A' 1178 1122        
14 A' 1113 1061        
15 A' 1052 1003        
16 A' 877 836        
17 A' 470 448        
18 A' 289 276        
19 A" 3147 2999        
20 A" 3037 2895        
21 A" 3000 2860        
22 A" 1476 1406        
23 A" 1472 1403        
24 A" 1289 1229        
25 A" 1199 1142        
26 A" 1163 1108        
27 A" 818 780        
28 A" 270 257        
29 A" 215 205        
30 A" 111 105        

Unscaled Zero Point Vibrational Energy (zpe) 23744.6 cm-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 22631.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD(T)/cc-pVDZ
ABC
0.92013 0.13851 0.12940

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD(T)/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.329 1.223 0.000
O2 0.008 0.719 0.000
C3 0.000 -0.700 0.000
C4 -1.452 -1.165 0.000
H5 1.263 2.325 0.000
H6 1.896 0.896 0.901
H7 1.896 0.896 -0.901
H8 0.536 -1.090 -0.897
H9 0.536 -1.090 0.897
H10 -1.506 -2.270 0.000
H11 -1.974 -0.784 0.896
H12 -1.974 -0.784 -0.896

Atom - Atom Distances (Å)
  C1 O2 C3 C4 H5 H6 H7 H8 H9 H10 H11 H12
C11.41402.33793.66551.10381.11321.11322.60482.60484.49853.96743.9674
O21.41401.41922.38332.03852.09902.09902.08722.08723.35012.64372.6437
C32.33791.41921.52433.27852.63682.63681.11521.11522.17502.16952.1695
C43.66552.38331.52434.42184.03294.03292.18212.18211.10611.10531.1053
H51.10382.03853.27854.42181.80361.80363.60533.60535.36474.57684.5768
H61.11322.09902.63684.03291.80361.80123.00402.40714.73324.21844.5852
H71.11322.09902.63684.03291.80361.80122.40713.00404.73324.58524.2184
H82.60482.08721.11522.18213.60533.00402.40711.79322.52273.09992.5287
H92.60482.08721.11522.18213.60532.40713.00401.79322.52272.52873.0999
H104.49853.35012.17501.10615.36474.73324.73322.52272.52271.79761.7976
H113.96742.64372.16951.10534.57684.21844.58523.09992.52871.79761.7930
H123.96742.64372.16951.10534.57684.58524.21842.52873.09991.79761.7930

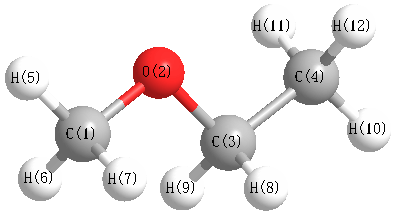
picture of Ethane, methoxy- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 O2 C3 111.214 O2 C1 H5 107.483
O2 C1 H6 111.764 O2 C1 H7 111.764
O2 C3 C4 108.076 O2 C3 H8 110.315
O2 C3 H9 110.315 C3 C4 H10 110.549
C3 C4 H11 110.160 C3 C4 H12 110.160
C4 C3 H8 110.562 C4 C3 H9 110.562
H5 C1 H6 108.882 H5 C1 H7 108.882
H6 C1 H7 108.002 H8 C3 H9 107.025
H10 C4 H11 108.759 H10 C4 H12 108.759
H11 C4 H12 108.402
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability