return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3NHNHCH3 (dimethyl hydrazine)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-189.934310
Energy at 298.15K-189.944767
Nuclear repulsion energy136.257249
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3519 3376 0.43      
2 A 3140 3013 2.59      
3 A 3099 2973 15.67      
4 A 3024 2901 92.52      
5 A 1562 1499 0.78      
6 A 1519 1458 1.15      
7 A 1493 1432 0.23      
8 A 1451 1392 0.08      
9 A 1280 1228 17.04      
10 A 1172 1124 0.08      
11 A 1142 1095 0.87      
12 A 951 913 21.67      
13 A 771 740 117.41      
14 A 343 329 7.23      
15 A 325 312 0.38      
16 A 182 175 1.11      
17 B 3535 3391 0.54      
18 B 3140 3012 55.65      
19 B 3097 2971 73.31      
20 B 3009 2887 8.64      
21 B 1519 1458 9.32      
22 B 1484 1424 14.80      
23 B 1458 1399 0.42      
24 B 1438 1380 1.32      
25 B 1181 1133 1.87      
26 B 1145 1099 20.68      
27 B 1040 998 0.25      
28 B 824 790 29.33      
29 B 514 493 2.16      
30 B 233 224 0.81      

Unscaled Zero Point Vibrational Energy (zpe) 24295.1 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 23308.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.48721 0.18650 0.15314

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 -0.162 0.696 -0.695
N2 0.162 -0.696 -0.695
C3 0.162 1.434 0.532
C4 -0.162 -1.434 0.532
H5 -1.170 0.747 -0.859
H6 1.170 -0.747 -0.859
H7 -0.100 2.494 0.369
H8 -0.352 1.089 1.454
H9 1.253 1.379 0.707
H10 0.100 -2.494 0.369
H11 0.352 -1.089 1.454
H12 -1.253 -1.379 0.707

Atom - Atom Distances (Å)
  N1 N2 C3 C4 H5 H6 H7 H8 H9 H10 H11 H12
N11.43021.46742.45821.02291.97162.08962.19222.10573.37352.84032.7320
N21.43022.45821.46741.97161.02293.37352.84032.73202.08962.19222.1057
C31.46742.45822.88632.04502.77641.10451.11081.10613.93222.69273.1541
C42.45821.46742.88632.77642.04503.93222.69273.15411.10451.11081.1061
H51.02291.97162.04502.77642.77722.38842.47712.95393.69183.32252.6423
H61.97161.02292.77642.04502.77723.69183.32252.64232.38842.47712.9539
H72.08963.37351.10453.93222.38843.69181.79351.78594.99273.77124.0557
H82.19222.84031.11082.69272.47713.32251.79351.79403.77122.28872.7314
H92.10572.73201.10613.15412.95392.64231.78591.79404.05572.73143.7269
H103.37352.08963.93221.10453.69182.38844.99273.77124.05571.79351.7859
H112.84032.19222.69271.11083.32252.47713.77122.28872.73141.79351.7940
H122.73202.10573.15411.10612.64232.95394.05572.73143.72691.78591.7940

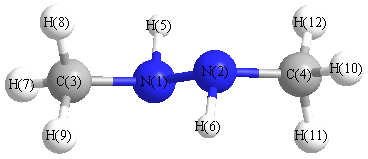
picture of dimethyl hydrazine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 N2 C4 116.065 N1 N2 H6 105.775
N1 C3 H7 107.846 N1 C3 H8 115.797
N1 C3 H9 109.006 N2 N1 C3 116.065
N2 N1 H5 105.775 N2 C4 H10 107.846
N2 C4 H11 115.797 N2 C4 H12 109.006
C3 N1 H5 109.109 C4 N2 H6 109.109
H7 C3 H8 108.112 H7 C3 H9 107.778
H8 C3 H9 108.041 H10 C4 H11 108.112
H10 C4 H12 107.778 H11 C4 H12 108.041
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability