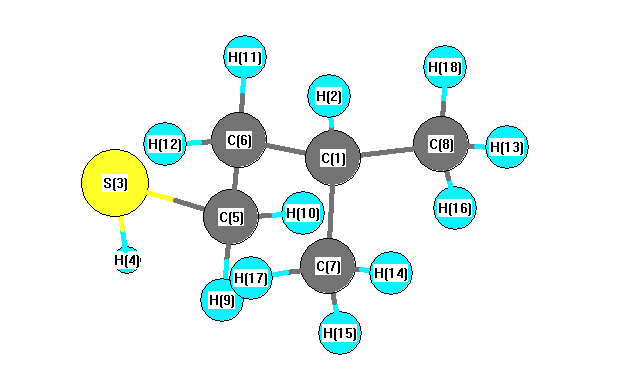
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3049 |
3031 |
42.41 |
|
|
|
| 2 |
A' |
3044 |
3026 |
57.98 |
|
|
|
| 3 |
A' |
2989 |
2972 |
26.20 |
|
|
|
| 4 |
A' |
2967 |
2950 |
5.72 |
|
|
|
| 5 |
A' |
2959 |
2942 |
36.10 |
|
|
|
| 6 |
A' |
2943 |
2926 |
8.37 |
|
|
|
| 7 |
A' |
2608 |
2592 |
10.01 |
|
|
|
| 8 |
A' |
1434 |
1425 |
5.35 |
|
|
|
| 9 |
A' |
1429 |
1421 |
10.02 |
|
|
|
| 10 |
A' |
1414 |
1406 |
1.71 |
|
|
|
| 11 |
A' |
1407 |
1399 |
3.65 |
|
|
|
| 12 |
A' |
1355 |
1348 |
6.04 |
|
|
|
| 13 |
A' |
1323 |
1316 |
0.41 |
|
|
|
| 14 |
A' |
1283 |
1276 |
20.81 |
|
|
|
| 15 |
A' |
1195 |
1188 |
9.08 |
|
|
|
| 16 |
A' |
1163 |
1156 |
3.02 |
|
|
|
| 17 |
A' |
1104 |
1098 |
2.36 |
|
|
|
| 18 |
A' |
994 |
988 |
0.63 |
|
|
|
| 19 |
A' |
957 |
951 |
1.55 |
|
|
|
| 20 |
A' |
846 |
841 |
0.18 |
|
|
|
| 21 |
A' |
768 |
763 |
1.20 |
|
|
|
| 22 |
A' |
737 |
732 |
6.18 |
|
|
|
| 23 |
A' |
507 |
504 |
0.91 |
|
|
|
| 24 |
A' |
367 |
365 |
0.53 |
|
|
|
| 25 |
A' |
244 |
242 |
0.57 |
|
|
|
| 26 |
A' |
239 |
237 |
0.77 |
|
|
|
| 27 |
A' |
174 |
173 |
1.38 |
|
|
|
| 28 |
A" |
3050 |
3033 |
17.41 |
|
|
|
| 29 |
A" |
3048 |
3030 |
8.66 |
|
|
|
| 30 |
A" |
3039 |
3021 |
12.54 |
|
|
|
| 31 |
A" |
2997 |
2980 |
13.23 |
|
|
|
| 32 |
A" |
2963 |
2946 |
30.71 |
|
|
|
| 33 |
A" |
1418 |
1410 |
0.39 |
|
|
|
| 34 |
A" |
1405 |
1397 |
0.49 |
|
|
|
| 35 |
A" |
1339 |
1331 |
8.77 |
|
|
|
| 36 |
A" |
1309 |
1302 |
1.12 |
|
|
|
| 37 |
A" |
1261 |
1253 |
0.08 |
|
|
|
| 38 |
A" |
1169 |
1162 |
2.43 |
|
|
|
| 39 |
A" |
1053 |
1047 |
0.16 |
|
|
|
| 40 |
A" |
949 |
944 |
0.06 |
|
|
|
| 41 |
A" |
913 |
907 |
0.97 |
|
|
|
| 42 |
A" |
888 |
883 |
0.19 |
|
|
|
| 43 |
A" |
748 |
744 |
2.55 |
|
|
|
| 44 |
A" |
356 |
354 |
0.02 |
|
|
|
| 45 |
A" |
225 |
224 |
0.64 |
|
|
|
| 46 |
A" |
198 |
197 |
15.00 |
|
|
|
| 47 |
A" |
98 |
98 |
0.76 |
|
|
|
| 48 |
A" |
60 |
60 |
4.03 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 33991.0 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 33793.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.151 |
|
|
|
| 2 |
H |
0.007 |
|
|
|
| 3 |
S |
-0.114 |
|
|
|
| 4 |
H |
0.082 |
|
|
|
| 5 |
C |
-0.150 |
|
|
|
| 6 |
C |
0.005 |
|
|
|
| 7 |
C |
-0.015 |
|
|
|
| 8 |
C |
-0.015 |
|
|
|
| 9 |
H |
0.064 |
|
|
|
| 10 |
H |
0.064 |
|
|
|
| 11 |
H |
0.028 |
|
|
|
| 12 |
H |
0.028 |
|
|
|
| 13 |
H |
0.027 |
|
|
|
| 14 |
H |
0.027 |
|
|
|
| 15 |
H |
0.029 |
|
|
|
| 16 |
H |
0.029 |
|
|
|
| 17 |
H |
0.028 |
|
|
|
| 18 |
H |
0.028 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.619 |
-1.517 |
0.000 |
1.639 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-45.305 |
3.180 |
0.000 |
| y |
3.180 |
-50.322 |
0.000 |
| z |
0.000 |
0.000 |
-48.924 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.318 |
3.180 |
0.000 |
| y |
3.180 |
-3.208 |
0.000 |
| z |
0.000 |
0.000 |
-1.110 |
|
| Polar |
|---|
| 3z2-r2 | -2.221 |
|---|
| x2-y2 | 5.017 |
|---|
| xy | 3.180 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.967 |
-1.652 |
0.000 |
| y |
-1.652 |
10.989 |
0.000 |
| z |
0.000 |
0.000 |
9.696 |
<r2> (average value of r
2) Å
2
| <r2> |
314.035 |
| (<r2>)1/2 |
17.721 |