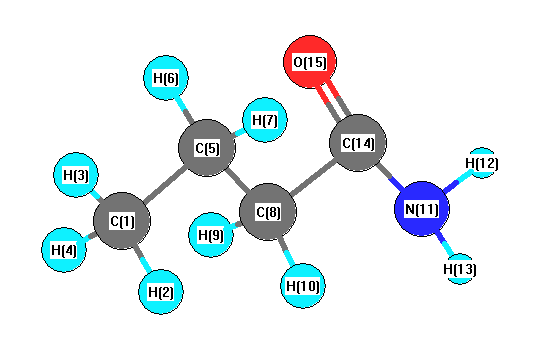
Vibrational Frequencies calculated at BLYP/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3582 |
3588 |
16.11 |
|
|
|
| 2 |
A |
3442 |
3448 |
11.01 |
|
|
|
| 3 |
A |
3016 |
3021 |
45.23 |
|
|
|
| 4 |
A |
3014 |
3018 |
54.08 |
|
|
|
| 5 |
A |
2990 |
2995 |
4.40 |
|
|
|
| 6 |
A |
2960 |
2965 |
25.76 |
|
|
|
| 7 |
A |
2950 |
2955 |
23.72 |
|
|
|
| 8 |
A |
2941 |
2946 |
32.98 |
|
|
|
| 9 |
A |
2924 |
2929 |
17.30 |
|
|
|
| 10 |
A |
1710 |
1712 |
230.19 |
|
|
|
| 11 |
A |
1535 |
1537 |
86.40 |
|
|
|
| 12 |
A |
1443 |
1445 |
3.62 |
|
|
|
| 13 |
A |
1430 |
1432 |
4.88 |
|
|
|
| 14 |
A |
1424 |
1426 |
2.21 |
|
|
|
| 15 |
A |
1400 |
1402 |
3.70 |
|
|
|
| 16 |
A |
1359 |
1361 |
0.44 |
|
|
|
| 17 |
A |
1347 |
1349 |
55.02 |
|
|
|
| 18 |
A |
1306 |
1308 |
24.50 |
|
|
|
| 19 |
A |
1272 |
1274 |
27.07 |
|
|
|
| 20 |
A |
1218 |
1219 |
54.78 |
|
|
|
| 21 |
A |
1200 |
1202 |
5.14 |
|
|
|
| 22 |
A |
1095 |
1097 |
0.84 |
|
|
|
| 23 |
A |
1071 |
1073 |
0.76 |
|
|
|
| 24 |
A |
1035 |
1036 |
0.28 |
|
|
|
| 25 |
A |
1015 |
1017 |
4.39 |
|
|
|
| 26 |
A |
900 |
901 |
1.78 |
|
|
|
| 27 |
A |
854 |
856 |
0.51 |
|
|
|
| 28 |
A |
807 |
809 |
5.15 |
|
|
|
| 29 |
A |
733 |
734 |
4.46 |
|
|
|
| 30 |
A |
646 |
647 |
8.56 |
|
|
|
| 31 |
A |
580 |
581 |
6.05 |
|
|
|
| 32 |
A |
502 |
503 |
11.57 |
|
|
|
| 33 |
A |
414 |
415 |
2.16 |
|
|
|
| 34 |
A |
333 |
333 |
1.87 |
|
|
|
| 35 |
A |
245 |
246 |
0.09 |
|
|
|
| 36 |
A |
181 |
181 |
5.71 |
|
|
|
| 37 |
A |
159 |
159 |
164.58 |
|
|
|
| 38 |
A |
88 |
88 |
1.67 |
|
|
|
| 39 |
A |
18 |
18 |
0.83 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 27568.6 cm
-1
Scaled (by 1.0016) Zero Point Vibrational Energy (zpe) 27612.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.021 |
|
|
|
| 2 |
H |
0.004 |
|
|
|
| 3 |
H |
0.001 |
|
|
|
| 4 |
H |
0.001 |
|
|
|
| 5 |
C |
0.006 |
|
|
|
| 6 |
H |
-0.005 |
|
|
|
| 7 |
H |
0.011 |
|
|
|
| 8 |
C |
0.014 |
|
|
|
| 9 |
H |
-0.010 |
|
|
|
| 10 |
H |
0.023 |
|
|
|
| 11 |
N |
-0.033 |
|
|
|
| 12 |
H |
0.081 |
|
|
|
| 13 |
H |
0.075 |
|
|
|
| 14 |
C |
0.038 |
|
|
|
| 15 |
O |
-0.228 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.439 |
-3.243 |
-0.502 |
3.311 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.234 |
-5.765 |
-1.536 |
| y |
-5.765 |
-39.018 |
0.497 |
| z |
-1.536 |
0.497 |
-37.737 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.143 |
-5.765 |
-1.536 |
| y |
-5.765 |
-3.532 |
0.497 |
| z |
-1.536 |
0.497 |
-1.611 |
|
| Polar |
|---|
| 3z2-r2 | -3.222 |
|---|
| x2-y2 | 5.784 |
|---|
| xy | -5.765 |
|---|
| xz | -1.536 |
|---|
| yz | 0.497 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.391 |
-0.435 |
-0.220 |
| y |
-0.435 |
8.577 |
0.080 |
| z |
-0.220 |
0.080 |
6.264 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |