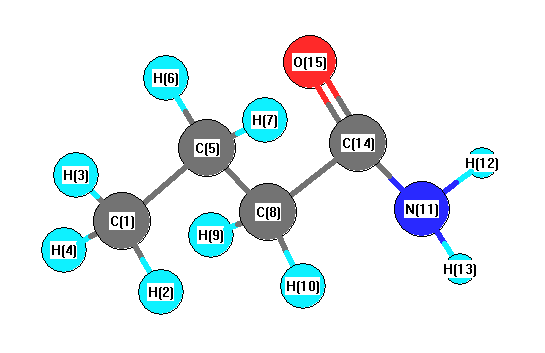
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3614 |
3593 |
21.92 |
|
|
|
| 2 |
A |
3470 |
3450 |
15.03 |
|
|
|
| 3 |
A |
3049 |
3032 |
30.53 |
|
|
|
| 4 |
A |
3042 |
3025 |
47.81 |
|
|
|
| 5 |
A |
3019 |
3002 |
0.28 |
|
|
|
| 6 |
A |
2988 |
2971 |
16.24 |
|
|
|
| 7 |
A |
2978 |
2960 |
23.20 |
|
|
|
| 8 |
A |
2960 |
2943 |
30.55 |
|
|
|
| 9 |
A |
2942 |
2925 |
14.98 |
|
|
|
| 10 |
A |
1743 |
1733 |
235.14 |
|
|
|
| 11 |
A |
1533 |
1524 |
108.04 |
|
|
|
| 12 |
A |
1431 |
1423 |
5.24 |
|
|
|
| 13 |
A |
1420 |
1412 |
6.47 |
|
|
|
| 14 |
A |
1414 |
1406 |
2.95 |
|
|
|
| 15 |
A |
1389 |
1381 |
3.97 |
|
|
|
| 16 |
A |
1358 |
1350 |
78.87 |
|
|
|
| 17 |
A |
1347 |
1340 |
2.67 |
|
|
|
| 18 |
A |
1305 |
1297 |
16.78 |
|
|
|
| 19 |
A |
1266 |
1259 |
20.89 |
|
|
|
| 20 |
A |
1207 |
1200 |
27.65 |
|
|
|
| 21 |
A |
1191 |
1184 |
2.60 |
|
|
|
| 22 |
A |
1103 |
1097 |
0.85 |
|
|
|
| 23 |
A |
1064 |
1058 |
1.36 |
|
|
|
| 24 |
A |
1054 |
1048 |
0.53 |
|
|
|
| 25 |
A |
1029 |
1023 |
2.15 |
|
|
|
| 26 |
A |
911 |
906 |
1.99 |
|
|
|
| 27 |
A |
852 |
847 |
0.76 |
|
|
|
| 28 |
A |
819 |
814 |
4.89 |
|
|
|
| 29 |
A |
729 |
724 |
6.01 |
|
|
|
| 30 |
A |
637 |
633 |
3.67 |
|
|
|
| 31 |
A |
601 |
598 |
15.95 |
|
|
|
| 32 |
A |
496 |
493 |
6.55 |
|
|
|
| 33 |
A |
414 |
411 |
1.25 |
|
|
|
| 34 |
A |
335 |
333 |
2.51 |
|
|
|
| 35 |
A |
247 |
246 |
4.22 |
|
|
|
| 36 |
A |
231 |
229 |
164.49 |
|
|
|
| 37 |
A |
180 |
179 |
8.27 |
|
|
|
| 38 |
A |
87 |
87 |
0.56 |
|
|
|
| 39 |
A |
31 |
30 |
3.81 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 27742.7 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 27581.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.051 |
|
|
|
| 2 |
H |
0.027 |
|
|
|
| 3 |
H |
0.029 |
|
|
|
| 4 |
H |
0.029 |
|
|
|
| 5 |
C |
-0.064 |
|
|
|
| 6 |
H |
0.040 |
|
|
|
| 7 |
H |
0.027 |
|
|
|
| 8 |
C |
-0.055 |
|
|
|
| 9 |
H |
0.050 |
|
|
|
| 10 |
H |
0.024 |
|
|
|
| 11 |
N |
-0.074 |
|
|
|
| 12 |
H |
0.097 |
|
|
|
| 13 |
H |
0.090 |
|
|
|
| 14 |
C |
0.066 |
|
|
|
| 15 |
O |
-0.236 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.518 |
-3.303 |
0.205 |
3.350 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.527 |
5.910 |
-0.469 |
| y |
5.910 |
-38.574 |
-0.537 |
| z |
-0.469 |
-0.537 |
-37.830 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.675 |
5.910 |
-0.469 |
| y |
5.910 |
-3.395 |
-0.537 |
| z |
-0.469 |
-0.537 |
-2.280 |
|
| Polar |
|---|
| 3z2-r2 | -4.560 |
|---|
| x2-y2 | 6.047 |
|---|
| xy | 5.910 |
|---|
| xz | -0.469 |
|---|
| yz | -0.537 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.313 |
0.430 |
0.144 |
| y |
0.430 |
8.530 |
0.034 |
| z |
0.144 |
0.034 |
6.237 |
<r2> (average value of r
2) Å
2
| <r2> |
219.518 |
| (<r2>)1/2 |
14.816 |