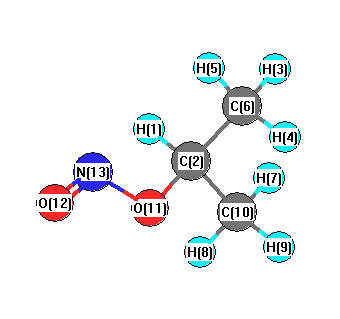
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3074 |
3056 |
20.04 |
|
|
|
| 2 |
A |
3072 |
3055 |
13.33 |
|
|
|
| 3 |
A |
3066 |
3048 |
35.48 |
|
|
|
| 4 |
A |
3061 |
3043 |
0.00 |
|
|
|
| 5 |
A |
2980 |
2963 |
14.52 |
|
|
|
| 6 |
A |
2977 |
2960 |
16.93 |
|
|
|
| 7 |
A |
2957 |
2940 |
14.79 |
|
|
|
| 8 |
A |
1735 |
1725 |
302.54 |
|
|
|
| 9 |
A |
1430 |
1422 |
5.93 |
|
|
|
| 10 |
A |
1414 |
1406 |
6.96 |
|
|
|
| 11 |
A |
1406 |
1398 |
0.05 |
|
|
|
| 12 |
A |
1400 |
1392 |
0.08 |
|
|
|
| 13 |
A |
1346 |
1338 |
6.61 |
|
|
|
| 14 |
A |
1338 |
1331 |
20.90 |
|
|
|
| 15 |
A |
1303 |
1295 |
3.89 |
|
|
|
| 16 |
A |
1297 |
1289 |
26.50 |
|
|
|
| 17 |
A |
1153 |
1147 |
12.42 |
|
|
|
| 18 |
A |
1128 |
1122 |
8.73 |
|
|
|
| 19 |
A |
1113 |
1106 |
31.81 |
|
|
|
| 20 |
A |
936 |
930 |
5.87 |
|
|
|
| 21 |
A |
909 |
903 |
0.04 |
|
|
|
| 22 |
A |
897 |
892 |
0.82 |
|
|
|
| 23 |
A |
873 |
867 |
47.71 |
|
|
|
| 24 |
A |
749 |
745 |
324.78 |
|
|
|
| 25 |
A |
553 |
550 |
49.12 |
|
|
|
| 26 |
A |
452 |
449 |
19.78 |
|
|
|
| 27 |
A |
417 |
414 |
4.49 |
|
|
|
| 28 |
A |
308 |
306 |
5.82 |
|
|
|
| 29 |
A |
273 |
271 |
0.98 |
|
|
|
| 30 |
A |
207 |
205 |
1.35 |
|
|
|
| 31 |
A |
196 |
195 |
0.05 |
|
|
|
| 32 |
A |
186 |
185 |
0.17 |
|
|
|
| 33 |
A |
56 |
55 |
0.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22129.0 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 22000.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.011 |
|
|
|
| 2 |
C |
-0.016 |
|
|
|
| 3 |
H |
0.035 |
|
|
|
| 4 |
H |
0.041 |
|
|
|
| 5 |
H |
0.036 |
|
|
|
| 6 |
C |
-0.008 |
|
|
|
| 7 |
H |
0.035 |
|
|
|
| 8 |
H |
0.036 |
|
|
|
| 9 |
H |
0.041 |
|
|
|
| 10 |
C |
-0.008 |
|
|
|
| 11 |
O |
-0.215 |
|
|
|
| 12 |
O |
-0.104 |
|
|
|
| 13 |
N |
0.117 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.029 |
0.000 |
-0.680 |
2.140 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.611 |
0.001 |
0.240 |
| y |
0.001 |
-34.787 |
-0.001 |
| z |
0.240 |
-0.001 |
-36.871 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.782 |
0.001 |
0.240 |
| y |
0.001 |
2.453 |
-0.001 |
| z |
0.240 |
-0.001 |
-0.672 |
|
| Polar |
|---|
| 3z2-r2 | -1.344 |
|---|
| x2-y2 | -2.823 |
|---|
| xy | 0.001 |
|---|
| xz | 0.240 |
|---|
| yz | -0.001 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.824 |
-0.000 |
0.050 |
| y |
-0.000 |
6.654 |
0.000 |
| z |
0.050 |
0.000 |
6.137 |
<r2> (average value of r
2) Å
2
| <r2> |
195.712 |
| (<r2>)1/2 |
13.990 |