return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H4Cl2 (1,3-dichlorobenzene)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-1149.658618
Energy at 298.15K-1149.662796
HF Energy-1148.558465
Nuclear repulsion energy449.287850
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3253 3121 0.33      
2 A1 3249 3117 2.73      
3 A1 3218 3088 4.50      
4 A1 1652 1585 21.31      
5 A1 1455 1396 13.94      
6 A1 1166 1118 20.08      
7 A1 1103 1058 9.02      
8 A1 1009 968 3.82      
9 A1 674 647 1.44      
10 A1 405 389 2.42      
11 A1 197 189 0.42      
12 A2 907 870 0.00      
13 A2 535 513 0.00      
14 A2 205 197 0.00      
15 B1 971 932 0.01      
16 B1 877 841 7.37      
17 B1 784 753 48.58      
18 B1 634 609 5.10      
19 B1 437 420 3.54      
20 B1 169 162 0.08      
21 B2 3243 3112 1.02      
22 B2 1658 1591 64.55      
23 B2 1509 1448 58.08      
24 B2 1316 1262 1.48      
25 B2 1267 1216 1.71      
26 B2 1179 1131 0.54      
27 B2 1115 1070 18.05      
28 B2 804 772 65.68      
29 B2 433 416 2.49      
30 B2 367 352 0.18      

Unscaled Zero Point Vibrational Energy (zpe) 17895.8 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 17169.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.09288 0.02824 0.02165

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 2.091
C2 0.000 1.222 1.398
C3 0.000 -1.222 1.398
C4 0.000 1.206 -0.005
C5 0.000 -1.206 -0.005
C6 0.000 0.000 -0.723
Cl7 0.000 2.718 -0.887
Cl8 0.000 -2.718 -0.887
H9 0.000 0.000 3.186
H10 0.000 2.175 1.935
H11 0.000 -2.175 1.935
H12 0.000 0.000 -1.815

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 Cl7 Cl8 H9 H10 H11 H12
C11.40461.40462.41792.41792.81344.03204.03201.09542.18052.18053.9061
C21.40462.44441.40352.80452.44802.73184.55542.16551.09363.43933.4384
C31.40462.44442.80451.40352.44804.55542.73182.16553.43931.09363.4384
C42.41791.40352.80452.41191.40331.75094.02213.41132.16873.89802.1753
C52.41792.80451.40352.41191.40334.02211.75093.41133.89802.16872.1753
C62.81342.44802.44801.40331.40332.72322.72323.90883.43443.43441.0928
Cl74.03202.73184.55541.75094.02212.72325.43644.89712.87445.64892.8723
Cl84.03204.55542.73184.02211.75092.72325.43644.89715.64892.87442.8723
H91.09542.16552.16553.41133.41133.90884.89714.89712.50902.50905.0015
H102.18051.09363.43932.16873.89803.43442.87445.64892.50904.34984.3356
H112.18053.43931.09363.89802.16873.43445.64892.87442.50904.34984.3356
H123.90613.43843.43842.17532.17531.09282.87232.87235.00154.33564.3356

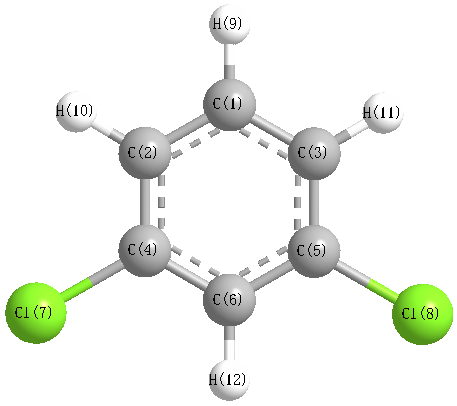
picture of 1,3-dichlorobenzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 118.864 C1 C2 H10 121.073
C1 C3 C5 118.864 C1 C3 H11 121.073
C2 C1 C3 120.944 C2 C1 H9 119.528
C2 C4 C6 121.423 C2 C4 Cl7 119.598
C3 C1 H9 119.528 C3 C5 C6 121.423
C3 C5 Cl8 119.598 C4 C2 H10 120.063
C4 C6 C5 118.482 C4 C6 H12 120.759
C5 C3 H11 120.063 C5 C6 H12 120.759
C6 C4 Cl7 118.979 C6 C5 Cl8 118.979
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability