Jump to
S1C2
Energy calculated at QCISD(T)/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -93.710562 |
| Energy at 298.15K | -93.711856 |
| HF Energy | -93.403993 |
| Nuclear repulsion energy | 27.773189 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3305 |
3150 |
|
|
|
|
| 2 |
A' |
2993 |
2852 |
|
|
|
|
| 3 |
A' |
1793 |
1709 |
|
|
|
|
| 4 |
A' |
1046 |
997 |
|
|
|
|
| 5 |
A' |
872 |
831 |
|
|
|
|
| 6 |
A" |
885 |
843 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5447.0 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 5191.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.115 |
0.645 |
0.000 |
| N2 |
0.115 |
-0.602 |
0.000 |
| H3 |
-0.684 |
1.420 |
0.000 |
| H4 |
-0.808 |
-1.079 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
H3 |
H4 |
| C1 | | 1.2476 | 1.1133 | 1.9552 |
N2 | 1.2476 | | 2.1748 | 1.0380 | H3 | 1.1133 | 2.1748 | | 2.5020 | H4 | 1.9552 | 1.0380 | 2.5020 | |
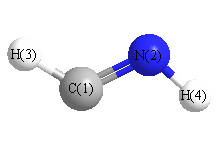 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
H4 |
117.319 |
|
H3 |
C1 |
N2 |
134.117 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD(T)/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -93.718766 |
| Energy at 298.15K | -93.720111 |
| HF Energy | -93.408850 |
| Nuclear repulsion energy | 27.695516 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3432 |
3271 |
|
|
|
|
| 2 |
A' |
3039 |
2897 |
|
|
|
|
| 3 |
A' |
1743 |
1661 |
|
|
|
|
| 4 |
A' |
1222 |
1164 |
|
|
|
|
| 5 |
A' |
918 |
875 |
|
|
|
|
| 6 |
A" |
974 |
928 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5663.6 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 5398.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.001 |
0.659 |
0.000 |
| N2 |
0.001 |
-0.599 |
0.000 |
| H3 |
0.926 |
1.275 |
0.000 |
| H4 |
-0.933 |
-1.039 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
H3 |
H4 |
| C1 | | 1.2584 | 1.1118 | 1.9383 |
N2 | 1.2584 | | 2.0904 | 1.0321 | H3 | 1.1118 | 2.0904 | | 2.9688 | H4 | 1.9383 | 1.0321 | 2.9688 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
H4 |
115.252 |
|
H3 |
C1 |
N2 |
123.645 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
