Jump to
S1C2
Energy calculated at BLYP/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -93.935592 |
| Energy at 298.15K | -93.936854 |
| HF Energy | -93.935592 |
| Nuclear repulsion energy | 27.847264 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at BLYP/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3075 |
3080 |
22.55 |
|
|
|
| 2 |
A' |
2821 |
2826 |
76.23 |
|
|
|
| 3 |
A' |
1802 |
1804 |
16.36 |
|
|
|
| 4 |
A' |
948 |
950 |
156.75 |
|
|
|
| 5 |
A' |
859 |
860 |
80.00 |
|
|
|
| 6 |
A" |
847 |
848 |
3.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5175.6 cm
-1
Scaled (by 1.0016) Zero Point Vibrational Energy (zpe) 5183.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at BLYP/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.113 |
0.644 |
0.000 |
| N2 |
0.113 |
-0.593 |
0.000 |
| H3 |
-0.692 |
1.424 |
0.000 |
| H4 |
-0.782 |
-1.138 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
H3 |
H4 |
| C1 | | 1.2364 | 1.1209 | 1.9939 |
N2 | 1.2364 | | 2.1712 | 1.0480 | H3 | 1.1209 | 2.1712 | | 2.5633 | H4 | 1.9939 | 1.0480 | 2.5633 | |
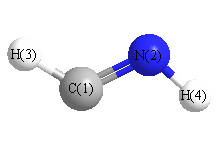 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
H4 |
121.356 |
|
H3 |
C1 |
N2 |
134.094 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.059 |
|
|
|
| 2 |
N |
-0.071 |
|
|
|
| 3 |
H |
0.039 |
|
|
|
| 4 |
H |
0.091 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.978 |
0.125 |
0.000 |
1.982 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-12.078 |
0.480 |
0.000 |
| y |
0.480 |
-11.164 |
0.000 |
| z |
0.000 |
0.000 |
-12.396 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.298 |
0.480 |
0.000 |
| y |
0.480 |
1.073 |
0.000 |
| z |
0.000 |
0.000 |
-0.775 |
|
| Polar |
|---|
| 3z2-r2 | -1.550 |
|---|
| x2-y2 | -0.914 |
|---|
| xy | 0.480 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.200 |
0.087 |
0.000 |
| y |
0.087 |
4.069 |
0.000 |
| z |
0.000 |
0.000 |
1.341 |
<r2> (average value of r
2) Å
2
| <r2> |
16.944 |
| (<r2>)1/2 |
4.116 |
Jump to
S1C1
Energy calculated at BLYP/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -93.942987 |
| Energy at 298.15K | -93.944314 |
| HF Energy | -93.942987 |
| Nuclear repulsion energy | 27.744787 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at BLYP/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3241 |
3246 |
1.50 |
|
|
|
| 2 |
A' |
2878 |
2883 |
42.61 |
|
|
|
| 3 |
A' |
1732 |
1734 |
16.00 |
|
|
|
| 4 |
A' |
1175 |
1177 |
8.66 |
|
|
|
| 5 |
A' |
892 |
894 |
188.01 |
|
|
|
| 6 |
A" |
938 |
939 |
96.21 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5427.6 cm
-1
Scaled (by 1.0016) Zero Point Vibrational Energy (zpe) 5436.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at BLYP/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.001 |
0.659 |
0.000 |
| N2 |
-0.001 |
-0.592 |
0.000 |
| H3 |
0.939 |
1.266 |
0.000 |
| H4 |
-0.923 |
-1.074 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
H3 |
H4 |
| C1 | | 1.2505 | 1.1194 | 1.9625 |
N2 | 1.2505 | | 2.0821 | 1.0400 | H3 | 1.1194 | 2.0821 | | 2.9902 | H4 | 1.9625 | 1.0400 | 2.9902 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
H4 |
117.622 |
|
H3 |
C1 |
N2 |
122.838 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.091 |
|
|
|
| 2 |
N |
-0.074 |
|
|
|
| 3 |
H |
0.053 |
|
|
|
| 4 |
H |
0.112 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.402 |
-0.110 |
0.000 |
0.416 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-11.188 |
2.821 |
0.000 |
| y |
2.821 |
-11.537 |
0.000 |
| z |
0.000 |
0.000 |
-12.423 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.792 |
2.821 |
0.000 |
| y |
2.821 |
0.268 |
0.000 |
| z |
0.000 |
0.000 |
-1.060 |
|
| Polar |
|---|
| 3z2-r2 | -2.121 |
|---|
| x2-y2 | 0.349 |
|---|
| xy | 2.821 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.423 |
0.633 |
0.000 |
| y |
0.633 |
3.718 |
0.000 |
| z |
0.000 |
0.000 |
1.392 |
<r2> (average value of r
2) Å
2
| <r2> |
16.862 |
| (<r2>)1/2 |
4.106 |
