Jump to
S1C2
Energy calculated at B3PW91/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -93.928019 |
| Energy at 298.15K | -93.929310 |
| HF Energy | -93.928019 |
| Nuclear repulsion energy | 28.088461 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3286 |
3171 |
5.99 |
|
|
|
| 2 |
A' |
2974 |
2870 |
52.93 |
|
|
|
| 3 |
A' |
1868 |
1803 |
25.59 |
|
|
|
| 4 |
A' |
1006 |
971 |
148.13 |
|
|
|
| 5 |
A' |
872 |
841 |
93.72 |
|
|
|
| 6 |
A" |
897 |
866 |
3.21 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5452.0 cm
-1
Scaled (by 0.965) Zero Point Vibrational Energy (zpe) 5261.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.113 |
0.637 |
0.000 |
| N2 |
0.113 |
-0.591 |
0.000 |
| H3 |
-0.678 |
1.416 |
0.000 |
| H4 |
-0.787 |
-1.103 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
H3 |
H4 |
| C1 | | 1.2279 | 1.1100 | 1.9584 |
N2 | 1.2279 | | 2.1572 | 1.0348 | H3 | 1.1100 | 2.1572 | | 2.5213 | H4 | 1.9584 | 1.0348 | 2.5213 | |
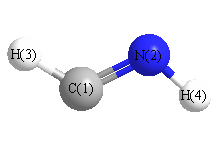 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
H4 |
119.642 |
|
H3 |
C1 |
N2 |
134.589 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.056 |
|
|
|
| 2 |
N |
-0.138 |
|
|
|
| 3 |
H |
0.076 |
|
|
|
| 4 |
H |
0.118 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.106 |
0.337 |
0.000 |
2.133 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-11.904 |
0.574 |
0.000 |
| y |
0.574 |
-10.992 |
0.000 |
| z |
0.000 |
0.000 |
-12.385 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.216 |
0.574 |
0.000 |
| y |
0.574 |
1.153 |
0.000 |
| z |
0.000 |
0.000 |
-0.938 |
|
| Polar |
|---|
| 3z2-r2 | -1.875 |
|---|
| x2-y2 | -0.913 |
|---|
| xy | 0.574 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.095 |
0.061 |
0.000 |
| y |
0.061 |
3.845 |
0.000 |
| z |
0.000 |
0.000 |
1.337 |
<r2> (average value of r
2) Å
2
| <r2> |
16.689 |
| (<r2>)1/2 |
4.085 |
Jump to
S1C1
Energy calculated at B3PW91/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -93.935388 |
| Energy at 298.15K | -93.936727 |
| HF Energy | -93.935388 |
| Nuclear repulsion energy | 28.014114 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3415 |
3295 |
5.59 |
|
|
|
| 2 |
A' |
3020 |
2914 |
29.12 |
|
|
|
| 3 |
A' |
1819 |
1755 |
24.96 |
|
|
|
| 4 |
A' |
1196 |
1154 |
9.08 |
|
|
|
| 5 |
A' |
904 |
872 |
212.62 |
|
|
|
| 6 |
A" |
969 |
935 |
109.03 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5660.7 cm
-1
Scaled (by 0.965) Zero Point Vibrational Energy (zpe) 5462.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.000 |
0.650 |
0.000 |
| N2 |
-0.000 |
-0.588 |
0.000 |
| H3 |
0.917 |
1.273 |
0.000 |
| H4 |
-0.916 |
-1.057 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
H3 |
H4 |
| C1 | | 1.2383 | 1.1088 | 1.9377 |
N2 | 1.2383 | | 2.0747 | 1.0290 | H3 | 1.1088 | 2.0747 | | 2.9649 | H4 | 1.9377 | 1.0290 | 2.9649 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
H4 |
117.133 |
|
H3 |
C1 |
N2 |
124.153 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.092 |
|
|
|
| 2 |
N |
-0.135 |
|
|
|
| 3 |
H |
0.090 |
|
|
|
| 4 |
H |
0.137 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.452 |
0.095 |
0.000 |
0.462 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-11.043 |
2.972 |
0.000 |
| y |
2.972 |
-11.318 |
0.000 |
| z |
0.000 |
0.000 |
-12.414 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.823 |
2.972 |
0.000 |
| y |
2.972 |
0.411 |
0.000 |
| z |
0.000 |
0.000 |
-1.233 |
|
| Polar |
|---|
| 3z2-r2 | -2.467 |
|---|
| x2-y2 | 0.275 |
|---|
| xy | 2.972 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.301 |
0.514 |
0.000 |
| y |
0.514 |
3.649 |
0.000 |
| z |
0.000 |
0.000 |
1.388 |
<r2> (average value of r
2) Å
2
| <r2> |
16.616 |
| (<r2>)1/2 |
4.076 |
