Jump to
S1C2
Energy calculated at QCISD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -93.700736 |
| Energy at 298.15K | -93.702043 |
| HF Energy | -93.404253 |
| Nuclear repulsion energy | 27.842131 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3343 |
3207 |
4.63 |
|
|
|
| 2 |
A' |
3013 |
2890 |
53.78 |
|
|
|
| 3 |
A' |
1822 |
1748 |
20.71 |
|
|
|
| 4 |
A' |
1063 |
1020 |
134.36 |
|
|
|
| 5 |
A' |
888 |
852 |
87.61 |
|
|
|
| 6 |
A" |
908 |
871 |
3.41 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5518.3 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 5294.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.115 |
0.644 |
0.000 |
| N2 |
0.115 |
-0.601 |
0.000 |
| H3 |
-0.685 |
1.416 |
0.000 |
| H4 |
-0.806 |
-1.075 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
H3 |
H4 |
| C1 | | 1.2443 | 1.1119 | 1.9496 |
N2 | 1.2443 | | 2.1696 | 1.0355 | H3 | 1.1119 | 2.1696 | | 2.4939 | H4 | 1.9496 | 1.0355 | 2.4939 | |
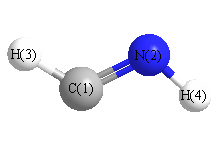 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
H4 |
117.255 |
|
H3 |
C1 |
N2 |
134.002 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -93.708582 |
| Energy at 298.15K | -93.709935 |
| HF Energy | -93.409240 |
| Nuclear repulsion energy | 27.770074 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3458 |
3317 |
4.49 |
|
|
|
| 2 |
A' |
3062 |
2938 |
32.26 |
|
|
|
| 3 |
A' |
1781 |
1708 |
23.95 |
|
|
|
| 4 |
A' |
1229 |
1180 |
9.63 |
|
|
|
| 5 |
A' |
931 |
893 |
199.31 |
|
|
|
| 6 |
A" |
991 |
951 |
98.83 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5726.1 cm
-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 5493.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.001 |
0.657 |
0.000 |
| N2 |
0.001 |
-0.598 |
0.000 |
| H3 |
0.919 |
1.280 |
0.000 |
| H4 |
-0.931 |
-1.037 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
H3 |
H4 |
| C1 | | 1.2542 | 1.1099 | 1.9331 |
N2 | 1.2542 | | 2.0900 | 1.0304 | H3 | 1.1099 | 2.0900 | | 2.9651 | H4 | 1.9331 | 1.0304 | 2.9651 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
H4 |
115.240 |
|
H3 |
C1 |
N2 |
124.163 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
