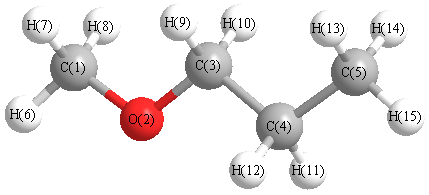
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3056 |
3038 |
26.10 |
|
|
|
| 2 |
A' |
3053 |
3036 |
27.65 |
|
|
|
| 3 |
A' |
2974 |
2957 |
27.94 |
|
|
|
| 4 |
A' |
2967 |
2950 |
25.57 |
|
|
|
| 5 |
A' |
2882 |
2865 |
87.65 |
|
|
|
| 6 |
A' |
2851 |
2834 |
46.50 |
|
|
|
| 7 |
A' |
1455 |
1446 |
1.58 |
|
|
|
| 8 |
A' |
1433 |
1424 |
6.72 |
|
|
|
| 9 |
A' |
1426 |
1417 |
7.11 |
|
|
|
| 10 |
A' |
1417 |
1409 |
2.70 |
|
|
|
| 11 |
A' |
1405 |
1397 |
0.03 |
|
|
|
| 12 |
A' |
1362 |
1354 |
21.61 |
|
|
|
| 13 |
A' |
1348 |
1340 |
1.30 |
|
|
|
| 14 |
A' |
1272 |
1264 |
5.45 |
|
|
|
| 15 |
A' |
1178 |
1171 |
41.35 |
|
|
|
| 16 |
A' |
1141 |
1134 |
144.81 |
|
|
|
| 17 |
A' |
1091 |
1085 |
6.91 |
|
|
|
| 18 |
A' |
1046 |
1040 |
2.33 |
|
|
|
| 19 |
A' |
968 |
962 |
22.55 |
|
|
|
| 20 |
A' |
885 |
879 |
7.15 |
|
|
|
| 21 |
A' |
428 |
426 |
0.62 |
|
|
|
| 22 |
A' |
396 |
393 |
2.44 |
|
|
|
| 23 |
A' |
184 |
183 |
1.04 |
|
|
|
| 24 |
A" |
3044 |
3026 |
47.78 |
|
|
|
| 25 |
A" |
3014 |
2996 |
7.57 |
|
|
|
| 26 |
A" |
2927 |
2910 |
65.67 |
|
|
|
| 27 |
A" |
2878 |
2862 |
81.20 |
|
|
|
| 28 |
A" |
1422 |
1414 |
6.70 |
|
|
|
| 29 |
A" |
1399 |
1391 |
5.87 |
|
|
|
| 30 |
A" |
1257 |
1250 |
0.20 |
|
|
|
| 31 |
A" |
1208 |
1201 |
1.93 |
|
|
|
| 32 |
A" |
1145 |
1139 |
4.73 |
|
|
|
| 33 |
A" |
1115 |
1108 |
0.24 |
|
|
|
| 34 |
A" |
868 |
863 |
2.10 |
|
|
|
| 35 |
A" |
748 |
744 |
2.47 |
|
|
|
| 36 |
A" |
240 |
239 |
2.54 |
|
|
|
| 37 |
A" |
235 |
234 |
0.01 |
|
|
|
| 38 |
A" |
112 |
111 |
1.94 |
|
|
|
| 39 |
A" |
102 |
101 |
0.63 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28963.0 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 28795.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.074 |
|
|
|
| 2 |
O |
-0.278 |
|
|
|
| 3 |
C |
0.107 |
|
|
|
| 4 |
C |
-0.073 |
|
|
|
| 5 |
C |
-0.070 |
|
|
|
| 6 |
H |
0.037 |
|
|
|
| 7 |
H |
0.015 |
|
|
|
| 8 |
H |
0.015 |
|
|
|
| 9 |
H |
0.007 |
|
|
|
| 10 |
H |
0.007 |
|
|
|
| 11 |
H |
0.030 |
|
|
|
| 12 |
H |
0.030 |
|
|
|
| 13 |
H |
0.034 |
|
|
|
| 14 |
H |
0.034 |
|
|
|
| 15 |
H |
0.032 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.203 |
0.866 |
0.000 |
0.890 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.283 |
-1.855 |
0.000 |
| y |
-1.855 |
-33.582 |
0.000 |
| z |
0.000 |
0.000 |
-33.338 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.177 |
-1.855 |
0.000 |
| y |
-1.855 |
-1.271 |
0.000 |
| z |
0.000 |
0.000 |
-0.906 |
|
| Polar |
|---|
| 3z2-r2 | -1.811 |
|---|
| x2-y2 | 2.299 |
|---|
| xy | -1.855 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.573 |
-0.528 |
0.000 |
| y |
-0.528 |
7.260 |
0.000 |
| z |
0.000 |
0.000 |
6.934 |
<r2> (average value of r
2) Å
2
| <r2> |
182.602 |
| (<r2>)1/2 |
13.513 |