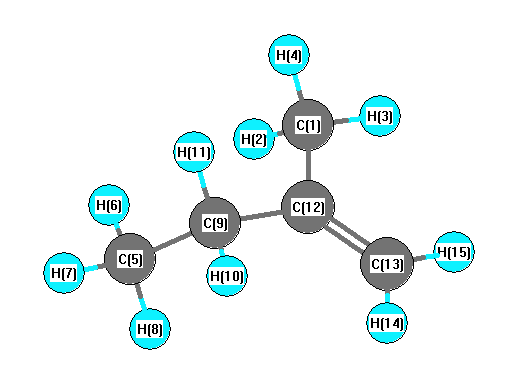
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3161 |
3143 |
18.18 |
|
|
|
| 2 |
A |
3067 |
3049 |
10.84 |
|
|
|
| 3 |
A |
3064 |
3046 |
15.28 |
|
|
|
| 4 |
A |
3057 |
3039 |
28.05 |
|
|
|
| 5 |
A |
3054 |
3036 |
30.53 |
|
|
|
| 6 |
A |
3016 |
2998 |
17.77 |
|
|
|
| 7 |
A |
3007 |
2990 |
13.92 |
|
|
|
| 8 |
A |
2973 |
2956 |
28.31 |
|
|
|
| 9 |
A |
2956 |
2939 |
25.50 |
|
|
|
| 10 |
A |
2944 |
2927 |
28.72 |
|
|
|
| 11 |
A |
1674 |
1665 |
16.46 |
|
|
|
| 12 |
A |
1432 |
1424 |
3.47 |
|
|
|
| 13 |
A |
1421 |
1413 |
4.41 |
|
|
|
| 14 |
A |
1416 |
1407 |
11.71 |
|
|
|
| 15 |
A |
1405 |
1397 |
9.74 |
|
|
|
| 16 |
A |
1393 |
1385 |
3.01 |
|
|
|
| 17 |
A |
1373 |
1365 |
0.17 |
|
|
|
| 18 |
A |
1340 |
1333 |
5.21 |
|
|
|
| 19 |
A |
1339 |
1331 |
0.75 |
|
|
|
| 20 |
A |
1296 |
1289 |
1.49 |
|
|
|
| 21 |
A |
1259 |
1251 |
2.54 |
|
|
|
| 22 |
A |
1223 |
1216 |
1.68 |
|
|
|
| 23 |
A |
1068 |
1062 |
0.92 |
|
|
|
| 24 |
A |
1054 |
1048 |
4.39 |
|
|
|
| 25 |
A |
1013 |
1007 |
0.89 |
|
|
|
| 26 |
A |
995 |
989 |
1.34 |
|
|
|
| 27 |
A |
954 |
949 |
1.52 |
|
|
|
| 28 |
A |
922 |
917 |
1.92 |
|
|
|
| 29 |
A |
872 |
867 |
32.02 |
|
|
|
| 30 |
A |
778 |
774 |
1.81 |
|
|
|
| 31 |
A |
763 |
758 |
1.54 |
|
|
|
| 32 |
A |
697 |
693 |
0.13 |
|
|
|
| 33 |
A |
519 |
516 |
6.28 |
|
|
|
| 34 |
A |
417 |
414 |
1.67 |
|
|
|
| 35 |
A |
389 |
387 |
0.65 |
|
|
|
| 36 |
A |
254 |
253 |
0.19 |
|
|
|
| 37 |
A |
220 |
218 |
0.42 |
|
|
|
| 38 |
A |
183 |
182 |
0.39 |
|
|
|
| 39 |
A |
66 |
66 |
0.07 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29016.3 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 28848.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.005 |
|
|
|
| 2 |
H |
0.045 |
|
|
|
| 3 |
H |
0.028 |
|
|
|
| 4 |
H |
0.041 |
|
|
|
| 5 |
C |
-0.062 |
|
|
|
| 6 |
H |
0.034 |
|
|
|
| 7 |
H |
0.030 |
|
|
|
| 8 |
H |
0.037 |
|
|
|
| 9 |
C |
-0.011 |
|
|
|
| 10 |
H |
0.013 |
|
|
|
| 11 |
H |
0.027 |
|
|
|
| 12 |
C |
-0.214 |
|
|
|
| 13 |
C |
0.011 |
|
|
|
| 14 |
H |
0.009 |
|
|
|
| 15 |
H |
0.008 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.385 |
0.416 |
0.122 |
0.580 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.115 |
0.599 |
-0.749 |
| y |
0.599 |
-32.596 |
0.126 |
| z |
-0.749 |
0.126 |
-34.447 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.407 |
0.599 |
-0.749 |
| y |
0.599 |
1.185 |
0.126 |
| z |
-0.749 |
0.126 |
-1.592 |
|
| Polar |
|---|
| 3z2-r2 | -3.184 |
|---|
| x2-y2 | -0.519 |
|---|
| xy | 0.599 |
|---|
| xz | -0.749 |
|---|
| yz | 0.126 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.976 |
-0.799 |
-0.704 |
| y |
-0.799 |
9.476 |
0.327 |
| z |
-0.704 |
0.327 |
6.619 |
<r2> (average value of r
2) Å
2
| <r2> |
145.111 |
| (<r2>)1/2 |
12.046 |