return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H12 (2,3-dimethylbut-1-ene)

using model chemistry: QCISD(T)/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A'
Energy calculated at QCISD(T)/cc-pVDZ
 hartrees
Energy at 0K-235.159524
Energy at 298.15K-235.171757
HF Energy-234.192446
Nuclear repulsion energy241.680449
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3233 3081        
2 A' 3140 2993        
3 A' 3135 2988        
4 A' 3124 2978        
5 A' 3118 2972        
6 A' 3040 2898        
7 A' 3036 2893        
8 A' 3033 2891        
9 A' 1699 1619        
10 A' 1500 1429        
11 A' 1493 1423        
12 A' 1482 1412        
13 A' 1443 1376        
14 A' 1409 1343        
15 A' 1404 1338        
16 A' 1340 1277        
17 A' 1311 1250        
18 A' 1187 1132        
19 A' 1105 1053        
20 A' 997 951        
21 A' 975 929        
22 A' 910 867        
23 A' 735 701        
24 A' 515 490        
25 A' 432 412        
26 A' 326 311        
27 A' 277 264        
28 A' 263 251        
29 A" 3122 2976        
30 A" 3115 2969        
31 A" 3106 2961        
32 A" 3032 2890        
33 A" 1486 1417        
34 A" 1476 1407        
35 A" 1467 1398        
36 A" 1390 1324        
37 A" 1339 1276        
38 A" 1137 1084        
39 A" 1052 1003        
40 A" 966 921        
41 A" 919 876        
42 A" 890 848        
43 A" 726 692        
44 A" 538 513        
45 A" 266 254        
46 A" 211 201        
47 A" 167 159        
48 A" 32 30        

Unscaled Zero Point Vibrational Energy (zpe) 36048.6 cm-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 34358.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD(T)/cc-pVDZ
ABC
0.14547 0.08211 0.08105

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD(T)/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.965 -1.833 0.000
C2 -0.063 -0.821 0.000
C3 -0.523 0.636 0.000
C4 1.431 -1.098 0.000
C5 -0.063 1.385 1.269
C6 -0.063 1.385 -1.269
H7 -0.647 -2.885 0.000
H8 -2.045 -1.634 0.000
H9 -1.632 0.622 0.000
H10 1.630 -2.185 0.000
H11 1.922 -0.659 -0.891
H12 1.922 -0.659 0.891
H13 -0.492 2.405 1.292
H14 -0.386 0.852 2.183
H15 1.040 1.485 1.299
H16 -0.492 2.405 -1.292
H17 -0.386 0.852 -2.183
H18 1.040 1.485 -1.299

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18
C11.35592.50872.50633.57483.57481.09891.09892.54492.61813.24143.24144.45623.50924.08904.45623.50924.0890
C21.35591.52821.51922.54462.54462.14532.14262.13252.17332.18152.18153.50172.76992.86753.50172.76992.8675
C32.50871.52822.61231.54321.54323.52372.73301.10963.54822.90672.90672.19062.19842.20252.19062.19842.2025
C42.50631.51922.61233.16283.16282.74133.51763.51331.10491.10761.10764.19963.44562.91754.19963.44562.9175
C53.57482.54461.54323.16282.53734.49273.82752.15744.14893.57472.87391.10741.10681.10772.78943.50792.7963
C63.57482.54461.54323.16282.53734.49273.82752.15744.14892.87393.57472.78943.50792.79631.10741.10681.1077
H71.09892.14533.52372.74134.49274.49271.87683.64352.38223.51443.51445.44824.33644.86165.44824.33644.8616
H81.09892.14262.73303.51763.82753.82751.87682.29333.71624.18144.18144.51583.70124.57534.51583.70124.5753
H92.54492.13251.10963.51332.15742.15743.64352.29334.30353.88203.88202.47942.52463.09412.47942.52463.0941
H102.61812.17333.54821.10494.14894.14892.38223.71624.30351.79051.79055.21884.24893.93735.21884.24893.9373
H113.24142.18152.90671.10763.57472.87393.51444.18143.88201.79051.78124.46984.13063.18913.92153.04692.3543
H123.24142.18152.90671.10762.87393.57473.51444.18143.88201.79051.78123.92153.04692.35434.46984.13063.1891
H134.45623.50172.19064.19961.10742.78945.44824.51582.47945.21884.46983.92151.79401.78702.58323.80783.1472
H143.50922.76992.19843.44561.10683.50794.33643.70122.52464.24894.13063.04691.79401.79343.80784.36693.8161
H154.08902.86752.20252.91751.10772.79634.86164.57533.09413.93733.18912.35431.78701.79343.14723.81612.5982
H164.45623.50172.19064.19962.78941.10745.44824.51582.47945.21883.92154.46982.58323.80783.14721.79401.7870
H173.50922.76992.19843.44563.50791.10684.33643.70122.52464.24893.04694.13063.80784.36693.81611.79401.7934
H184.08902.86752.20252.91752.79631.10774.86164.57533.09413.93732.35433.18913.14723.81612.59821.78701.7934

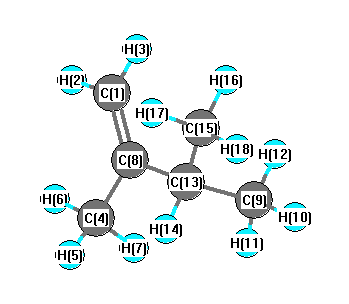
picture of 2,3-dimethylbut-1-ene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 120.767 C1 C2 C4 121.220
C2 C1 H7 121.487 C2 C1 H8 121.227
C2 C3 C5 111.884 C2 C3 C6 111.884
C2 C3 H9 106.814 C2 C4 H10 110.839
C2 C4 H11 111.337 C2 C4 H12 111.337
C3 C2 C4 118.013 C3 C5 H13 110.393
C3 C5 H14 111.038 C3 C5 H15 111.313
C3 C6 H16 110.393 C3 C6 H17 111.038
C3 C6 H18 111.313 C5 C3 C6 110.587
C5 C3 H9 107.710 C6 C3 H9 107.710
H7 C1 H8 117.286 H10 C4 H11 108.057
H10 C4 H12 108.057 H11 C4 H12 107.045
H13 C5 H14 108.240 H13 C5 H15 107.558
H14 C5 H15 108.169 H16 C6 H17 108.240
H16 C6 H18 107.558 H17 C6 H18 108.169
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability