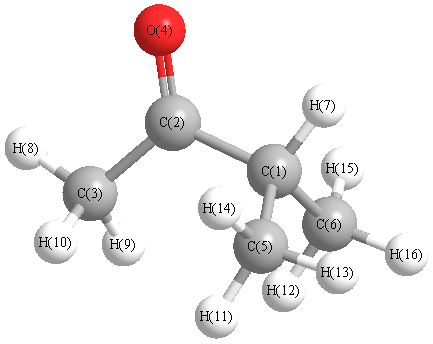
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3101 |
3083 |
7.52 |
|
|
|
| 2 |
A |
3060 |
3043 |
16.40 |
|
|
|
| 3 |
A |
3051 |
3034 |
28.92 |
|
|
|
| 4 |
A |
3049 |
3031 |
1.48 |
|
|
|
| 5 |
A |
3048 |
3031 |
41.75 |
|
|
|
| 6 |
A |
3043 |
3026 |
3.69 |
|
|
|
| 7 |
A |
2989 |
2971 |
6.51 |
|
|
|
| 8 |
A |
2975 |
2958 |
4.06 |
|
|
|
| 9 |
A |
2967 |
2950 |
16.12 |
|
|
|
| 10 |
A |
2965 |
2948 |
24.41 |
|
|
|
| 11 |
A |
1741 |
1731 |
148.94 |
|
|
|
| 12 |
A |
1434 |
1425 |
18.26 |
|
|
|
| 13 |
A |
1425 |
1417 |
7.80 |
|
|
|
| 14 |
A |
1412 |
1404 |
2.47 |
|
|
|
| 15 |
A |
1408 |
1400 |
0.84 |
|
|
|
| 16 |
A |
1391 |
1383 |
9.01 |
|
|
|
| 17 |
A |
1380 |
1372 |
11.45 |
|
|
|
| 18 |
A |
1346 |
1338 |
4.19 |
|
|
|
| 19 |
A |
1324 |
1317 |
4.15 |
|
|
|
| 20 |
A |
1308 |
1300 |
52.36 |
|
|
|
| 21 |
A |
1287 |
1279 |
2.12 |
|
|
|
| 22 |
A |
1264 |
1257 |
1.95 |
|
|
|
| 23 |
A |
1208 |
1201 |
40.17 |
|
|
|
| 24 |
A |
1136 |
1129 |
2.32 |
|
|
|
| 25 |
A |
1098 |
1091 |
0.68 |
|
|
|
| 26 |
A |
1045 |
1039 |
23.25 |
|
|
|
| 27 |
A |
972 |
966 |
0.94 |
|
|
|
| 28 |
A |
927 |
921 |
1.55 |
|
|
|
| 29 |
A |
901 |
895 |
16.05 |
|
|
|
| 30 |
A |
894 |
888 |
4.32 |
|
|
|
| 31 |
A |
881 |
876 |
5.20 |
|
|
|
| 32 |
A |
706 |
702 |
6.36 |
|
|
|
| 33 |
A |
573 |
570 |
0.30 |
|
|
|
| 34 |
A |
510 |
507 |
6.54 |
|
|
|
| 35 |
A |
472 |
469 |
3.56 |
|
|
|
| 36 |
A |
322 |
320 |
1.11 |
|
|
|
| 37 |
A |
268 |
266 |
1.39 |
|
|
|
| 38 |
A |
248 |
247 |
0.31 |
|
|
|
| 39 |
A |
233 |
232 |
0.11 |
|
|
|
| 40 |
A |
206 |
205 |
0.25 |
|
|
|
| 41 |
A |
142 |
141 |
0.31 |
|
|
|
| 42 |
A |
42 |
42 |
2.48 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29874.7 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 29701.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.183 |
|
|
|
| 2 |
C |
0.018 |
|
|
|
| 3 |
H |
0.062 |
|
|
|
| 4 |
H |
0.046 |
|
|
|
| 5 |
H |
0.048 |
|
|
|
| 6 |
C |
-0.037 |
|
|
|
| 7 |
H |
0.014 |
|
|
|
| 8 |
C |
-0.131 |
|
|
|
| 9 |
H |
0.037 |
|
|
|
| 10 |
H |
0.043 |
|
|
|
| 11 |
H |
0.030 |
|
|
|
| 12 |
H |
0.031 |
|
|
|
| 13 |
H |
0.036 |
|
|
|
| 14 |
H |
0.035 |
|
|
|
| 15 |
C |
-0.024 |
|
|
|
| 16 |
C |
-0.026 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.754 |
1.511 |
1.156 |
2.588 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.608 |
3.116 |
2.399 |
| y |
3.116 |
-38.687 |
-1.234 |
| z |
2.399 |
-1.234 |
-38.490 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.020 |
3.116 |
2.399 |
| y |
3.116 |
0.862 |
-1.234 |
| z |
2.399 |
-1.234 |
1.158 |
|
| Polar |
|---|
| 3z2-r2 | 2.315 |
|---|
| x2-y2 | -1.921 |
|---|
| xy | 3.116 |
|---|
| xz | 2.399 |
|---|
| yz | -1.234 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.746 |
0.037 |
0.067 |
| y |
0.037 |
8.843 |
0.372 |
| z |
0.067 |
0.372 |
8.130 |
<r2> (average value of r
2) Å
2
| <r2> |
181.877 |
| (<r2>)1/2 |
13.486 |