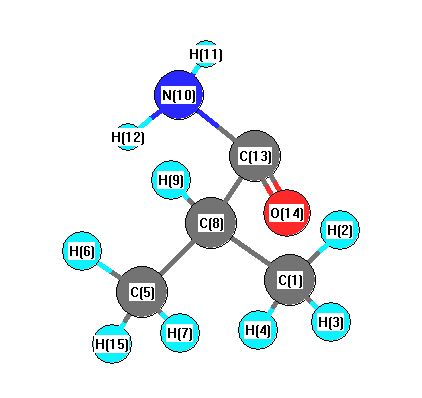
Vibrational Frequencies calculated at B3PW91/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3738 |
3608 |
36.09 |
|
|
|
| 2 |
A |
3588 |
3463 |
32.27 |
|
|
|
| 3 |
A |
3148 |
3038 |
14.82 |
|
|
|
| 4 |
A |
3138 |
3028 |
11.53 |
|
|
|
| 5 |
A |
3130 |
3020 |
27.40 |
|
|
|
| 6 |
A |
3125 |
3016 |
22.66 |
|
|
|
| 7 |
A |
3048 |
2941 |
20.14 |
|
|
|
| 8 |
A |
3042 |
2936 |
29.11 |
|
|
|
| 9 |
A |
3029 |
2923 |
36.43 |
|
|
|
| 10 |
A |
1806 |
1743 |
265.38 |
|
|
|
| 11 |
A |
1586 |
1530 |
128.98 |
|
|
|
| 12 |
A |
1487 |
1435 |
18.59 |
|
|
|
| 13 |
A |
1468 |
1417 |
10.30 |
|
|
|
| 14 |
A |
1460 |
1409 |
1.36 |
|
|
|
| 15 |
A |
1451 |
1401 |
0.92 |
|
|
|
| 16 |
A |
1409 |
1360 |
56.16 |
|
|
|
| 17 |
A |
1387 |
1338 |
4.58 |
|
|
|
| 18 |
A |
1371 |
1323 |
7.24 |
|
|
|
| 19 |
A |
1325 |
1278 |
6.39 |
|
|
|
| 20 |
A |
1272 |
1228 |
68.67 |
|
|
|
| 21 |
A |
1187 |
1145 |
3.02 |
|
|
|
| 22 |
A |
1134 |
1095 |
0.43 |
|
|
|
| 23 |
A |
1119 |
1080 |
5.45 |
|
|
|
| 24 |
A |
1053 |
1016 |
5.02 |
|
|
|
| 25 |
A |
973 |
939 |
0.45 |
|
|
|
| 26 |
A |
928 |
895 |
3.49 |
|
|
|
| 27 |
A |
922 |
890 |
1.95 |
|
|
|
| 28 |
A |
776 |
748 |
3.90 |
|
|
|
| 29 |
A |
761 |
734 |
4.73 |
|
|
|
| 30 |
A |
611 |
590 |
6.44 |
|
|
|
| 31 |
A |
594 |
573 |
11.20 |
|
|
|
| 32 |
A |
471 |
455 |
2.84 |
|
|
|
| 33 |
A |
326 |
315 |
0.58 |
|
|
|
| 34 |
A |
293 |
283 |
4.35 |
|
|
|
| 35 |
A |
242 |
234 |
5.52 |
|
|
|
| 36 |
A |
235 |
227 |
1.06 |
|
|
|
| 37 |
A |
213 |
206 |
0.47 |
|
|
|
| 38 |
A |
113 |
109 |
174.16 |
|
|
|
| 39 |
A |
18 |
18 |
8.41 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28489.0 cm
-1
Scaled (by 0.965) Zero Point Vibrational Energy (zpe) 27491.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.026 |
|
|
|
| 2 |
H |
0.041 |
|
|
|
| 3 |
H |
0.061 |
|
|
|
| 4 |
H |
0.032 |
|
|
|
| 5 |
C |
-0.030 |
|
|
|
| 6 |
H |
0.036 |
|
|
|
| 7 |
H |
0.059 |
|
|
|
| 8 |
C |
-0.200 |
|
|
|
| 9 |
H |
0.007 |
|
|
|
| 10 |
N |
-0.117 |
|
|
|
| 11 |
H |
0.113 |
|
|
|
| 12 |
H |
0.106 |
|
|
|
| 13 |
C |
0.156 |
|
|
|
| 14 |
O |
-0.279 |
|
|
|
| 15 |
H |
0.041 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.334 |
2.347 |
-2.622 |
3.535 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.440 |
3.219 |
-3.380 |
| y |
3.219 |
-38.031 |
0.397 |
| z |
-3.380 |
0.397 |
-38.278 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.714 |
3.219 |
-3.380 |
| y |
3.219 |
-2.672 |
0.397 |
| z |
-3.380 |
0.397 |
-3.043 |
|
| Polar |
|---|
| 3z2-r2 | -6.085 |
|---|
| x2-y2 | 5.590 |
|---|
| xy | 3.219 |
|---|
| xz | -3.380 |
|---|
| yz | 0.397 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
172.556 |
| (<r2>)1/2 |
13.136 |