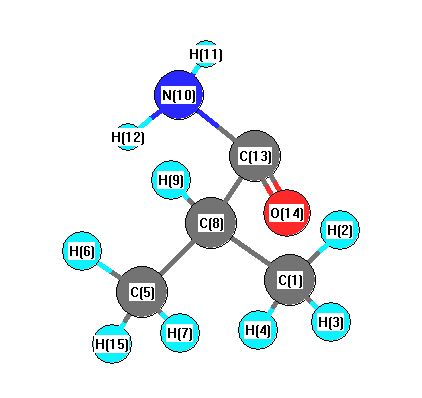
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3625 |
3604 |
23.36 |
|
|
|
| 2 |
A |
3476 |
3456 |
16.65 |
|
|
|
| 3 |
A |
3069 |
3051 |
15.22 |
|
|
|
| 4 |
A |
3061 |
3043 |
10.38 |
|
|
|
| 5 |
A |
3057 |
3039 |
25.76 |
|
|
|
| 6 |
A |
3052 |
3034 |
23.38 |
|
|
|
| 7 |
A |
2975 |
2958 |
21.30 |
|
|
|
| 8 |
A |
2969 |
2952 |
27.84 |
|
|
|
| 9 |
A |
2949 |
2932 |
36.65 |
|
|
|
| 10 |
A |
1740 |
1730 |
225.20 |
|
|
|
| 11 |
A |
1527 |
1518 |
111.91 |
|
|
|
| 12 |
A |
1439 |
1430 |
15.15 |
|
|
|
| 13 |
A |
1420 |
1412 |
9.95 |
|
|
|
| 14 |
A |
1412 |
1404 |
1.12 |
|
|
|
| 15 |
A |
1402 |
1394 |
0.66 |
|
|
|
| 16 |
A |
1355 |
1347 |
52.27 |
|
|
|
| 17 |
A |
1335 |
1327 |
4.62 |
|
|
|
| 18 |
A |
1320 |
1313 |
7.37 |
|
|
|
| 19 |
A |
1278 |
1271 |
7.02 |
|
|
|
| 20 |
A |
1228 |
1221 |
57.97 |
|
|
|
| 21 |
A |
1149 |
1142 |
2.10 |
|
|
|
| 22 |
A |
1100 |
1094 |
0.41 |
|
|
|
| 23 |
A |
1080 |
1074 |
4.83 |
|
|
|
| 24 |
A |
1016 |
1010 |
4.14 |
|
|
|
| 25 |
A |
944 |
939 |
0.36 |
|
|
|
| 26 |
A |
901 |
896 |
3.89 |
|
|
|
| 27 |
A |
894 |
888 |
1.81 |
|
|
|
| 28 |
A |
746 |
742 |
2.23 |
|
|
|
| 29 |
A |
736 |
732 |
6.05 |
|
|
|
| 30 |
A |
591 |
588 |
6.76 |
|
|
|
| 31 |
A |
582 |
578 |
9.30 |
|
|
|
| 32 |
A |
457 |
455 |
2.71 |
|
|
|
| 33 |
A |
319 |
317 |
0.53 |
|
|
|
| 34 |
A |
288 |
286 |
3.47 |
|
|
|
| 35 |
A |
239 |
237 |
5.74 |
|
|
|
| 36 |
A |
230 |
228 |
1.76 |
|
|
|
| 37 |
A |
210 |
209 |
0.95 |
|
|
|
| 38 |
A |
127 |
127 |
157.58 |
|
|
|
| 39 |
A |
17 |
17 |
7.36 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 27657.0 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 27496.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.001 |
|
|
|
| 2 |
H |
0.028 |
|
|
|
| 3 |
H |
0.047 |
|
|
|
| 4 |
H |
0.020 |
|
|
|
| 5 |
C |
-0.003 |
|
|
|
| 6 |
H |
0.023 |
|
|
|
| 7 |
H |
0.046 |
|
|
|
| 8 |
C |
-0.164 |
|
|
|
| 9 |
H |
-0.011 |
|
|
|
| 10 |
N |
-0.067 |
|
|
|
| 11 |
H |
0.097 |
|
|
|
| 12 |
H |
0.092 |
|
|
|
| 13 |
C |
0.103 |
|
|
|
| 14 |
O |
-0.241 |
|
|
|
| 15 |
H |
0.029 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.457 |
2.271 |
-2.515 |
3.419 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.735 |
3.194 |
-3.265 |
| y |
3.194 |
-38.217 |
0.278 |
| z |
-3.265 |
0.278 |
-38.600 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.674 |
3.194 |
-3.265 |
| y |
3.194 |
-2.549 |
0.278 |
| z |
-3.265 |
0.278 |
-3.124 |
|
| Polar |
|---|
| 3z2-r2 | -6.249 |
|---|
| x2-y2 | 5.482 |
|---|
| xy | 3.194 |
|---|
| xz | -3.265 |
|---|
| yz | 0.278 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.439 |
0.323 |
-0.299 |
| y |
0.323 |
7.821 |
-0.596 |
| z |
-0.299 |
-0.596 |
7.654 |
<r2> (average value of r
2) Å
2
| <r2> |
174.289 |
| (<r2>)1/2 |
13.202 |