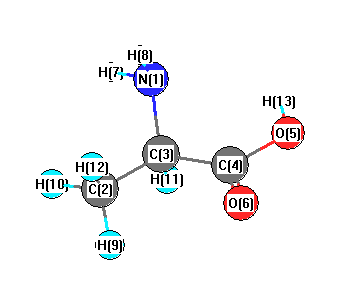
Vibrational Frequencies calculated at B3PW91/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3582 |
3457 |
8.43 |
|
|
|
| 2 |
A |
3493 |
3371 |
0.50 |
|
|
|
| 3 |
A |
3381 |
3263 |
332.75 |
|
|
|
| 4 |
A |
3162 |
3052 |
4.39 |
|
|
|
| 5 |
A |
3128 |
3018 |
22.57 |
|
|
|
| 6 |
A |
3048 |
2942 |
21.82 |
|
|
|
| 7 |
A |
3045 |
2939 |
6.68 |
|
|
|
| 8 |
A |
1880 |
1814 |
299.35 |
|
|
|
| 9 |
A |
1637 |
1580 |
31.06 |
|
|
|
| 10 |
A |
1471 |
1419 |
130.56 |
|
|
|
| 11 |
A |
1460 |
1408 |
291.26 |
|
|
|
| 12 |
A |
1458 |
1407 |
3.86 |
|
|
|
| 13 |
A |
1388 |
1339 |
14.81 |
|
|
|
| 14 |
A |
1374 |
1326 |
1.84 |
|
|
|
| 15 |
A |
1312 |
1266 |
6.57 |
|
|
|
| 16 |
A |
1246 |
1202 |
16.53 |
|
|
|
| 17 |
A |
1214 |
1171 |
8.08 |
|
|
|
| 18 |
A |
1158 |
1117 |
10.47 |
|
|
|
| 19 |
A |
1105 |
1066 |
41.26 |
|
|
|
| 20 |
A |
1011 |
976 |
1.74 |
|
|
|
| 21 |
A |
940 |
907 |
49.37 |
|
|
|
| 22 |
A |
915 |
883 |
43.80 |
|
|
|
| 23 |
A |
883 |
852 |
98.37 |
|
|
|
| 24 |
A |
811 |
783 |
7.81 |
|
|
|
| 25 |
A |
741 |
715 |
6.14 |
|
|
|
| 26 |
A |
568 |
548 |
3.77 |
|
|
|
| 27 |
A |
535 |
516 |
0.64 |
|
|
|
| 28 |
A |
408 |
394 |
5.18 |
|
|
|
| 29 |
A |
369 |
356 |
18.89 |
|
|
|
| 30 |
A |
313 |
302 |
13.85 |
|
|
|
| 31 |
A |
274 |
265 |
13.23 |
|
|
|
| 32 |
A |
225 |
217 |
0.27 |
|
|
|
| 33 |
A |
83 |
80 |
1.21 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23808.7 cm
-1
Scaled (by 0.965) Zero Point Vibrational Energy (zpe) 22975.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.215 |
|
|
|
| 2 |
C |
-0.034 |
|
|
|
| 3 |
C |
-0.153 |
|
|
|
| 4 |
C |
0.215 |
|
|
|
| 5 |
O |
-0.171 |
|
|
|
| 6 |
O |
-0.249 |
|
|
|
| 7 |
H |
0.108 |
|
|
|
| 8 |
H |
0.109 |
|
|
|
| 9 |
H |
0.072 |
|
|
|
| 10 |
H |
0.046 |
|
|
|
| 11 |
H |
0.077 |
|
|
|
| 12 |
H |
0.036 |
|
|
|
| 13 |
H |
0.158 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
4.793 |
2.185 |
0.268 |
5.275 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.363 |
-0.020 |
-0.373 |
| y |
-0.020 |
-36.861 |
-0.170 |
| z |
-0.373 |
-0.170 |
-33.234 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.315 |
-0.020 |
-0.373 |
| y |
-0.020 |
-1.063 |
-0.170 |
| z |
-0.373 |
-0.170 |
4.378 |
|
| Polar |
|---|
| 3z2-r2 | 8.756 |
|---|
| x2-y2 | -1.501 |
|---|
| xy | -0.020 |
|---|
| xz | -0.373 |
|---|
| yz | -0.170 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.066 |
0.192 |
0.085 |
| y |
0.192 |
7.466 |
-0.057 |
| z |
0.085 |
-0.057 |
5.305 |
<r2> (average value of r
2) Å
2
| <r2> |
162.223 |
| (<r2>)1/2 |
12.737 |