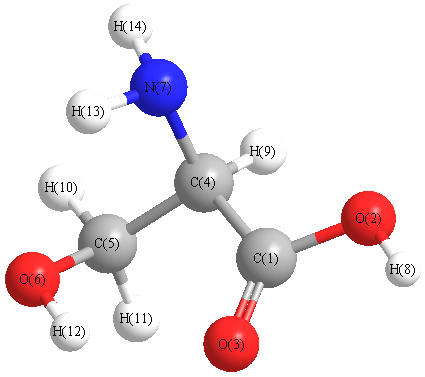
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3561 |
3540 |
34.59 |
|
|
|
| 2 |
A |
3511 |
3491 |
146.02 |
|
|
|
| 3 |
A |
3455 |
3435 |
1.15 |
|
|
|
| 4 |
A |
3376 |
3357 |
0.27 |
|
|
|
| 5 |
A |
3027 |
3009 |
25.15 |
|
|
|
| 6 |
A |
2959 |
2942 |
12.89 |
|
|
|
| 7 |
A |
2850 |
2834 |
82.96 |
|
|
|
| 8 |
A |
1733 |
1723 |
207.29 |
|
|
|
| 9 |
A |
1602 |
1593 |
25.53 |
|
|
|
| 10 |
A |
1435 |
1427 |
3.43 |
|
|
|
| 11 |
A |
1419 |
1411 |
63.29 |
|
|
|
| 12 |
A |
1336 |
1328 |
35.54 |
|
|
|
| 13 |
A |
1331 |
1323 |
16.06 |
|
|
|
| 14 |
A |
1317 |
1310 |
28.88 |
|
|
|
| 15 |
A |
1262 |
1255 |
10.47 |
|
|
|
| 16 |
A |
1189 |
1182 |
2.69 |
|
|
|
| 17 |
A |
1170 |
1164 |
13.68 |
|
|
|
| 18 |
A |
1146 |
1139 |
129.58 |
|
|
|
| 19 |
A |
1121 |
1115 |
78.63 |
|
|
|
| 20 |
A |
1079 |
1072 |
101.41 |
|
|
|
| 21 |
A |
999 |
993 |
35.33 |
|
|
|
| 22 |
A |
962 |
956 |
2.48 |
|
|
|
| 23 |
A |
876 |
871 |
149.84 |
|
|
|
| 24 |
A |
783 |
779 |
25.84 |
|
|
|
| 25 |
A |
722 |
718 |
36.74 |
|
|
|
| 26 |
A |
646 |
643 |
58.07 |
|
|
|
| 27 |
A |
602 |
599 |
144.29 |
|
|
|
| 28 |
A |
551 |
548 |
2.30 |
|
|
|
| 29 |
A |
528 |
525 |
48.19 |
|
|
|
| 30 |
A |
404 |
401 |
7.63 |
|
|
|
| 31 |
A |
311 |
309 |
2.71 |
|
|
|
| 32 |
A |
294 |
293 |
8.53 |
|
|
|
| 33 |
A |
240 |
239 |
11.01 |
|
|
|
| 34 |
A |
222 |
220 |
24.20 |
|
|
|
| 35 |
A |
164 |
164 |
1.29 |
|
|
|
| 36 |
A |
47 |
47 |
0.15 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24114.7 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 23974.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.171 |
|
|
|
| 2 |
O |
-0.139 |
|
|
|
| 3 |
O |
-0.239 |
|
|
|
| 4 |
C |
-0.118 |
|
|
|
| 5 |
C |
0.152 |
|
|
|
| 6 |
O |
-0.245 |
|
|
|
| 7 |
N |
-0.131 |
|
|
|
| 8 |
H |
0.153 |
|
|
|
| 9 |
H |
0.062 |
|
|
|
| 10 |
H |
0.002 |
|
|
|
| 11 |
H |
0.036 |
|
|
|
| 12 |
H |
0.133 |
|
|
|
| 13 |
H |
0.085 |
|
|
|
| 14 |
H |
0.079 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.486 |
0.322 |
0.600 |
3.552 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.410 |
-2.279 |
-0.184 |
| y |
-2.279 |
-44.159 |
0.454 |
| z |
-0.184 |
0.454 |
-39.178 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.259 |
-2.279 |
-0.184 |
| y |
-2.279 |
-3.865 |
0.454 |
| z |
-0.184 |
0.454 |
3.606 |
|
| Polar |
|---|
| 3z2-r2 | 7.212 |
|---|
| x2-y2 | 2.749 |
|---|
| xy | -2.279 |
|---|
| xz | -0.184 |
|---|
| yz | 0.454 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.739 |
0.080 |
0.113 |
| y |
0.080 |
8.015 |
0.068 |
| z |
0.113 |
0.068 |
5.899 |
<r2> (average value of r
2) Å
2
| <r2> |
217.260 |
| (<r2>)1/2 |
14.740 |