return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H6O (2,5-Cyclohexadienone)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-306.574739
Energy at 298.15K-306.581294
HF Energy-305.562845
Nuclear repulsion energy266.622839
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3226 3095 0.32      
2 A1 3188 3058 7.22      
3 A1 3039 2916 15.94      
4 A1 1791 1719 230.91      
5 A1 1718 1648 8.35      
6 A1 1448 1389 8.30      
7 A1 1413 1356 2.60      
8 A1 1186 1138 5.03      
9 A1 959 920 3.59      
10 A1 895 859 5.54      
11 A1 774 743 2.12      
12 A1 498 477 1.64      
13 A2 1212 1163 0.00      
14 A2 1003 962 0.00      
15 A2 753 722 0.00      
16 A2 353 339 0.00      
17 B1 3071 2946 10.59      
18 B1 1012 971 1.67      
19 B1 961 922 21.15      
20 B1 846 812 36.96      
21 B1 577 553 16.95      
22 B1 289 277 2.22      
23 B1 119 114 0.87      
24 B2 3224 3093 16.65      
25 B2 3187 3057 13.92      
26 B2 1696 1627 1.70      
27 B2 1439 1381 35.65      
28 B2 1394 1337 5.18      
29 B2 1275 1224 22.48      
30 B2 1140 1093 7.82      
31 B2 1004 963 5.31      
32 B2 571 548 0.35      
33 B2 448 430 13.86      

Unscaled Zero Point Vibrational Energy (zpe) 22853.4 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 21925.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.17201 0.08758 0.05866

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.126
C2 0.000 0.000 -1.840
C3 0.000 1.266 0.335
C4 0.000 -1.266 0.335
C5 0.000 1.264 -1.018
C6 0.000 -1.264 -1.018
O7 0.000 0.000 2.352
H8 0.000 2.197 0.913
H9 0.000 -2.197 0.913
H10 0.000 2.215 -1.567
H11 0.000 -2.215 -1.567
H12 0.880 0.000 -2.516
H13 -0.880 0.000 -2.516

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 O7 H8 H9 H10 H11 H12 H13
C12.96691.49351.49352.48892.48891.22602.20752.20753.48743.48743.74753.7475
C22.96692.51702.51701.50811.50814.19293.52283.52282.23172.23171.10941.1094
C31.49352.51702.53271.35252.86902.38221.09593.51152.12543.96703.24143.2414
C41.49352.51702.53272.86901.35252.38223.51151.09593.96702.12543.24143.2414
C52.48891.50811.35252.86902.52783.59942.14463.96321.09843.52202.14882.1488
C62.48891.50812.86901.35252.52783.59943.96322.14463.52201.09842.14882.1488
O71.22604.19292.38222.38223.59943.59942.62662.62664.50224.50224.94764.9476
H82.20753.52281.09593.51152.14463.96322.62664.39432.48055.06154.16694.1669
H92.20753.52283.51151.09593.96322.14462.62664.39435.06152.48054.16694.1669
H103.48742.23172.12543.96701.09843.52204.50222.48055.06154.42992.56532.5653
H113.48742.23173.96702.12543.52201.09844.50225.06152.48054.42992.56532.5653
H123.74751.10943.24143.24142.14882.14884.94764.16694.16692.56532.56531.7596
H133.74751.10943.24143.24142.14882.14884.94764.16694.16692.56532.56531.7596

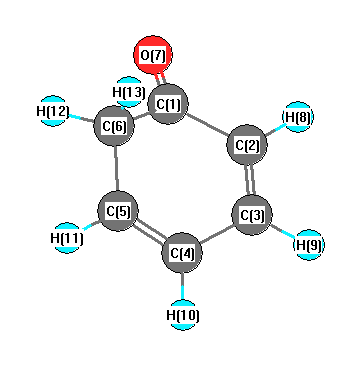
picture of 2,5-Cyclohexadienone state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C3 C5 121.907 C1 C3 H8 116.130
C1 C4 C6 121.907 C1 C4 H9 116.130
C2 C5 C3 123.168 C2 C5 H10 116.920
C2 C6 C4 123.168 C2 C6 H11 116.920
C3 C1 C4 115.976 C3 C1 O7 122.012
C3 C5 H10 119.912 C4 C1 O7 122.012
C4 C6 H11 119.912 C5 C2 C6 113.873
C5 C2 H12 109.412 C5 C2 H13 109.412
C5 C3 H8 121.963 C6 C2 H12 109.412
C6 C2 H13 109.412 C6 C4 H9 121.963
H12 C2 H13 104.939
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability