return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H6O (2,5-Cyclohexadienone)

using model chemistry: CCD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at CCD/cc-pVDZ
 hartrees
Energy at 0K-306.562680
Energy at 298.15K-306.569271
HF Energy-305.563814
Nuclear repulsion energy266.991380
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3234 3094 0.28      
2 A1 3196 3058 6.02      
3 A1 3046 2915 15.11      
4 A1 1845 1765 203.12      
5 A1 1741 1666 0.61      
6 A1 1451 1388 8.19      
7 A1 1420 1358 3.13      
8 A1 1191 1139 3.80      
9 A1 962 921 3.24      
10 A1 899 860 6.42      
11 A1 779 746 2.03      
12 A1 500 478 1.50      
13 A2 1216 1163 0.00      
14 A2 1010 966 0.00      
15 A2 755 722 0.00      
16 A2 357 341 0.00      
17 B1 3079 2946 10.33      
18 B1 1019 975 1.59      
19 B1 965 923 21.60      
20 B1 852 815 35.70      
21 B1 579 554 17.86      
22 B1 294 281 2.51      
23 B1 120 115 0.93      
24 B2 3232 3092 15.87      
25 B2 3196 3057 13.08      
26 B2 1718 1644 1.62      
27 B2 1448 1385 36.37      
28 B2 1399 1338 4.76      
29 B2 1283 1227 21.88      
30 B2 1145 1095 7.10      
31 B2 1008 965 4.88      
32 B2 576 551 0.22      
33 B2 453 433 14.33      

Unscaled Zero Point Vibrational Energy (zpe) 22982.2 cm-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 21987.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/cc-pVDZ
ABC
0.17211 0.08795 0.05884

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/cc-pVDZ

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.126
C2 0.000 0.000 -1.838
C3 0.000 1.266 0.334
C4 0.000 -1.266 0.334
C5 0.000 1.264 -1.015
C6 0.000 -1.264 -1.015
O7 0.000 0.000 2.346
H8 0.000 2.197 0.913
H9 0.000 -2.197 0.913
H10 0.000 2.214 -1.564
H11 0.000 -2.214 -1.564
H12 0.879 0.000 -2.514
H13 -0.879 0.000 -2.514

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 O7 H8 H9 H10 H11 H12 H13
C12.96401.49321.49322.48642.48641.21962.20692.20693.48463.48463.74463.7446
C22.96402.51422.51421.50761.50764.18363.51993.51992.23112.23111.10911.1091
C31.49322.51422.53201.34972.86712.37671.09563.51052.12243.96453.23853.2385
C41.49322.51422.53202.86711.34972.37673.51051.09563.96452.12243.23853.2385
C52.48641.50761.34972.86712.52713.59072.14183.96091.09793.52092.14832.1483
C62.48641.50762.86711.34972.52713.59073.96092.14183.52091.09792.14832.1483
O71.21964.18362.37672.37673.59073.59072.62272.62274.49364.49364.93854.9385
H82.20693.51991.09563.51052.14183.96092.62274.39312.47715.05884.16394.1639
H92.20693.51993.51051.09563.96092.14182.62274.39315.05882.47714.16394.1639
H103.48462.23112.12243.96451.09793.52094.49362.47715.05884.42862.56472.5647
H113.48462.23113.96452.12243.52091.09794.49365.05882.47714.42862.56472.5647
H123.74461.10913.23853.23852.14832.14834.93854.16394.16392.56472.56471.7587
H133.74461.10913.23853.23852.14832.14834.93854.16394.16392.56472.56471.7587

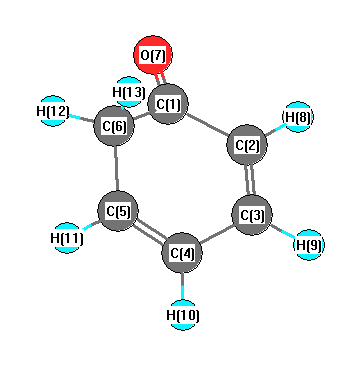
picture of 2,5-Cyclohexadienone state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C3 C5 121.919 C1 C3 H8 116.119
C1 C4 C6 121.919 C1 C4 H9 116.119
C2 C5 C3 123.167 C2 C5 H10 116.930
C2 C6 C4 123.167 C2 C6 H11 116.930
C3 C1 C4 115.953 C3 C1 O7 122.023
C3 C5 H10 119.903 C4 C1 O7 122.023
C4 C6 H11 119.903 C5 C2 C6 113.875
C5 C2 H12 109.419 C5 C2 H13 109.419
C5 C3 H8 121.962 C6 C2 H12 109.419
C6 C2 H13 109.419 C6 C4 H9 121.962
H12 C2 H13 104.903
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability