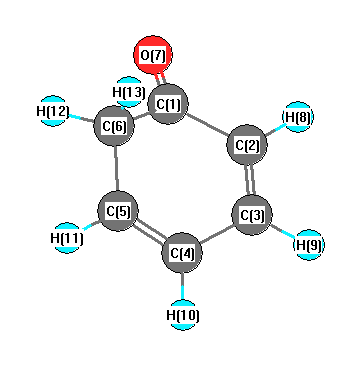
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3127 |
3109 |
0.58 |
|
|
|
| 2 |
A1 |
3089 |
3071 |
9.05 |
|
|
|
| 3 |
A1 |
2924 |
2907 |
8.86 |
|
|
|
| 4 |
A1 |
1692 |
1682 |
219.81 |
|
|
|
| 5 |
A1 |
1652 |
1643 |
12.39 |
|
|
|
| 6 |
A1 |
1357 |
1349 |
0.06 |
|
|
|
| 7 |
A1 |
1336 |
1328 |
26.55 |
|
|
|
| 8 |
A1 |
1137 |
1131 |
5.59 |
|
|
|
| 9 |
A1 |
929 |
924 |
6.80 |
|
|
|
| 10 |
A1 |
878 |
873 |
11.50 |
|
|
|
| 11 |
A1 |
761 |
757 |
0.56 |
|
|
|
| 12 |
A1 |
490 |
487 |
2.74 |
|
|
|
| 13 |
A2 |
1142 |
1136 |
0.00 |
|
|
|
| 14 |
A2 |
977 |
971 |
0.00 |
|
|
|
| 15 |
A2 |
727 |
723 |
0.00 |
|
|
|
| 16 |
A2 |
348 |
346 |
0.00 |
|
|
|
| 17 |
B1 |
2939 |
2922 |
4.19 |
|
|
|
| 18 |
B1 |
988 |
983 |
0.10 |
|
|
|
| 19 |
B1 |
903 |
898 |
22.74 |
|
|
|
| 20 |
B1 |
836 |
831 |
18.95 |
|
|
|
| 21 |
B1 |
554 |
551 |
21.91 |
|
|
|
| 22 |
B1 |
306 |
304 |
3.48 |
|
|
|
| 23 |
B1 |
127 |
127 |
0.35 |
|
|
|
| 24 |
B2 |
3126 |
3108 |
17.29 |
|
|
|
| 25 |
B2 |
3088 |
3070 |
13.89 |
|
|
|
| 26 |
B2 |
1619 |
1610 |
3.16 |
|
|
|
| 27 |
B2 |
1375 |
1367 |
33.94 |
|
|
|
| 28 |
B2 |
1324 |
1316 |
1.79 |
|
|
|
| 29 |
B2 |
1219 |
1211 |
17.46 |
|
|
|
| 30 |
B2 |
1091 |
1085 |
8.16 |
|
|
|
| 31 |
B2 |
975 |
970 |
7.61 |
|
|
|
| 32 |
B2 |
559 |
556 |
1.31 |
|
|
|
| 33 |
B2 |
436 |
434 |
11.95 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22015.9 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 21888.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.052 |
|
|
|
| 2 |
C |
0.038 |
|
|
|
| 3 |
C |
0.025 |
|
|
|
| 4 |
C |
0.025 |
|
|
|
| 5 |
C |
-0.033 |
|
|
|
| 6 |
C |
-0.033 |
|
|
|
| 7 |
O |
-0.208 |
|
|
|
| 8 |
H |
-0.007 |
|
|
|
| 9 |
H |
-0.007 |
|
|
|
| 10 |
H |
0.002 |
|
|
|
| 11 |
H |
0.002 |
|
|
|
| 12 |
H |
0.072 |
|
|
|
| 13 |
H |
0.072 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-4.276 |
4.276 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.804 |
0.000 |
0.000 |
| y |
0.000 |
-36.650 |
0.000 |
| z |
0.000 |
0.000 |
-44.017 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.471 |
0.000 |
0.000 |
| y |
0.000 |
6.261 |
0.000 |
| z |
0.000 |
0.000 |
-4.790 |
|
| Polar |
|---|
| 3z2-r2 | -9.580 |
|---|
| x2-y2 | -5.154 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.735 |
0.000 |
0.000 |
| y |
0.000 |
10.116 |
0.000 |
| z |
0.000 |
0.000 |
14.170 |
<r2> (average value of r
2) Å
2
| <r2> |
189.741 |
| (<r2>)1/2 |
13.775 |