Jump to
S1C2
Energy calculated at CCD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -550.204456 |
| Energy at 298.15K | |
| HF Energy | -548.966427 |
| Nuclear repulsion energy | 352.524045 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1g |
1864 |
1783 |
0.00 |
|
|
|
| 2 |
A1g |
705 |
674 |
0.00 |
|
|
|
| 3 |
A2g |
560 |
535 |
0.00 |
|
|
|
| 4 |
A2u |
380i |
363i |
82.81 |
|
|
|
| 5 |
B1g |
1548 |
1481 |
0.00 |
|
|
|
| 6 |
B1g |
620 |
593 |
0.00 |
|
|
|
| 7 |
B2g |
173 |
165 |
0.00 |
|
|
|
| 8 |
B2g |
3781i |
3618i |
0.00 |
|
|
|
| 9 |
B2u |
401 |
384 |
0.00 |
|
|
|
| 10 |
B2u |
205i |
196i |
0.00 |
|
|
|
| 11 |
Eg |
762i |
729i |
0.00 |
|
|
|
| 11 |
Eg |
762i |
729i |
0.00 |
|
|
|
| 12 |
Eu |
1693 |
1620 |
148.27 |
|
|
|
| 12 |
Eu |
1693 |
1620 |
148.27 |
|
|
|
| 13 |
Eu |
971 |
929 |
7.81 |
|
|
|
| 13 |
Eu |
971 |
929 |
7.81 |
|
|
|
| 14 |
Eu |
200 |
192 |
19.68 |
|
|
|
| 14 |
Eu |
200 |
192 |
19.68 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 2854.0 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 2730.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/cc-pVDZ
Point Group is D4h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
1.019 |
0.000 |
| C2 |
1.019 |
0.000 |
0.000 |
| C3 |
0.000 |
-1.019 |
0.000 |
| C4 |
-1.019 |
0.000 |
0.000 |
| F5 |
0.000 |
2.335 |
0.000 |
| F6 |
2.335 |
0.000 |
0.000 |
| F7 |
0.000 |
-2.335 |
0.000 |
| F8 |
-2.335 |
0.000 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
F5 |
F6 |
F7 |
F8 |
| C1 | | 1.4409 | 2.0377 | 1.4409 | 1.3161 | 2.5476 | 3.3538 | 2.5476 |
C2 | 1.4409 | | 1.4409 | 2.0377 | 2.5476 | 1.3161 | 2.5476 | 3.3538 | C3 | 2.0377 | 1.4409 | | 1.4409 | 3.3538 | 2.5476 | 1.3161 | 2.5476 | C4 | 1.4409 | 2.0377 | 1.4409 | | 2.5476 | 3.3538 | 2.5476 | 1.3161 | F5 | 1.3161 | 2.5476 | 3.3538 | 2.5476 | | 3.3022 | 4.6700 | 3.3022 | F6 | 2.5476 | 1.3161 | 2.5476 | 3.3538 | 3.3022 | | 3.3022 | 4.6700 | F7 | 3.3538 | 2.5476 | 1.3161 | 2.5476 | 4.6700 | 3.3022 | | 3.3022 | F8 | 2.5476 | 3.3538 | 2.5476 | 1.3161 | 3.3022 | 4.6700 | 3.3022 | |
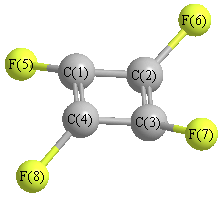 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
90.000 |
|
C1 |
C2 |
F6 |
135.000 |
| C1 |
C4 |
C3 |
90.000 |
|
C1 |
C4 |
F8 |
135.000 |
| C2 |
C1 |
C4 |
90.000 |
|
C2 |
C1 |
F5 |
135.000 |
| C2 |
C3 |
C4 |
90.000 |
|
C2 |
C3 |
F7 |
135.000 |
| C3 |
C2 |
F6 |
135.000 |
|
C3 |
C4 |
F8 |
135.000 |
| C4 |
C1 |
F5 |
135.000 |
|
C4 |
C3 |
F7 |
135.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCD/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -550.296817 |
| Energy at 298.15K | -550.296725 |
| HF Energy | -549.038866 |
| Nuclear repulsion energy | 351.041462 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
1909 |
1827 |
0.00 |
|
|
|
| 2 |
Ag |
1261 |
1207 |
0.00 |
|
|
|
| 3 |
Ag |
663 |
634 |
0.00 |
|
|
|
| 4 |
Ag |
240 |
229 |
0.00 |
|
|
|
| 5 |
Ag |
115 |
110 |
0.00 |
|
|
|
| 6 |
Au |
1377 |
1318 |
432.58 |
|
|
|
| 7 |
Au |
941 |
900 |
66.43 |
|
|
|
| 8 |
Au |
608 |
581 |
16.41 |
|
|
|
| 9 |
Au |
228 |
218 |
0.26 |
|
|
|
| 10 |
Au |
164 |
156 |
0.03 |
|
|
|
| 11 |
Bg |
1413 |
1352 |
0.00 |
|
|
|
| 12 |
Bg |
775 |
741 |
0.00 |
|
|
|
| 13 |
Bg |
498 |
476 |
0.00 |
|
|
|
| 14 |
Bg |
494 |
473 |
0.00 |
|
|
|
| 15 |
Bu |
1898 |
1816 |
113.94 |
|
|
|
| 16 |
Bu |
991 |
948 |
183.61 |
|
|
|
| 17 |
Bu |
305 |
292 |
7.21 |
|
|
|
| 18 |
Bu |
209 |
200 |
1.62 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 7043.7 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 6738.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/cc-pVDZ
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.062 |
0.787 |
0.669 |
| C2 |
-0.062 |
-0.787 |
0.669 |
| C3 |
-0.062 |
-0.787 |
-0.669 |
| C4 |
0.062 |
0.787 |
-0.669 |
| F5 |
-0.062 |
1.662 |
1.649 |
| F6 |
0.062 |
-1.662 |
1.649 |
| F7 |
0.062 |
-1.662 |
-1.649 |
| F8 |
-0.062 |
1.662 |
-1.649 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
F5 |
F6 |
F7 |
F8 |
| C1 | | 1.5787 | 2.0691 | 1.3374 | 1.3197 | 2.6375 | 3.3716 | 2.4804 |
C2 | 1.5787 | | 1.3374 | 2.0691 | 2.6375 | 1.3197 | 2.4804 | 3.3716 | C3 | 2.0691 | 1.3374 | | 1.5787 | 3.3716 | 2.4804 | 1.3197 | 2.6375 | C4 | 1.3374 | 2.0691 | 1.5787 | | 2.4804 | 3.3716 | 2.6375 | 1.3197 | F5 | 1.3197 | 2.6375 | 3.3716 | 2.4804 | | 3.3256 | 4.6837 | 3.2981 | F6 | 2.6375 | 1.3197 | 2.4804 | 3.3716 | 3.3256 | | 3.2981 | 4.6837 | F7 | 3.3716 | 2.4804 | 1.3197 | 2.6375 | 4.6837 | 3.2981 | | 3.3256 | F8 | 2.4804 | 3.3716 | 2.6375 | 1.3197 | 3.2981 | 4.6837 | 3.3256 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
90.000 |
|
C1 |
C2 |
F6 |
130.794 |
| C1 |
C4 |
C3 |
90.000 |
|
C1 |
C4 |
F8 |
137.972 |
| C2 |
C1 |
C4 |
90.000 |
|
C2 |
C1 |
F5 |
130.794 |
| C2 |
C3 |
C4 |
90.000 |
|
C2 |
C3 |
F7 |
137.972 |
| C3 |
C2 |
F6 |
137.972 |
|
C3 |
C4 |
F8 |
130.794 |
| C4 |
C1 |
F5 |
137.972 |
|
C4 |
C3 |
F7 |
130.794 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
