return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H12O (3-Pentanol)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-272.200009
Energy at 298.15K-272.213428
Nuclear repulsion energy257.603221
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3868 3711 21.04      
2 A 3162 3034 23.42      
3 A 3140 3013 30.83      
4 A 3132 3005 60.51      
5 A 3130 3003 38.65      
6 A 3105 2979 24.36      
7 A 3097 2972 0.90      
8 A 3059 2935 21.30      
9 A 3056 2931 44.72      
10 A 3052 2928 10.89      
11 A 3048 2924 33.16      
12 A 2979 2859 49.65      
13 A 1510 1449 4.95      
14 A 1509 1448 5.00      
15 A 1498 1437 6.07      
16 A 1494 1433 4.58      
17 A 1482 1422 1.75      
18 A 1473 1413 1.79      
19 A 1462 1403 5.01      
20 A 1425 1367 1.06      
21 A 1415 1357 7.27      
22 A 1410 1352 6.07      
23 A 1374 1318 0.19      
24 A 1340 1286 21.71      
25 A 1310 1257 4.82      
26 A 1288 1235 9.82      
27 A 1268 1216 22.40      
28 A 1194 1145 25.75      
29 A 1166 1119 3.23      
30 A 1116 1071 8.01      
31 A 1094 1050 19.78      
32 A 1051 1008 0.65      
33 A 1034 992 1.60      
34 A 993 952 38.60      
35 A 951 912 6.45      
36 A 868 833 2.75      
37 A 788 756 1.53      
38 A 774 743 0.13      
39 A 498 478 3.32      
40 A 481 461 7.64      
41 A 401 385 2.77      
42 A 313 301 3.24      
43 A 288 277 58.91      
44 A 242 232 41.45      
45 A 224 215 10.40      
46 A 190 182 0.58      
47 A 105 101 2.41      
48 A 96 92 0.03      

Unscaled Zero Point Vibrational Energy (zpe) 36476.3 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 34995.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.24668 0.06285 0.05470

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.008 0.020 0.217
C2 -1.281 -0.689 -0.249
C3 1.263 -0.727 -0.210
C4 -2.561 -0.009 0.255
C5 2.555 -0.024 0.230
O6 -0.049 1.336 -0.341
H7 -0.026 0.082 1.330
H8 -1.245 -1.741 0.093
H9 -1.273 -0.707 -1.356
H10 1.251 -0.831 -1.312
H11 1.233 -1.749 0.216
H12 -3.462 -0.528 -0.119
H13 -2.601 -0.015 1.361
H14 -2.594 1.040 -0.082
H15 3.448 -0.605 -0.062
H16 2.644 0.973 -0.239
H17 2.583 0.106 1.329
H18 0.630 1.856 0.107

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 O6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18
C11.53001.53552.55312.56321.43021.11482.15562.14622.15622.16073.51312.83372.79643.52282.85482.82091.9464
C21.53002.54541.53353.92252.37212.15961.10711.10772.75002.76772.19022.18732.17754.73344.26274.24923.2019
C31.53552.54543.91871.53462.44902.16522.72272.78391.10741.10764.73044.23154.24552.19252.19042.19212.6782
C42.55311.53353.91875.11552.91092.75552.18062.17714.20214.17331.10481.10671.10306.04615.31965.25613.6985
C52.56323.92251.53465.11552.99282.80704.17194.19962.17502.17326.04775.27795.26731.10431.10581.10732.6936
O61.43022.37212.44902.91092.99282.08993.32982.58942.70753.38693.89493.35142.57544.00902.71943.35110.9648
H71.11482.15962.16522.75552.80702.08992.51763.06553.07342.48493.77902.57723.08453.80433.22232.60862.2526
H82.15561.10712.72272.18064.17193.32982.51761.78103.00562.48072.53582.53403.09634.83094.75424.42654.0559
H92.14621.10772.78392.17714.19962.58943.06551.78102.52743.13642.52033.10202.53434.89624.40664.76923.5114
H102.15622.75001.10744.20212.17502.70753.07343.00562.52741.78314.87064.75854.44982.53802.52003.10363.1013
H112.16072.76771.10764.17332.17323.38692.48492.48073.13641.78314.86214.36004.74502.50873.09992.55063.6564
H123.51312.19024.73041.10486.04773.89493.77902.53582.52034.87064.86211.78731.79266.91016.28886.24824.7406
H132.83372.18734.23151.10675.27793.35142.57722.53403.10204.75854.36001.78731.78786.24135.57155.18523.9380
H142.79642.17754.24551.10305.26732.57543.08453.09632.53434.44984.74501.79261.78786.26205.24125.44703.3310
H153.52284.73342.19256.04611.10434.00903.80434.83094.89622.53802.50876.91016.24136.26201.77961.78583.7447
H162.85484.26272.19045.31961.10582.71943.22234.75424.40662.52003.09996.28885.57155.24121.77961.79252.2261
H172.82094.24922.19215.25611.10733.35112.60864.42654.76923.10362.55066.24825.18525.44701.78581.79252.8931
H181.94643.20192.67823.69852.69360.96482.25264.05593.51143.10133.65644.74063.93803.33103.74472.22612.8931

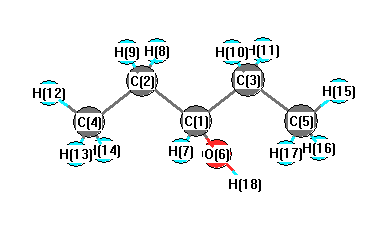
picture of 3-Pentanol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 112.896 C1 C2 H8 108.592
C1 C2 H9 107.834 C1 C3 C5 113.211
C1 C3 H10 108.249 C1 C3 H11 108.583
C1 O6 H18 107.113 C2 C1 C3 112.269
C2 C1 O6 106.469 C2 C1 H7 108.463
C2 C4 H12 111.185 C2 C4 H13 110.845
C2 C4 H14 110.292 C3 C1 O6 111.281
C3 C1 H7 108.520 C3 C5 H15 111.331
C3 C5 H16 111.074 C3 C5 H17 111.115
C4 C2 H8 110.291 C4 C2 H9 109.978
C5 C3 H10 109.771 C5 C3 H11 109.618
O6 C1 H7 109.784 H8 C2 H9 107.048
H10 C3 H11 107.221 H12 C4 H13 107.834
H12 C4 H14 108.571 H13 C4 H14 108.007
H15 C5 H16 107.262 H15 C5 H17 107.700
H16 C5 H17 108.189
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability