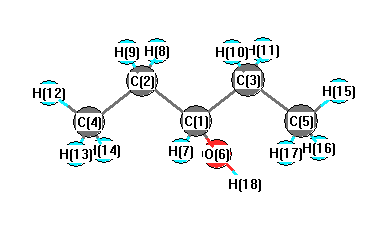
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3672 |
3651 |
12.19 |
|
|
|
| 2 |
A |
3071 |
3053 |
18.51 |
|
|
|
| 3 |
A |
3053 |
3035 |
23.73 |
|
|
|
| 4 |
A |
3047 |
3029 |
39.05 |
|
|
|
| 5 |
A |
3035 |
3017 |
47.05 |
|
|
|
| 6 |
A |
3009 |
2991 |
23.03 |
|
|
|
| 7 |
A |
3001 |
2984 |
0.87 |
|
|
|
| 8 |
A |
2971 |
2953 |
31.12 |
|
|
|
| 9 |
A |
2963 |
2946 |
50.16 |
|
|
|
| 10 |
A |
2959 |
2942 |
28.97 |
|
|
|
| 11 |
A |
2957 |
2940 |
3.29 |
|
|
|
| 12 |
A |
2850 |
2833 |
59.29 |
|
|
|
| 13 |
A |
1433 |
1425 |
5.80 |
|
|
|
| 14 |
A |
1431 |
1423 |
5.43 |
|
|
|
| 15 |
A |
1420 |
1412 |
8.05 |
|
|
|
| 16 |
A |
1415 |
1407 |
5.03 |
|
|
|
| 17 |
A |
1402 |
1394 |
2.86 |
|
|
|
| 18 |
A |
1391 |
1383 |
2.58 |
|
|
|
| 19 |
A |
1380 |
1372 |
6.78 |
|
|
|
| 20 |
A |
1345 |
1338 |
0.90 |
|
|
|
| 21 |
A |
1335 |
1327 |
3.33 |
|
|
|
| 22 |
A |
1333 |
1325 |
11.80 |
|
|
|
| 23 |
A |
1304 |
1296 |
0.18 |
|
|
|
| 24 |
A |
1276 |
1269 |
15.01 |
|
|
|
| 25 |
A |
1250 |
1243 |
4.62 |
|
|
|
| 26 |
A |
1231 |
1224 |
9.88 |
|
|
|
| 27 |
A |
1207 |
1200 |
19.52 |
|
|
|
| 28 |
A |
1130 |
1124 |
22.27 |
|
|
|
| 29 |
A |
1110 |
1104 |
4.32 |
|
|
|
| 30 |
A |
1066 |
1060 |
3.70 |
|
|
|
| 31 |
A |
1050 |
1044 |
19.06 |
|
|
|
| 32 |
A |
1004 |
998 |
1.58 |
|
|
|
| 33 |
A |
992 |
986 |
2.90 |
|
|
|
| 34 |
A |
944 |
938 |
52.55 |
|
|
|
| 35 |
A |
916 |
910 |
10.85 |
|
|
|
| 36 |
A |
836 |
831 |
4.17 |
|
|
|
| 37 |
A |
756 |
752 |
1.64 |
|
|
|
| 38 |
A |
744 |
740 |
0.31 |
|
|
|
| 39 |
A |
479 |
476 |
2.69 |
|
|
|
| 40 |
A |
465 |
462 |
6.31 |
|
|
|
| 41 |
A |
386 |
384 |
1.74 |
|
|
|
| 42 |
A |
304 |
302 |
5.33 |
|
|
|
| 43 |
A |
270 |
268 |
71.60 |
|
|
|
| 44 |
A |
233 |
232 |
25.41 |
|
|
|
| 45 |
A |
214 |
213 |
4.93 |
|
|
|
| 46 |
A |
185 |
183 |
0.22 |
|
|
|
| 47 |
A |
107 |
106 |
2.16 |
|
|
|
| 48 |
A |
91 |
91 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35009.7 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 34806.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.015 |
|
|
|
| 2 |
C |
-0.029 |
|
|
|
| 3 |
C |
-0.024 |
|
|
|
| 4 |
C |
-0.043 |
|
|
|
| 5 |
C |
-0.068 |
|
|
|
| 6 |
O |
-0.232 |
|
|
|
| 7 |
H |
-0.008 |
|
|
|
| 8 |
H |
0.014 |
|
|
|
| 9 |
H |
0.023 |
|
|
|
| 10 |
H |
0.022 |
|
|
|
| 11 |
H |
0.015 |
|
|
|
| 12 |
H |
0.023 |
|
|
|
| 13 |
H |
0.024 |
|
|
|
| 14 |
H |
0.042 |
|
|
|
| 15 |
H |
0.035 |
|
|
|
| 16 |
H |
0.023 |
|
|
|
| 17 |
H |
0.032 |
|
|
|
| 18 |
H |
0.136 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.076 |
-0.351 |
0.784 |
1.376 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.541 |
1.989 |
0.215 |
| y |
1.989 |
-39.225 |
1.229 |
| z |
0.215 |
1.229 |
-40.316 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.230 |
1.989 |
0.215 |
| y |
1.989 |
0.703 |
1.229 |
| z |
0.215 |
1.229 |
-0.933 |
|
| Polar |
|---|
| 3z2-r2 | -1.866 |
|---|
| x2-y2 | -0.315 |
|---|
| xy | 1.989 |
|---|
| xz | 0.215 |
|---|
| yz | 1.229 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.253 |
0.123 |
0.059 |
| y |
0.123 |
8.877 |
0.140 |
| z |
0.059 |
0.140 |
8.424 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |