return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CHCHCH3 (2-Butene, (Z)-)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-156.738422
Energy at 298.15K-156.746071
HF Energy-156.119343
Nuclear repulsion energy116.437242
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3183 3054 58.53      
2 A 3167 3038 2.69      
3 A 3056 2932 14.15      
4 A 1758 1686 2.96      
5 A 1494 1434 4.10      
6 A 1424 1366 1.61      
7 A 1288 1236 0.00      
8 A 1026 984 0.65      
9 A 881 845 0.44      
10 A 290 278 0.00      
11 A 3117 2990 0.00      
12 A 1480 1420 0.00      
13 A 1054 1011 0.00      
14 A 997 956 0.00      
15 A 384 368 0.00      
16 A 115 111 0.00      
17 A 3117 2991 46.76      
18 A 1486 1426 9.02      
19 A 1068 1025 3.15      
20 A 699 671 30.38      
21 A 120 115 0.66      
22 A 3163 3034 23.98      
23 A 3148 3020 0.94      
24 A 3053 2929 43.33      
25 A 1489 1428 2.35      
26 A 1450 1391 5.44      
27 A 1387 1331 5.14      
28 A 1161 1114 0.10      
29 A 996 955 9.54      
30 A 561 539 5.16      

Unscaled Zero Point Vibrational Energy (zpe) 23805.2 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 22838.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.53403 0.16688 0.13357

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.676 0.670
C2 0.000 -0.676 0.670
C3 0.000 1.598 -0.527
C4 0.000 -1.598 -0.527
H5 0.000 1.178 1.649
H6 0.000 -1.178 1.649
H7 0.000 1.054 -1.486
H8 0.000 -1.054 -1.486
H9 -0.889 2.257 -0.510
H10 0.889 2.257 -0.510
H11 0.889 -2.257 -0.510
H12 -0.889 -2.257 -0.510

Atom - Atom Distances (Å)
  C1 C2 C3 C4 H5 H6 H7 H8 H9 H10 H11 H12
C11.35101.51192.56991.10022.09622.18902.76392.16402.16403.28353.2835
C21.35102.56991.51192.09621.10022.76392.18903.28353.28352.16402.1640
C31.51192.56993.19652.21643.52771.10222.82001.10661.10663.95643.9564
C42.56991.51193.19653.52772.21642.82001.10223.95643.95641.10661.1066
H51.10022.09622.21643.52772.35653.13713.84822.57142.57144.15324.1532
H62.09621.10023.52772.21642.35653.84823.13714.15324.15322.57142.5714
H72.18902.76391.10222.82003.13713.84822.10791.78621.78623.56453.5645
H82.76392.18902.82001.10223.84823.13712.10793.56453.56451.78621.7862
H92.16403.28351.10663.95642.57144.15321.78623.56451.77804.85144.5139
H102.16403.28351.10663.95642.57144.15321.78623.56451.77804.51394.8514
H113.28352.16403.95641.10664.15322.57143.56451.78624.85144.51391.7780
H123.28352.16403.95641.10664.15322.57143.56451.78624.51394.85141.7780

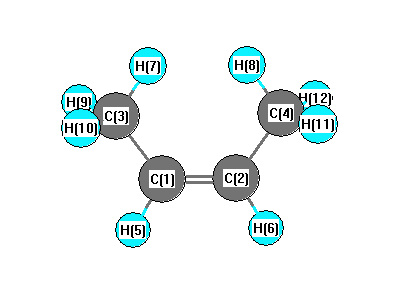
picture of 2-Butene, (Z)- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 127.613 C1 C2 H6 117.192
C1 C3 H7 112.792 C1 C3 H9 110.517
C1 C3 H10 110.517 C2 C1 C3 127.613
C2 C1 H5 117.192 C2 C4 H8 112.792
C2 C4 H11 110.517 C2 C4 H12 110.517
C3 C1 H5 115.195 C4 C2 H6 115.195
H7 C3 H9 107.942 H7 C3 H10 107.942
H8 C4 H11 107.942 H8 C4 H12 107.942
H9 C3 H10 106.905 H11 C4 H12 106.905
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability