Jump to
S1C2
Energy calculated at PBEPBE/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -616.419228 |
| Energy at 298.15K | -616.426194 |
| HF Energy | -616.419228 |
| Nuclear repulsion energy | 203.920772 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3092 |
3074 |
8.32 |
|
|
|
| 2 |
A' |
3054 |
3036 |
20.00 |
|
|
|
| 3 |
A' |
3053 |
3036 |
5.53 |
|
|
|
| 4 |
A' |
2982 |
2965 |
21.07 |
|
|
|
| 5 |
A' |
2958 |
2941 |
29.07 |
|
|
|
| 6 |
A' |
1712 |
1702 |
0.64 |
|
|
|
| 7 |
A' |
1412 |
1404 |
11.96 |
|
|
|
| 8 |
A' |
1391 |
1383 |
1.54 |
|
|
|
| 9 |
A' |
1345 |
1337 |
1.45 |
|
|
|
| 10 |
A' |
1273 |
1266 |
0.05 |
|
|
|
| 11 |
A' |
1255 |
1247 |
30.73 |
|
|
|
| 12 |
A' |
1238 |
1231 |
9.45 |
|
|
|
| 13 |
A' |
1100 |
1094 |
1.17 |
|
|
|
| 14 |
A' |
1022 |
1016 |
13.72 |
|
|
|
| 15 |
A' |
887 |
882 |
11.95 |
|
|
|
| 16 |
A' |
697 |
693 |
20.19 |
|
|
|
| 17 |
A' |
571 |
567 |
5.01 |
|
|
|
| 18 |
A' |
327 |
325 |
1.22 |
|
|
|
| 19 |
A' |
165 |
164 |
0.82 |
|
|
|
| 20 |
A" |
3028 |
3011 |
5.54 |
|
|
|
| 21 |
A" |
3019 |
3001 |
14.65 |
|
|
|
| 22 |
A" |
1399 |
1391 |
6.53 |
|
|
|
| 23 |
A" |
1138 |
1131 |
1.81 |
|
|
|
| 24 |
A" |
1014 |
1008 |
0.02 |
|
|
|
| 25 |
A" |
953 |
947 |
26.49 |
|
|
|
| 26 |
A" |
899 |
894 |
0.07 |
|
|
|
| 27 |
A" |
700 |
696 |
0.47 |
|
|
|
| 28 |
A" |
238 |
237 |
3.11 |
|
|
|
| 29 |
A" |
207 |
206 |
3.44 |
|
|
|
| 30 |
A" |
123 |
122 |
1.10 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21125.2 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 21002.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.318 |
0.309 |
0.000 |
| C2 |
0.000 |
1.025 |
0.000 |
| C3 |
1.229 |
0.481 |
0.000 |
| C4 |
2.511 |
1.259 |
0.000 |
| Cl5 |
-1.206 |
-1.499 |
0.000 |
| H6 |
-1.918 |
0.578 |
0.894 |
| H7 |
-1.918 |
0.578 |
-0.894 |
| H8 |
-0.108 |
2.126 |
0.000 |
| H9 |
1.309 |
-0.619 |
0.000 |
| H10 |
2.336 |
2.353 |
0.000 |
| H11 |
3.131 |
1.008 |
0.888 |
| H12 |
3.131 |
1.008 |
-0.888 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
Cl5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.4999 | 2.5532 | 3.9453 | 1.8114 | 1.1099 | 1.1099 | 2.1830 | 2.7864 | 4.1871 | 4.5902 | 4.5902 |
C2 | 1.4999 | | 1.3442 | 2.5222 | 2.7970 | 2.1632 | 2.1632 | 1.1067 | 2.1018 | 2.6875 | 3.2544 | 3.2544 | C3 | 2.5532 | 1.3442 | | 1.4992 | 3.1387 | 3.2737 | 3.2737 | 2.1203 | 1.1037 | 2.1746 | 2.1638 | 2.1638 | C4 | 3.9453 | 2.5222 | 1.4992 | | 4.6283 | 4.5701 | 4.5701 | 2.7596 | 2.2299 | 1.1083 | 1.1115 | 1.1115 | Cl5 | 1.8114 | 2.7970 | 3.1387 | 4.6283 | | 2.3705 | 2.3705 | 3.7876 | 2.6643 | 5.2330 | 5.0873 | 5.0873 | H6 | 1.1099 | 2.1632 | 3.2737 | 4.5701 | 2.3705 | | 1.7881 | 2.5443 | 3.5566 | 4.6961 | 5.0676 | 5.3718 | H7 | 1.1099 | 2.1632 | 3.2737 | 4.5701 | 2.3705 | 1.7881 | | 2.5443 | 3.5566 | 4.6961 | 5.3718 | 5.0676 | H8 | 2.1830 | 1.1067 | 2.1203 | 2.7596 | 3.7876 | 2.5443 | 2.5443 | | 3.0901 | 2.4552 | 3.5400 | 3.5400 | H9 | 2.7864 | 2.1018 | 1.1037 | 2.2299 | 2.6643 | 3.5566 | 3.5566 | 3.0901 | | 3.1450 | 2.5993 | 2.5993 | H10 | 4.1871 | 2.6875 | 2.1746 | 1.1083 | 5.2330 | 4.6961 | 4.6961 | 2.4552 | 3.1450 | | 1.7969 | 1.7969 | H11 | 4.5902 | 3.2544 | 2.1638 | 1.1115 | 5.0873 | 5.0676 | 5.3718 | 3.5400 | 2.5993 | 1.7969 | | 1.7763 | H12 | 4.5902 | 3.2544 | 2.1638 | 1.1115 | 5.0873 | 5.3718 | 5.0676 | 3.5400 | 2.5993 | 1.7969 | 1.7763 | |
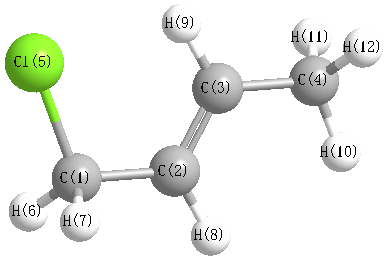 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
127.640 |
|
C1 |
C2 |
H8 |
112.888 |
| C2 |
C1 |
Cl5 |
114.956 |
|
C2 |
C1 |
H6 |
111.088 |
| C2 |
C1 |
H7 |
111.088 |
|
C2 |
C3 |
C4 |
124.914 |
| C2 |
C3 |
H9 |
117.998 |
|
C3 |
C2 |
H8 |
119.472 |
| C3 |
C4 |
H10 |
112.148 |
|
C3 |
C4 |
H11 |
111.084 |
| C3 |
C4 |
H12 |
111.084 |
|
C4 |
C3 |
H9 |
117.088 |
| Cl5 |
C1 |
H6 |
105.970 |
|
Cl5 |
C1 |
H7 |
105.970 |
| H6 |
C1 |
H7 |
107.314 |
|
H10 |
C4 |
H11 |
108.095 |
| H10 |
C4 |
H12 |
108.095 |
|
H11 |
C4 |
H12 |
106.080 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.047 |
|
|
|
| 2 |
C |
-0.049 |
|
|
|
| 3 |
C |
-0.063 |
|
|
|
| 4 |
C |
-0.014 |
|
|
|
| 5 |
Cl |
-0.129 |
|
|
|
| 6 |
H |
0.089 |
|
|
|
| 7 |
H |
0.089 |
|
|
|
| 8 |
H |
-0.006 |
|
|
|
| 9 |
H |
0.002 |
|
|
|
| 10 |
H |
0.027 |
|
|
|
| 11 |
H |
0.050 |
|
|
|
| 12 |
H |
0.050 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.310 |
1.803 |
0.000 |
1.830 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.279 |
-2.315 |
0.000 |
| y |
-2.315 |
-38.317 |
0.000 |
| z |
0.000 |
0.000 |
-39.186 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.473 |
-2.315 |
0.000 |
| y |
-2.315 |
-1.085 |
0.000 |
| z |
0.000 |
0.000 |
-2.388 |
|
| Polar |
|---|
| 3z2-r2 | -4.776 |
|---|
| x2-y2 | 3.038 |
|---|
| xy | -2.315 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.806 |
0.569 |
0.000 |
| y |
0.569 |
9.024 |
0.000 |
| z |
0.000 |
0.000 |
5.202 |
<r2> (average value of r
2) Å
2
| <r2> |
211.960 |
| (<r2>)1/2 |
14.559 |
Jump to
S1C1
Energy calculated at PBEPBE/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -616.421076 |
| Energy at 298.15K | -616.427970 |
| HF Energy | -616.421076 |
| Nuclear repulsion energy | 199.450565 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3087 |
3069 |
19.41 |
|
|
|
| 2 |
A |
3070 |
3053 |
3.60 |
|
|
|
| 3 |
A |
3068 |
3051 |
22.02 |
|
|
|
| 4 |
A |
3052 |
3034 |
1.80 |
|
|
|
| 5 |
A |
3018 |
3000 |
11.49 |
|
|
|
| 6 |
A |
3007 |
2990 |
16.15 |
|
|
|
| 7 |
A |
2958 |
2941 |
23.47 |
|
|
|
| 8 |
A |
1692 |
1682 |
16.86 |
|
|
|
| 9 |
A |
1410 |
1402 |
15.70 |
|
|
|
| 10 |
A |
1401 |
1393 |
2.36 |
|
|
|
| 11 |
A |
1397 |
1389 |
7.95 |
|
|
|
| 12 |
A |
1341 |
1334 |
1.34 |
|
|
|
| 13 |
A |
1276 |
1268 |
1.11 |
|
|
|
| 14 |
A |
1271 |
1264 |
1.56 |
|
|
|
| 15 |
A |
1200 |
1194 |
38.01 |
|
|
|
| 16 |
A |
1155 |
1148 |
3.85 |
|
|
|
| 17 |
A |
1088 |
1081 |
1.64 |
|
|
|
| 18 |
A |
1072 |
1066 |
0.40 |
|
|
|
| 19 |
A |
1011 |
1005 |
0.31 |
|
|
|
| 20 |
A |
960 |
954 |
25.52 |
|
|
|
| 21 |
A |
911 |
906 |
15.16 |
|
|
|
| 22 |
A |
856 |
851 |
3.67 |
|
|
|
| 23 |
A |
770 |
766 |
10.10 |
|
|
|
| 24 |
A |
654 |
650 |
65.92 |
|
|
|
| 25 |
A |
479 |
476 |
1.46 |
|
|
|
| 26 |
A |
341 |
339 |
6.21 |
|
|
|
| 27 |
A |
268 |
267 |
1.99 |
|
|
|
| 28 |
A |
212 |
211 |
1.38 |
|
|
|
| 29 |
A |
154 |
153 |
2.71 |
|
|
|
| 30 |
A |
87 |
87 |
0.27 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21133.9 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 21011.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.774 |
0.841 |
0.092 |
| C2 |
0.521 |
0.203 |
0.458 |
| C3 |
1.610 |
0.218 |
-0.338 |
| C4 |
2.938 |
-0.387 |
-0.001 |
| Cl5 |
-2.112 |
-0.408 |
-0.096 |
| H6 |
-1.146 |
1.528 |
0.877 |
| H7 |
-0.718 |
1.380 |
-0.872 |
| H8 |
0.563 |
-0.301 |
1.439 |
| H9 |
1.531 |
0.718 |
-1.321 |
| H10 |
2.932 |
-0.871 |
0.995 |
| H11 |
3.229 |
-1.150 |
-0.756 |
| H12 |
3.743 |
0.378 |
-0.011 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
Cl5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.4892 | 2.5010 | 3.9105 | 1.8398 | 1.1072 | 1.1063 | 2.2149 | 2.7068 | 4.1806 | 4.5498 | 4.5419 |
C2 | 1.4892 | | 1.3484 | 2.5294 | 2.7593 | 2.1703 | 2.1653 | 1.1041 | 2.1096 | 2.6931 | 3.2607 | 3.2605 | C3 | 2.5010 | 1.3484 | | 1.4976 | 3.7819 | 3.2841 | 2.6556 | 2.1266 | 1.1062 | 2.1704 | 2.1601 | 2.1642 | C4 | 3.9105 | 2.5294 | 1.4976 | | 5.0505 | 4.5951 | 4.1523 | 2.7784 | 2.2233 | 1.1077 | 1.1115 | 1.1115 | Cl5 | 1.8398 | 2.7593 | 3.7819 | 5.0505 | | 2.3716 | 2.3966 | 3.0862 | 4.0053 | 5.1810 | 5.4320 | 5.9084 | H6 | 1.1072 | 2.1703 | 3.2841 | 4.5951 | 2.3716 | | 1.8067 | 2.5654 | 3.5574 | 4.7325 | 5.3825 | 5.1005 | H7 | 1.1063 | 2.1653 | 2.6556 | 4.1523 | 2.3966 | 1.8067 | | 3.1318 | 2.3867 | 4.6767 | 4.6887 | 4.6523 | H8 | 2.2149 | 1.1041 | 2.1266 | 2.7784 | 3.0862 | 2.5654 | 3.1318 | | 3.0978 | 2.4762 | 3.5557 | 3.5603 | H9 | 2.7068 | 2.1096 | 1.1062 | 2.2233 | 4.0053 | 3.5574 | 2.3867 | 3.0978 | | 3.1391 | 2.5865 | 2.5936 | H10 | 4.1806 | 2.6931 | 2.1704 | 1.1077 | 5.1810 | 4.7325 | 4.6767 | 2.4762 | 3.1391 | | 1.7978 | 1.7978 | H11 | 4.5498 | 3.2607 | 2.1601 | 1.1115 | 5.4320 | 5.3825 | 4.6887 | 3.5557 | 2.5865 | 1.7978 | | 1.7763 | H12 | 4.5419 | 3.2605 | 2.1642 | 1.1115 | 5.9084 | 5.1005 | 4.6523 | 3.5603 | 2.5936 | 1.7978 | 1.7763 | |
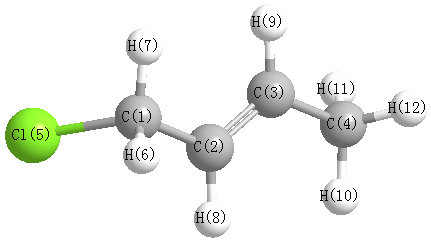 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
123.542 |
|
C1 |
C2 |
H8 |
116.532 |
| C2 |
C1 |
Cl5 |
111.535 |
|
C2 |
C1 |
H6 |
112.583 |
| C2 |
C1 |
H7 |
112.231 |
|
C2 |
C3 |
C4 |
125.356 |
| C2 |
C3 |
H9 |
118.177 |
|
C3 |
C2 |
H8 |
119.926 |
| C3 |
C4 |
H10 |
111.965 |
|
C3 |
C4 |
H11 |
110.905 |
| C3 |
C4 |
H12 |
111.233 |
|
C4 |
C3 |
H9 |
116.467 |
| Cl5 |
C1 |
H6 |
104.412 |
|
Cl5 |
C1 |
H7 |
106.190 |
| H6 |
C1 |
H7 |
109.418 |
|
H10 |
C4 |
H11 |
108.210 |
| H10 |
C4 |
H12 |
108.213 |
|
H11 |
C4 |
H12 |
106.077 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.061 |
|
|
|
| 2 |
C |
0.003 |
|
|
|
| 3 |
C |
-0.067 |
|
|
|
| 4 |
C |
-0.017 |
|
|
|
| 5 |
Cl |
-0.145 |
|
|
|
| 6 |
H |
0.087 |
|
|
|
| 7 |
H |
0.070 |
|
|
|
| 8 |
H |
-0.004 |
|
|
|
| 9 |
H |
-0.006 |
|
|
|
| 10 |
H |
0.035 |
|
|
|
| 11 |
H |
0.055 |
|
|
|
| 12 |
H |
0.052 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.221 |
1.106 |
0.102 |
2.483 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.969 |
-1.937 |
-0.970 |
| y |
-1.937 |
-37.767 |
-0.651 |
| z |
-0.970 |
-0.651 |
-37.131 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.520 |
-1.937 |
-0.970 |
| y |
-1.937 |
0.283 |
-0.651 |
| z |
-0.970 |
-0.651 |
1.237 |
|
| Polar |
|---|
| 3z2-r2 | 2.474 |
|---|
| x2-y2 | -1.202 |
|---|
| xy | -1.937 |
|---|
| xz | -0.970 |
|---|
| yz | -0.651 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.660 |
0.048 |
-0.767 |
| y |
0.048 |
6.540 |
-0.388 |
| z |
-0.767 |
-0.388 |
6.856 |
<r2> (average value of r
2) Å
2
| <r2> |
234.234 |
| (<r2>)1/2 |
15.305 |
