Jump to
S1C2
Energy calculated at PBEPBEultrafine/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -616.419211 |
| Energy at 298.15K | -616.426161 |
| HF Energy | -616.419211 |
| Nuclear repulsion energy | 203.910652 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3090 |
3072 |
8.60 |
|
|
|
| 2 |
A' |
3053 |
3036 |
21.47 |
|
|
|
| 3 |
A' |
3053 |
3035 |
3.78 |
|
|
|
| 4 |
A' |
2980 |
2963 |
21.10 |
|
|
|
| 5 |
A' |
2957 |
2940 |
29.09 |
|
|
|
| 6 |
A' |
1711 |
1701 |
0.64 |
|
|
|
| 7 |
A' |
1412 |
1403 |
11.92 |
|
|
|
| 8 |
A' |
1390 |
1382 |
1.59 |
|
|
|
| 9 |
A' |
1343 |
1336 |
1.42 |
|
|
|
| 10 |
A' |
1271 |
1264 |
0.04 |
|
|
|
| 11 |
A' |
1253 |
1246 |
31.70 |
|
|
|
| 12 |
A' |
1236 |
1229 |
8.52 |
|
|
|
| 13 |
A' |
1100 |
1094 |
1.19 |
|
|
|
| 14 |
A' |
1021 |
1015 |
13.72 |
|
|
|
| 15 |
A' |
886 |
881 |
11.96 |
|
|
|
| 16 |
A' |
696 |
692 |
20.16 |
|
|
|
| 17 |
A' |
570 |
567 |
5.02 |
|
|
|
| 18 |
A' |
326 |
324 |
1.22 |
|
|
|
| 19 |
A' |
164 |
163 |
0.82 |
|
|
|
| 20 |
A" |
3026 |
3009 |
5.51 |
|
|
|
| 21 |
A" |
3017 |
3000 |
14.68 |
|
|
|
| 22 |
A" |
1398 |
1390 |
6.54 |
|
|
|
| 23 |
A" |
1137 |
1130 |
1.84 |
|
|
|
| 24 |
A" |
1014 |
1008 |
0.03 |
|
|
|
| 25 |
A" |
953 |
948 |
26.52 |
|
|
|
| 26 |
A" |
898 |
893 |
0.07 |
|
|
|
| 27 |
A" |
699 |
695 |
0.45 |
|
|
|
| 28 |
A" |
236 |
235 |
3.04 |
|
|
|
| 29 |
A" |
205 |
204 |
3.49 |
|
|
|
| 30 |
A" |
119 |
118 |
1.12 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21108.3 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 20985.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.318 |
0.308 |
0.000 |
| C2 |
0.000 |
1.025 |
0.000 |
| C3 |
1.230 |
0.482 |
0.000 |
| C4 |
2.511 |
1.260 |
0.000 |
| Cl5 |
-1.206 |
-1.499 |
0.000 |
| H6 |
-1.918 |
0.577 |
0.894 |
| H7 |
-1.918 |
0.577 |
-0.894 |
| H8 |
-0.109 |
2.126 |
0.000 |
| H9 |
1.310 |
-0.619 |
0.000 |
| H10 |
2.336 |
2.354 |
0.000 |
| H11 |
3.131 |
1.010 |
0.888 |
| H12 |
3.131 |
1.010 |
-0.888 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
Cl5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.4999 | 2.5533 | 3.9454 | 1.8110 | 1.1099 | 1.1099 | 2.1830 | 2.7869 | 4.1873 | 4.5904 | 4.5904 |
C2 | 1.4999 | | 1.3442 | 2.5222 | 2.7974 | 2.1630 | 2.1630 | 1.1068 | 2.1020 | 2.6876 | 3.2545 | 3.2545 | C3 | 2.5533 | 1.3442 | | 1.4992 | 3.1398 | 3.2736 | 3.2736 | 2.1203 | 1.1037 | 2.1746 | 2.1638 | 2.1638 | C4 | 3.9454 | 2.5222 | 1.4992 | | 4.6294 | 4.5699 | 4.5699 | 2.7596 | 2.2298 | 1.1083 | 1.1115 | 1.1115 | Cl5 | 1.8110 | 2.7974 | 3.1398 | 4.6294 | | 2.3703 | 2.3703 | 3.7878 | 2.6660 | 5.2339 | 5.0886 | 5.0886 | H6 | 1.1099 | 2.1630 | 3.2736 | 4.5699 | 2.3703 | | 1.7880 | 2.5439 | 3.5571 | 4.6959 | 5.0675 | 5.3717 | H7 | 1.1099 | 2.1630 | 3.2736 | 4.5699 | 2.3703 | 1.7880 | | 2.5439 | 3.5571 | 4.6959 | 5.3717 | 5.0675 | H8 | 2.1830 | 1.1068 | 2.1203 | 2.7596 | 3.7878 | 2.5439 | 2.5439 | | 3.0902 | 2.4552 | 3.5401 | 3.5401 | H9 | 2.7869 | 2.1020 | 1.1037 | 2.2298 | 2.6660 | 3.5571 | 3.5571 | 3.0902 | | 3.1449 | 2.5991 | 2.5991 | H10 | 4.1873 | 2.6876 | 2.1746 | 1.1083 | 5.2339 | 4.6959 | 4.6959 | 2.4552 | 3.1449 | | 1.7969 | 1.7969 | H11 | 4.5904 | 3.2545 | 2.1638 | 1.1115 | 5.0886 | 5.0675 | 5.3717 | 3.5401 | 2.5991 | 1.7969 | | 1.7762 | H12 | 4.5904 | 3.2545 | 2.1638 | 1.1115 | 5.0886 | 5.3717 | 5.0675 | 3.5401 | 2.5991 | 1.7969 | 1.7762 | |
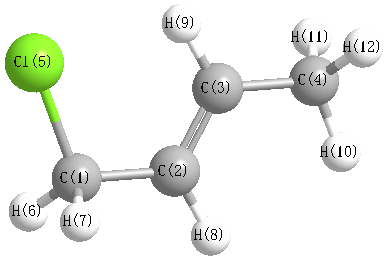 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
127.648 |
|
C1 |
C2 |
H8 |
112.885 |
| C2 |
C1 |
Cl5 |
114.998 |
|
C2 |
C1 |
H6 |
111.065 |
| C2 |
C1 |
H7 |
111.065 |
|
C2 |
C3 |
C4 |
124.916 |
| C2 |
C3 |
H9 |
118.011 |
|
C3 |
C2 |
H8 |
119.468 |
| C3 |
C4 |
H10 |
112.152 |
|
C3 |
C4 |
H11 |
111.086 |
| C3 |
C4 |
H12 |
111.086 |
|
C4 |
C3 |
H9 |
117.073 |
| Cl5 |
C1 |
H6 |
105.974 |
|
Cl5 |
C1 |
H7 |
105.974 |
| H6 |
C1 |
H7 |
107.307 |
|
H10 |
C4 |
H11 |
108.096 |
| H10 |
C4 |
H12 |
108.096 |
|
H11 |
C4 |
H12 |
106.070 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.047 |
|
|
|
| 2 |
C |
-0.049 |
|
|
|
| 3 |
C |
-0.063 |
|
|
|
| 4 |
C |
-0.014 |
|
|
|
| 5 |
Cl |
-0.129 |
|
|
|
| 6 |
H |
0.089 |
|
|
|
| 7 |
H |
0.089 |
|
|
|
| 8 |
H |
-0.006 |
|
|
|
| 9 |
H |
0.002 |
|
|
|
| 10 |
H |
0.027 |
|
|
|
| 11 |
H |
0.050 |
|
|
|
| 12 |
H |
0.050 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.310 |
1.802 |
0.000 |
1.829 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.279 |
-2.312 |
0.000 |
| y |
-2.312 |
-38.318 |
0.000 |
| z |
0.000 |
0.000 |
-39.187 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.473 |
-2.312 |
0.000 |
| y |
-2.312 |
-1.085 |
0.000 |
| z |
0.000 |
0.000 |
-2.388 |
|
| Polar |
|---|
| 3z2-r2 | -4.776 |
|---|
| x2-y2 | 3.038 |
|---|
| xy | -2.312 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.805 |
0.571 |
0.000 |
| y |
0.571 |
9.024 |
0.000 |
| z |
0.000 |
0.000 |
5.201 |
<r2> (average value of r
2) Å
2
| <r2> |
212.017 |
| (<r2>)1/2 |
14.561 |
Jump to
S1C1
Energy calculated at PBEPBEultrafine/cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -616.421082 |
| Energy at 298.15K | -616.427979 |
| HF Energy | -616.421082 |
| Nuclear repulsion energy | 199.430140 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3088 |
3070 |
19.25 |
|
|
|
| 2 |
A |
3071 |
3053 |
3.59 |
|
|
|
| 3 |
A |
3069 |
3051 |
21.90 |
|
|
|
| 4 |
A |
3052 |
3034 |
1.97 |
|
|
|
| 5 |
A |
3019 |
3001 |
11.51 |
|
|
|
| 6 |
A |
3007 |
2990 |
16.14 |
|
|
|
| 7 |
A |
2959 |
2942 |
23.49 |
|
|
|
| 8 |
A |
1691 |
1681 |
16.82 |
|
|
|
| 9 |
A |
1410 |
1402 |
15.58 |
|
|
|
| 10 |
A |
1402 |
1394 |
2.56 |
|
|
|
| 11 |
A |
1397 |
1389 |
7.91 |
|
|
|
| 12 |
A |
1342 |
1334 |
1.34 |
|
|
|
| 13 |
A |
1276 |
1269 |
1.14 |
|
|
|
| 14 |
A |
1270 |
1263 |
1.57 |
|
|
|
| 15 |
A |
1200 |
1193 |
37.98 |
|
|
|
| 16 |
A |
1155 |
1148 |
3.80 |
|
|
|
| 17 |
A |
1088 |
1081 |
1.63 |
|
|
|
| 18 |
A |
1072 |
1066 |
0.42 |
|
|
|
| 19 |
A |
1011 |
1005 |
0.32 |
|
|
|
| 20 |
A |
961 |
955 |
25.50 |
|
|
|
| 21 |
A |
912 |
907 |
15.19 |
|
|
|
| 22 |
A |
857 |
852 |
3.51 |
|
|
|
| 23 |
A |
770 |
766 |
9.87 |
|
|
|
| 24 |
A |
653 |
649 |
66.39 |
|
|
|
| 25 |
A |
480 |
477 |
1.47 |
|
|
|
| 26 |
A |
342 |
340 |
6.24 |
|
|
|
| 27 |
A |
269 |
268 |
2.01 |
|
|
|
| 28 |
A |
212 |
210 |
1.47 |
|
|
|
| 29 |
A |
155 |
154 |
2.61 |
|
|
|
| 30 |
A |
88 |
87 |
0.30 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21137.1 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 21014.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.773 |
0.840 |
0.095 |
| C2 |
0.521 |
0.199 |
0.458 |
| C3 |
1.611 |
0.220 |
-0.337 |
| C4 |
2.938 |
-0.387 |
-0.002 |
| Cl5 |
-2.113 |
-0.407 |
-0.097 |
| H6 |
-1.144 |
1.524 |
0.883 |
| H7 |
-0.717 |
1.383 |
-0.867 |
| H8 |
0.563 |
-0.310 |
1.436 |
| H9 |
1.533 |
0.726 |
-1.317 |
| H10 |
2.931 |
-0.876 |
0.992 |
| H11 |
3.230 |
-1.145 |
-0.761 |
| H12 |
3.744 |
0.379 |
-0.006 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
Cl5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.4892 | 2.5009 | 3.9103 | 1.8401 | 1.1072 | 1.1063 | 2.2150 | 2.7068 | 4.1798 | 4.5499 | 4.5416 |
C2 | 1.4892 | | 1.3484 | 2.5292 | 2.7597 | 2.1702 | 2.1654 | 1.1040 | 2.1097 | 2.6923 | 3.2604 | 3.2605 | C3 | 2.5009 | 1.3484 | | 1.4976 | 3.7841 | 3.2830 | 2.6555 | 2.1265 | 1.1062 | 2.1702 | 2.1602 | 2.1643 | C4 | 3.9103 | 2.5292 | 1.4976 | | 5.0525 | 4.5938 | 4.1523 | 2.7778 | 2.2235 | 1.1077 | 1.1115 | 1.1115 | Cl5 | 1.8401 | 2.7597 | 3.7841 | 5.0525 | | 2.3715 | 2.3967 | 3.0858 | 4.0084 | 5.1817 | 5.4352 | 5.9102 | H6 | 1.1072 | 2.1702 | 3.2830 | 4.5938 | 2.3715 | | 1.8069 | 2.5660 | 3.5560 | 4.7309 | 5.3817 | 5.0986 | H7 | 1.1063 | 2.1654 | 2.6555 | 4.1523 | 2.3967 | 1.8069 | | 3.1319 | 2.3867 | 4.6762 | 4.6887 | 4.6527 | H8 | 2.2150 | 1.1040 | 2.1265 | 2.7778 | 3.0858 | 2.5660 | 3.1319 | | 3.0977 | 2.4750 | 3.5549 | 3.5597 | H9 | 2.7068 | 2.1097 | 1.1062 | 2.2235 | 4.0084 | 3.5560 | 2.3867 | 3.0977 | | 3.1391 | 2.5871 | 2.5938 | H10 | 4.1798 | 2.6923 | 2.1702 | 1.1077 | 5.1817 | 4.7309 | 4.6762 | 2.4750 | 3.1391 | | 1.7978 | 1.7978 | H11 | 4.5499 | 3.2604 | 2.1602 | 1.1115 | 5.4352 | 5.3817 | 4.6887 | 3.5549 | 2.5871 | 1.7978 | | 1.7764 | H12 | 4.5416 | 3.2605 | 2.1643 | 1.1115 | 5.9102 | 5.0986 | 4.6527 | 3.5597 | 2.5938 | 1.7978 | 1.7764 | |
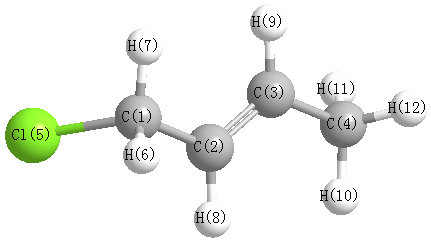 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
123.533 |
|
C1 |
C2 |
H8 |
116.549 |
| C2 |
C1 |
Cl5 |
111.542 |
|
C2 |
C1 |
H6 |
112.583 |
| C2 |
C1 |
H7 |
112.244 |
|
C2 |
C3 |
C4 |
125.329 |
| C2 |
C3 |
H9 |
118.187 |
|
C3 |
C2 |
H8 |
119.918 |
| C3 |
C4 |
H10 |
111.946 |
|
C3 |
C4 |
H11 |
110.912 |
| C3 |
C4 |
H12 |
111.234 |
|
C4 |
C3 |
H9 |
116.484 |
| Cl5 |
C1 |
H6 |
104.383 |
|
Cl5 |
C1 |
H7 |
106.178 |
| H6 |
C1 |
H7 |
109.436 |
|
H10 |
C4 |
H11 |
108.211 |
| H10 |
C4 |
H12 |
108.213 |
|
H11 |
C4 |
H12 |
106.088 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.061 |
|
|
|
| 2 |
C |
0.003 |
|
|
|
| 3 |
C |
-0.068 |
|
|
|
| 4 |
C |
-0.017 |
|
|
|
| 5 |
Cl |
-0.145 |
|
|
|
| 6 |
H |
0.087 |
|
|
|
| 7 |
H |
0.070 |
|
|
|
| 8 |
H |
-0.004 |
|
|
|
| 9 |
H |
-0.006 |
|
|
|
| 10 |
H |
0.035 |
|
|
|
| 11 |
H |
0.055 |
|
|
|
| 12 |
H |
0.052 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.223 |
1.106 |
0.107 |
2.486 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.975 |
-1.928 |
-0.975 |
| y |
-1.928 |
-37.759 |
-0.657 |
| z |
-0.975 |
-0.657 |
-37.139 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.526 |
-1.928 |
-0.975 |
| y |
-1.928 |
0.298 |
-0.657 |
| z |
-0.975 |
-0.657 |
1.228 |
|
| Polar |
|---|
| 3z2-r2 | 2.456 |
|---|
| x2-y2 | -1.216 |
|---|
| xy | -1.928 |
|---|
| xz | -0.975 |
|---|
| yz | -0.657 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.667 |
0.055 |
-0.767 |
| y |
0.055 |
6.543 |
-0.393 |
| z |
-0.767 |
-0.393 |
6.848 |
<r2> (average value of r
2) Å
2
| <r2> |
234.342 |
| (<r2>)1/2 |
15.308 |
