return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H12 (hex-1-ene)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-235.124585
Energy at 298.15K-235.136998
HF Energy-234.192194
Nuclear repulsion energy226.491385
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3256 3124 17.79      
2 A 3167 3038 5.45      
3 A 3154 3025 22.91      
4 A 3136 3008 38.03      
5 A 3131 3004 55.17      
6 A 3101 2975 54.25      
7 A 3085 2959 1.91      
8 A 3071 2946 1.71      
9 A 3051 2927 22.63      
10 A 3044 2921 84.30      
11 A 3038 2914 11.09      
12 A 3037 2913 2.14      
13 A 1725 1655 6.96      
14 A 1508 1447 3.76      
15 A 1497 1437 1.27      
16 A 1497 1436 4.98      
17 A 1487 1427 0.53      
18 A 1482 1422 1.96      
19 A 1458 1399 0.80      
20 A 1427 1369 1.08      
21 A 1416 1359 0.58      
22 A 1362 1307 0.59      
23 A 1333 1279 0.53      
24 A 1319 1265 0.07      
25 A 1314 1261 0.56      
26 A 1268 1216 6.96      
27 A 1252 1201 0.08      
28 A 1203 1154 0.26      
29 A 1137 1090 1.61      
30 A 1098 1053 0.20      
31 A 1060 1017 1.54      
32 A 1046 1003 2.90      
33 A 1018 977 11.93      
34 A 950 911 0.31      
35 A 929 891 25.79      
36 A 926 889 5.23      
37 A 915 878 0.91      
38 A 798 766 0.57      
39 A 740 710 1.70      
40 A 645 618 8.11      
41 A 456 438 0.87      
42 A 361 347 0.07      
43 A 356 342 0.41      
44 A 255 245 0.00      
45 A 174 167 0.02      
46 A 130 124 0.04      
47 A 94 90 0.01      
48 A 75 72 0.01      

Unscaled Zero Point Vibrational Energy (zpe) 36489.7 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 35008.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.43936 0.04122 0.04049

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 3.087 -0.187 -0.451
C2 2.045 -0.199 0.403
C3 0.773 0.598 0.222
C4 -0.474 -0.296 0.102
C5 -1.777 0.504 -0.042
C6 -3.015 -0.394 -0.167
H7 3.077 0.436 -1.356
H8 3.982 -0.795 -0.273
H9 2.095 -0.839 1.297
H10 0.641 1.279 1.088
H11 0.862 1.238 -0.677
H12 -0.354 -0.970 -0.770
H13 -0.543 -0.951 0.994
H14 -1.891 1.175 0.832
H15 -1.704 1.162 -0.931
H16 -3.939 0.203 -0.270
H17 -2.937 -1.054 -1.052
H18 -3.126 -1.040 0.724

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18
C11.34772.53523.60624.93016.11241.09821.09652.11403.24112.65243.54363.98225.31845.00047.03946.11586.3810
C21.34771.51192.53923.91135.09562.13552.13651.10142.15082.15192.77932.75984.19074.20516.03495.25975.2490
C32.53521.51191.53952.56533.93502.79753.53352.22911.10941.10742.17122.17472.79282.78964.75394.25624.2590
C43.60622.53921.53951.53502.55653.90824.50012.88522.16762.17861.10871.10962.16872.16853.51982.82282.8236
C54.93013.91132.56531.53501.53445.02915.90824.31122.77952.81192.17412.17081.10861.10852.19462.18862.1889
C66.11245.09563.93502.55651.53446.26227.00935.33374.21214.23742.78852.78702.17402.17361.10501.10621.1063
H71.09822.13552.79753.90825.02916.26221.87273.10333.55212.45183.75394.53395.47854.85427.10336.20326.7071
H81.09652.13653.53354.50015.90827.00931.87272.45564.16163.74614.36794.70236.29246.04907.98366.96747.1820
H92.11401.10142.22912.88524.31125.33373.10332.45562.57803.11993.20712.65774.48964.83706.32025.55685.2559
H103.24112.15081.10942.16762.77954.21213.55214.16162.57801.77913.08262.52732.54673.09664.89684.77744.4390
H112.65242.15191.10742.17862.81194.23742.45183.74613.11991.77912.52273.09243.14002.57944.92814.45264.8021
H123.54362.77932.17121.10872.17412.78853.75394.36793.20713.08262.52271.77493.08742.52833.80502.59963.1501
H133.98222.75982.17471.10962.17082.78704.53394.70232.65772.52733.09241.77492.52243.08523.80263.15052.5983
H145.31844.19072.79282.16871.10862.17405.47856.29244.48962.54673.14003.08742.52241.77362.52113.10052.5389
H155.00044.20512.78962.16851.10852.17364.85426.04904.83703.09662.57942.52833.08521.77362.52042.53853.1004
H167.03946.03494.75393.51982.19461.10507.10337.98366.32024.89684.92813.80503.80262.52112.52041.78731.7873
H176.11585.25974.25622.82282.18861.10626.20326.96745.55684.77744.45262.59963.15053.10052.53851.78731.7860
H186.38105.24904.25902.82362.18891.10636.70717.18205.25594.43904.80213.15012.59832.53893.10041.78731.7860

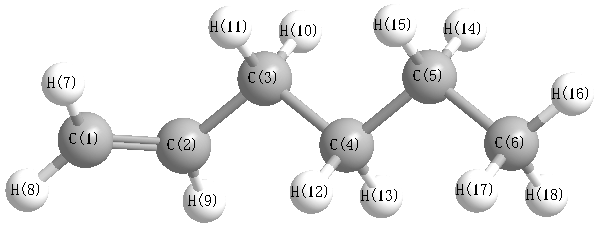
picture of hex-1-ene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 124.787 C1 C2 H9 119.017
C2 C1 H7 121.308 C2 C1 H8 121.541
C2 C3 C4 112.637 C2 C3 H10 109.311
C2 C3 H11 109.508 C3 C2 H9 116.190
C3 C4 C5 113.105 C3 C4 H12 109.064
C3 C4 H13 109.287 C4 C3 H10 108.748
C4 C3 H11 109.710 C4 C5 C6 112.794
C4 C5 H14 109.184 C4 C5 H15 109.167
C5 C4 H12 109.589 C5 C4 H13 109.284
C5 C6 H16 111.466 C5 C6 H17 110.919
C5 C6 H18 110.932 C6 C5 H14 109.630
C6 C5 H15 109.600 H7 C1 H8 117.150
H10 C3 H11 106.749 H12 C4 H13 106.284
H14 C5 H15 106.252 H16 C6 H17 107.861
H16 C6 H18 107.851 H17 C6 H18 107.651
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability