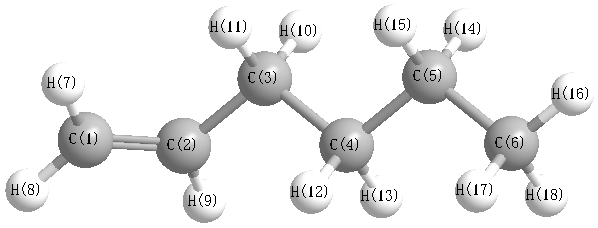
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3162 |
3144 |
16.56 |
|
|
|
| 2 |
A |
3068 |
3050 |
4.10 |
|
|
|
| 3 |
A |
3054 |
3036 |
27.79 |
|
|
|
| 4 |
A |
3050 |
3032 |
31.84 |
|
|
|
| 5 |
A |
3042 |
3024 |
43.13 |
|
|
|
| 6 |
A |
3006 |
2988 |
51.42 |
|
|
|
| 7 |
A |
2989 |
2971 |
2.51 |
|
|
|
| 8 |
A |
2975 |
2957 |
5.48 |
|
|
|
| 9 |
A |
2964 |
2947 |
35.10 |
|
|
|
| 10 |
A |
2952 |
2935 |
63.12 |
|
|
|
| 11 |
A |
2944 |
2927 |
2.13 |
|
|
|
| 12 |
A |
2932 |
2915 |
15.68 |
|
|
|
| 13 |
A |
1671 |
1661 |
13.85 |
|
|
|
| 14 |
A |
1433 |
1424 |
4.58 |
|
|
|
| 15 |
A |
1420 |
1412 |
2.84 |
|
|
|
| 16 |
A |
1420 |
1411 |
4.31 |
|
|
|
| 17 |
A |
1409 |
1400 |
0.52 |
|
|
|
| 18 |
A |
1400 |
1392 |
2.36 |
|
|
|
| 19 |
A |
1379 |
1371 |
0.69 |
|
|
|
| 20 |
A |
1346 |
1338 |
1.32 |
|
|
|
| 21 |
A |
1337 |
1329 |
0.49 |
|
|
|
| 22 |
A |
1293 |
1286 |
1.47 |
|
|
|
| 23 |
A |
1277 |
1269 |
0.84 |
|
|
|
| 24 |
A |
1266 |
1259 |
0.26 |
|
|
|
| 25 |
A |
1255 |
1248 |
0.45 |
|
|
|
| 26 |
A |
1209 |
1202 |
5.87 |
|
|
|
| 27 |
A |
1193 |
1186 |
0.13 |
|
|
|
| 28 |
A |
1150 |
1143 |
0.55 |
|
|
|
| 29 |
A |
1087 |
1081 |
2.14 |
|
|
|
| 30 |
A |
1056 |
1050 |
0.17 |
|
|
|
| 31 |
A |
1022 |
1016 |
3.66 |
|
|
|
| 32 |
A |
1005 |
1000 |
4.56 |
|
|
|
| 33 |
A |
990 |
985 |
8.33 |
|
|
|
| 34 |
A |
916 |
910 |
0.77 |
|
|
|
| 35 |
A |
892 |
887 |
14.85 |
|
|
|
| 36 |
A |
889 |
883 |
16.95 |
|
|
|
| 37 |
A |
881 |
876 |
1.43 |
|
|
|
| 38 |
A |
771 |
766 |
1.03 |
|
|
|
| 39 |
A |
721 |
717 |
3.90 |
|
|
|
| 40 |
A |
630 |
627 |
8.74 |
|
|
|
| 41 |
A |
444 |
441 |
1.19 |
|
|
|
| 42 |
A |
350 |
348 |
0.04 |
|
|
|
| 43 |
A |
342 |
340 |
0.42 |
|
|
|
| 44 |
A |
246 |
245 |
0.00 |
|
|
|
| 45 |
A |
170 |
169 |
0.02 |
|
|
|
| 46 |
A |
123 |
122 |
0.06 |
|
|
|
| 47 |
A |
90 |
89 |
0.02 |
|
|
|
| 48 |
A |
71 |
71 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35144.8 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 34941.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.006 |
|
|
|
| 2 |
C |
-0.088 |
|
|
|
| 3 |
C |
0.016 |
|
|
|
| 4 |
C |
-0.050 |
|
|
|
| 5 |
C |
-0.072 |
|
|
|
| 6 |
C |
-0.047 |
|
|
|
| 7 |
H |
0.015 |
|
|
|
| 8 |
H |
0.023 |
|
|
|
| 9 |
H |
-0.008 |
|
|
|
| 10 |
H |
0.031 |
|
|
|
| 11 |
H |
0.012 |
|
|
|
| 12 |
H |
0.024 |
|
|
|
| 13 |
H |
0.018 |
|
|
|
| 14 |
H |
0.020 |
|
|
|
| 15 |
H |
0.022 |
|
|
|
| 16 |
H |
0.027 |
|
|
|
| 17 |
H |
0.032 |
|
|
|
| 18 |
H |
0.031 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.439 |
0.081 |
0.121 |
0.463 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.136 |
-0.781 |
-0.373 |
| y |
-0.781 |
-40.670 |
-1.003 |
| z |
-0.373 |
-1.003 |
-39.521 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.041 |
-0.781 |
-0.373 |
| y |
-0.781 |
-0.341 |
-1.003 |
| z |
-0.373 |
-1.003 |
1.382 |
|
| Polar |
|---|
| 3z2-r2 | 2.764 |
|---|
| x2-y2 | -0.466 |
|---|
| xy | -0.781 |
|---|
| xz | -0.373 |
|---|
| yz | -1.003 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.814 |
-0.449 |
-1.087 |
| y |
-0.449 |
8.767 |
-0.559 |
| z |
-1.087 |
-0.559 |
9.186 |
<r2> (average value of r
2) Å
2
| <r2> |
288.774 |
| (<r2>)1/2 |
16.993 |