return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H10 (1,5-Hexadiene)

using model chemistry: CCD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1Ag
Energy calculated at CCD/cc-pVDZ
 hartrees
Energy at 0K-233.892012
Energy at 298.15K-233.901757
HF Energy-233.001500
Nuclear repulsion energy210.807360
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3267 3125 6.96      
2 A 3177 3039 0.70      
3 A 3163 3026 33.42      
4 A 3120 2985 28.10      
5 A 3051 2919 6.02      
6 A 1738 1663 2.77      
7 A 1484 1420 3.56      
8 A 1461 1397 1.41      
9 A 1388 1328 0.06      
10 A 1322 1265 1.27      
11 A 1249 1195 0.02      
12 A 1094 1047 0.10      
13 A 1073 1027 2.19      
14 A 1023 978 7.31      
15 A 958 917 0.00      
16 A 941 900 13.14      
17 A 785 751 0.21      
18 A 634 606 5.50      
19 A 417 399 0.03      
20 A 354 338 0.05      
21 A 97 92 0.02      
22 A 75 72 0.01      
23 B 3267 3125 25.38      
24 B 3176 3039 9.80      
25 B 3163 3026 9.22      
26 B 3102 2968 3.01      
27 B 3054 2922 45.31      
28 B 1741 1666 7.28      
29 B 1495 1430 5.50      
30 B 1460 1397 0.11      
31 B 1331 1273 0.39      
32 B 1314 1257 0.03      
33 B 1271 1216 6.77      
34 B 1188 1137 0.96      
35 B 1025 981 22.82      
36 B 969 927 2.13      
37 B 946 905 0.87      
38 B 941 900 47.54      
39 B 657 629 15.00      
40 B 434 416 1.43      
41 B 216 207 0.16      
42 B 110 105 0.11      

Unscaled Zero Point Vibrational Energy (zpe) 31364.4 cm-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 30006.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/cc-pVDZ
ABC
0.41307 0.04620 0.04482

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/cc-pVDZ

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.425 0.645 -0.309
C2 0.425 -0.645 -0.309
C3 0.425 1.894 -0.345
C4 -0.425 -1.894 -0.345
C5 0.443 2.835 0.617
C6 -0.443 -2.835 0.617
H7 -1.095 0.630 -1.192
H8 -1.074 0.658 0.587
H9 1.095 -0.630 -1.192
H10 1.074 -0.658 0.587
H11 1.077 2.014 -1.224
H12 -1.077 -2.014 -1.224
H13 1.087 3.719 0.542
H14 -0.190 2.751 1.509
H15 -1.087 -3.719 0.542
H16 0.190 -2.751 1.509

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
C11.54421.51142.53952.53103.60061.10881.10612.17122.17822.22892.88673.52942.79214.49463.9007
C21.54422.53951.51143.60062.53102.17122.17821.10881.10612.88672.22894.49463.90073.52942.7921
C31.51142.53953.88291.34514.90342.15112.15482.74552.79341.10104.27852.13362.13295.88035.0073
C42.53951.51143.88294.90341.34512.74552.79342.15112.15484.27851.10105.88035.00732.13362.1329
C52.53103.60061.34514.90345.73853.24042.65383.96203.54922.11225.40481.09601.09776.73015.6624
C63.60062.53104.90341.34515.73853.96203.54923.24042.65385.40482.11226.73015.66241.09601.0977
H71.10882.17122.15112.74553.24043.96201.77852.52693.08642.57652.64414.16053.55154.68094.5140
H81.10612.17822.15482.79342.65383.54921.77853.08642.51903.12163.22743.74722.45244.37663.7511
H92.17121.10882.74552.15113.96203.24042.52693.08641.77852.64412.57654.68094.51404.16053.5515
H102.17821.10612.79342.15483.54922.65383.08642.51901.77853.22743.12164.37663.75113.74722.4524
H112.22892.88671.10104.27852.11225.40482.57653.12162.64413.22744.56842.45373.10116.37705.5644
H122.88672.22894.27851.10105.40482.11222.64413.22742.57653.12164.56846.37705.56442.45373.1011
H133.52944.49462.13365.88031.09606.73014.16053.74724.68094.37662.45376.37701.87187.74846.6028
H142.79213.90072.13295.00731.09775.66243.55152.45244.51403.75113.10115.56441.87186.60285.5154
H154.49463.52945.88032.13366.73011.09604.68094.37664.16053.74726.37702.45377.74846.60281.8718
H163.90072.79215.00732.13295.66241.09774.51403.75113.55152.45245.56443.10116.60285.51541.8718

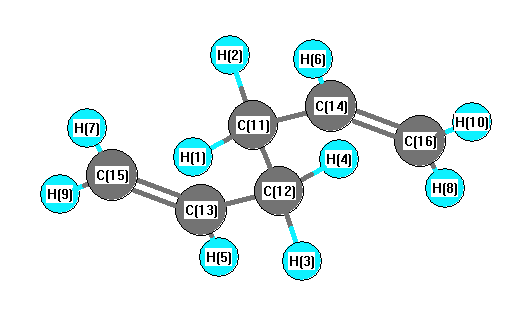
picture of 1,5-Hexadiene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 112.419 C1 C2 H9 108.739
C1 C2 H10 109.434 C1 C3 C5 124.660
C1 C3 H11 116.239 C2 C1 C3 112.419
C2 C1 H7 108.739 C2 C1 H8 109.434
C2 C4 C6 124.660 C2 C4 H12 116.239
C3 C1 H7 109.405 C3 C1 H8 109.853
C3 C5 H13 121.531 C3 C5 H14 121.327
C4 C2 H9 109.405 C4 C2 H10 109.853
C4 C6 H15 121.531 C4 C6 H16 121.327
C5 C3 H11 119.094 C6 C4 H12 119.094
H7 C1 H8 106.827 H9 C2 H10 106.827
H13 C5 H14 117.141 H15 C6 H16 117.141
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability