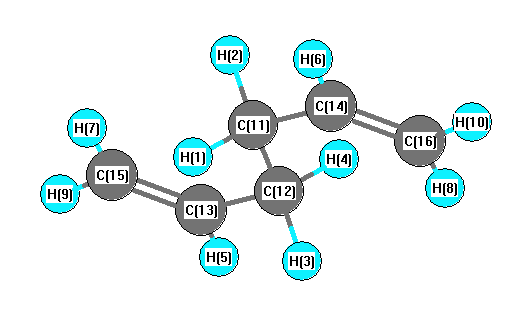
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3165 |
3147 |
6.94 |
|
|
|
| 2 |
A |
3070 |
3052 |
0.55 |
|
|
|
| 3 |
A |
3055 |
3037 |
39.06 |
|
|
|
| 4 |
A |
3017 |
2999 |
19.10 |
|
|
|
| 5 |
A |
2940 |
2923 |
7.69 |
|
|
|
| 6 |
A |
1669 |
1659 |
5.07 |
|
|
|
| 7 |
A |
1401 |
1393 |
3.15 |
|
|
|
| 8 |
A |
1376 |
1368 |
1.44 |
|
|
|
| 9 |
A |
1303 |
1296 |
0.05 |
|
|
|
| 10 |
A |
1267 |
1260 |
1.77 |
|
|
|
| 11 |
A |
1193 |
1186 |
0.02 |
|
|
|
| 12 |
A |
1053 |
1047 |
0.07 |
|
|
|
| 13 |
A |
1019 |
1013 |
1.77 |
|
|
|
| 14 |
A |
990 |
984 |
7.16 |
|
|
|
| 15 |
A |
918 |
913 |
0.01 |
|
|
|
| 16 |
A |
893 |
888 |
13.94 |
|
|
|
| 17 |
A |
765 |
760 |
0.54 |
|
|
|
| 18 |
A |
618 |
615 |
6.73 |
|
|
|
| 19 |
A |
402 |
399 |
0.00 |
|
|
|
| 20 |
A |
340 |
338 |
0.06 |
|
|
|
| 21 |
A |
97 |
97 |
0.05 |
|
|
|
| 22 |
A |
73 |
73 |
0.03 |
|
|
|
| 23 |
B |
3165 |
3147 |
23.65 |
|
|
|
| 24 |
B |
3070 |
3052 |
7.20 |
|
|
|
| 25 |
B |
3054 |
3037 |
14.67 |
|
|
|
| 26 |
B |
3000 |
2982 |
6.49 |
|
|
|
| 27 |
B |
2943 |
2926 |
37.34 |
|
|
|
| 28 |
B |
1673 |
1663 |
20.39 |
|
|
|
| 29 |
B |
1414 |
1406 |
6.14 |
|
|
|
| 30 |
B |
1380 |
1372 |
0.11 |
|
|
|
| 31 |
B |
1274 |
1266 |
0.91 |
|
|
|
| 32 |
B |
1256 |
1249 |
0.53 |
|
|
|
| 33 |
B |
1210 |
1203 |
5.01 |
|
|
|
| 34 |
B |
1137 |
1130 |
2.68 |
|
|
|
| 35 |
B |
992 |
986 |
17.82 |
|
|
|
| 36 |
B |
933 |
927 |
6.33 |
|
|
|
| 37 |
B |
907 |
901 |
1.02 |
|
|
|
| 38 |
B |
893 |
888 |
46.57 |
|
|
|
| 39 |
B |
640 |
637 |
16.57 |
|
|
|
| 40 |
B |
424 |
421 |
1.94 |
|
|
|
| 41 |
B |
211 |
210 |
0.16 |
|
|
|
| 42 |
B |
114 |
114 |
0.16 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30155.0 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 29980.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.006 |
|
|
|
| 2 |
C |
0.006 |
|
|
|
| 3 |
C |
-0.092 |
|
|
|
| 4 |
C |
-0.092 |
|
|
|
| 5 |
C |
-0.003 |
|
|
|
| 6 |
C |
-0.003 |
|
|
|
| 7 |
H |
0.034 |
|
|
|
| 8 |
H |
0.019 |
|
|
|
| 9 |
H |
0.034 |
|
|
|
| 10 |
H |
0.019 |
|
|
|
| 11 |
H |
-0.007 |
|
|
|
| 12 |
H |
-0.007 |
|
|
|
| 13 |
H |
0.025 |
|
|
|
| 14 |
H |
0.017 |
|
|
|
| 15 |
H |
0.025 |
|
|
|
| 16 |
H |
0.017 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.286 |
0.286 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.231 |
1.364 |
0.000 |
| y |
1.364 |
-39.616 |
0.000 |
| z |
0.000 |
0.000 |
-37.385 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.730 |
1.364 |
0.000 |
| y |
1.364 |
-1.308 |
0.000 |
| z |
0.000 |
0.000 |
2.038 |
|
| Polar |
|---|
| 3z2-r2 | 4.076 |
|---|
| x2-y2 | 0.385 |
|---|
| xy | 1.364 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.679 |
1.771 |
0.000 |
| y |
1.771 |
13.853 |
0.000 |
| z |
0.000 |
0.000 |
9.549 |
<r2> (average value of r
2) Å
2
| <r2> |
257.598 |
| (<r2>)1/2 |
16.050 |