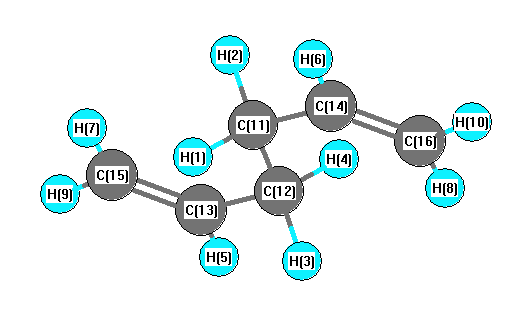
Vibrational Frequencies calculated at PBEPBEultrafine/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3164 |
3146 |
6.94 |
|
|
|
| 2 |
A |
3069 |
3051 |
0.43 |
|
|
|
| 3 |
A |
3054 |
3037 |
39.09 |
|
|
|
| 4 |
A |
3015 |
2998 |
19.42 |
|
|
|
| 5 |
A |
2939 |
2922 |
7.57 |
|
|
|
| 6 |
A |
1669 |
1659 |
5.09 |
|
|
|
| 7 |
A |
1401 |
1393 |
3.24 |
|
|
|
| 8 |
A |
1377 |
1370 |
1.34 |
|
|
|
| 9 |
A |
1303 |
1295 |
0.05 |
|
|
|
| 10 |
A |
1269 |
1261 |
1.79 |
|
|
|
| 11 |
A |
1193 |
1186 |
0.01 |
|
|
|
| 12 |
A |
1052 |
1046 |
0.07 |
|
|
|
| 13 |
A |
1019 |
1013 |
1.79 |
|
|
|
| 14 |
A |
990 |
984 |
7.10 |
|
|
|
| 15 |
A |
919 |
914 |
0.00 |
|
|
|
| 16 |
A |
893 |
887 |
13.79 |
|
|
|
| 17 |
A |
764 |
759 |
0.52 |
|
|
|
| 18 |
A |
618 |
614 |
6.72 |
|
|
|
| 19 |
A |
400 |
398 |
0.00 |
|
|
|
| 20 |
A |
340 |
338 |
0.06 |
|
|
|
| 21 |
A |
95 |
94 |
0.05 |
|
|
|
| 22 |
A |
72 |
72 |
0.02 |
|
|
|
| 23 |
B |
3164 |
3146 |
23.70 |
|
|
|
| 24 |
B |
3069 |
3051 |
7.20 |
|
|
|
| 25 |
B |
3054 |
3036 |
14.50 |
|
|
|
| 26 |
B |
2998 |
2981 |
6.45 |
|
|
|
| 27 |
B |
2942 |
2925 |
37.49 |
|
|
|
| 28 |
B |
1672 |
1663 |
20.46 |
|
|
|
| 29 |
B |
1413 |
1405 |
6.10 |
|
|
|
| 30 |
B |
1381 |
1373 |
0.09 |
|
|
|
| 31 |
B |
1275 |
1267 |
0.86 |
|
|
|
| 32 |
B |
1254 |
1247 |
0.63 |
|
|
|
| 33 |
B |
1208 |
1201 |
4.94 |
|
|
|
| 34 |
B |
1137 |
1130 |
2.68 |
|
|
|
| 35 |
B |
991 |
986 |
17.92 |
|
|
|
| 36 |
B |
932 |
926 |
6.32 |
|
|
|
| 37 |
B |
909 |
903 |
0.98 |
|
|
|
| 38 |
B |
893 |
888 |
46.80 |
|
|
|
| 39 |
B |
640 |
636 |
16.53 |
|
|
|
| 40 |
B |
423 |
420 |
1.94 |
|
|
|
| 41 |
B |
210 |
209 |
0.17 |
|
|
|
| 42 |
B |
111 |
110 |
0.16 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30143.2 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 29968.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.006 |
|
|
|
| 2 |
C |
0.006 |
|
|
|
| 3 |
C |
-0.092 |
|
|
|
| 4 |
C |
-0.092 |
|
|
|
| 5 |
C |
-0.003 |
|
|
|
| 6 |
C |
-0.003 |
|
|
|
| 7 |
H |
0.034 |
|
|
|
| 8 |
H |
0.019 |
|
|
|
| 9 |
H |
0.034 |
|
|
|
| 10 |
H |
0.019 |
|
|
|
| 11 |
H |
-0.007 |
|
|
|
| 12 |
H |
-0.007 |
|
|
|
| 13 |
H |
0.025 |
|
|
|
| 14 |
H |
0.017 |
|
|
|
| 15 |
H |
0.025 |
|
|
|
| 16 |
H |
0.017 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.285 |
0.285 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.239 |
1.367 |
0.000 |
| y |
1.367 |
-39.618 |
0.000 |
| z |
0.000 |
0.000 |
-37.375 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.743 |
1.367 |
0.000 |
| y |
1.367 |
-1.311 |
0.000 |
| z |
0.000 |
0.000 |
2.054 |
|
| Polar |
|---|
| 3z2-r2 | 4.107 |
|---|
| x2-y2 | 0.378 |
|---|
| xy | 1.367 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.676 |
1.781 |
0.000 |
| y |
1.781 |
13.849 |
0.000 |
| z |
0.000 |
0.000 |
9.556 |
<r2> (average value of r
2) Å
2
| <r2> |
257.592 |
| (<r2>)1/2 |
16.050 |