return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H8 (methylcyclopropane)

using model chemistry: QCISD(T)/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD(T)/cc-pVDZ
 hartrees
Energy at 0K-156.746169
Energy at 298.15K-156.754730
HF Energy-156.105337
Nuclear repulsion energy123.113495
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3239 3087        
2 A' 3155 3007        
3 A' 3142 2995        
4 A' 3114 2968        
5 A' 3032 2890        
6 A' 1514 1443        
7 A' 1487 1417        
8 A' 1427 1360        
9 A' 1381 1316        
10 A' 1227 1170        
11 A' 1207 1150        
12 A' 1056 1006        
13 A' 1003 956        
14 A' 933 890        
15 A' 795 757        
16 A' 767 731        
17 A' 357 340        
18 A" 3225 3074        
19 A" 3135 2988        
20 A" 3114 2968        
21 A" 1480 1411        
22 A" 1452 1384        
23 A" 1188 1132        
24 A" 1140 1087        
25 A" 1103 1052        
26 A" 1062 1012        
27 A" 879 838        
28 A" 812 774        
29 A" 337 322        
30 A" 225 215        

Unscaled Zero Point Vibrational Energy (zpe) 23993.4 cm-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 22868.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD(T)/cc-pVDZ
ABC
0.50982 0.20870 0.18331

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD(T)/cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.351 0.411 0.000
C2 -0.892 1.284 0.000
H3 1.305 0.959 0.000
C4 0.351 -0.904 0.763
C5 0.351 -0.904 -0.763
H6 1.270 -1.210 1.278
H7 1.270 -1.210 -1.278
H8 -0.578 -1.193 1.271
H9 -0.578 -1.193 -1.271
H10 -1.804 0.656 0.000
H11 -0.928 1.933 0.897
H12 -0.928 1.933 -0.897

Atom - Atom Distances (Å)
  C1 C2 H3 C4 C5 H6 H7 H8 H9 H10 H11 H12
C11.51921.09961.52041.52042.25932.25932.24772.24772.16952.18082.1808
C21.51922.22052.62992.62993.53953.53952.80182.80181.10761.10741.1074
H31.09962.22052.22772.22772.51772.51773.12923.12923.12372.59532.5953
C41.52042.62992.22771.52531.09692.25851.09802.25442.76823.11513.5269
C51.52042.62992.22771.52532.25851.09692.25441.09802.76823.52693.1151
H62.25933.53952.51771.09692.25852.55551.84843.14833.81663.85404.4087
H72.25933.53952.51772.25851.09692.55553.14831.84843.81664.40873.8540
H82.24772.80183.12921.09802.25441.84843.14832.54172.55683.16773.8200
H92.24772.80183.12922.25441.09803.14831.84842.54172.55683.82003.1677
H102.16951.10763.12372.76822.76823.81663.81662.55682.55681.78981.7898
H112.18081.10742.59533.11513.52693.85404.40873.16773.82001.78981.7934
H122.18081.10742.59533.52693.11514.40873.85403.82003.16771.78981.7934

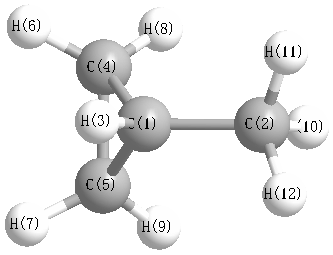
picture of methylcyclopropane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 H10 110.377 C1 C2 H11 111.286
C1 C2 H12 111.286 C1 C4 H5 59.895
C1 C4 H6 118.465 C1 C4 H8 117.367
C1 H5 C4 59.895 C1 H5 H7 118.465
C1 H5 H9 117.367 C2 C1 C3 115.029
C2 C1 C4 119.810 C2 C1 H5 119.810
C3 C1 C4 115.544 C3 C1 H5 115.544
C4 C1 H5 60.211 C4 H5 H7 118.011
C4 H5 H9 117.571 H5 C4 H6 118.011
H5 C4 H8 117.571 H6 C4 H8 114.740
H7 H5 H9 114.740 H10 C2 H11 107.805
H10 C2 H12 107.805 H11 C2 H12 108.138
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability