return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CCl2CH3 (Propane, 2,2-dichloro-)

using model chemistry: QCISD/cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at QCISD/cc-pVDZ
 hartrees
Energy at 0K-1036.882800
Energy at 298.15K 
HF Energy-1036.115653
Nuclear repulsion energy293.444183
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3184 3055 9.05      
2 A1 3076 2951 13.38      
3 A1 1489 1428 2.58      
4 A1 1425 1368 3.36      
5 A1 1198 1149 32.45      
6 A1 954 915 7.24      
7 A1 575 551 14.13      
8 A1 367 352 0.41      
9 A1 261 250 1.06      
10 A2 3161 3033 0.00      
11 A2 1477 1417 0.00      
12 A2 1026 984 0.00      
13 A2 289 277 0.00      
14 A2 279 268 0.00      
15 B1 3166 3038 22.20      
16 B1 1498 1437 4.85      
17 B1 1150 1104 72.39      
18 B1 699 670 79.06      
19 B1 367 352 1.81      
20 B1 315 302 0.00      
21 B2 3182 3053 5.36      
22 B2 3072 2948 6.52      
23 B2 1476 1416 3.07      
24 B2 1411 1354 9.33      
25 B2 1241 1191 5.79      
26 B2 961 922 0.01      
27 B2 395 379 3.47      

Unscaled Zero Point Vibrational Energy (zpe) 18845.8 cm-1
Scaled (by 0.9594) Zero Point Vibrational Energy (zpe) 18080.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/cc-pVDZ
ABC
0.12108 0.08028 0.07063

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/cc-pVDZ

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.339
Cl2 1.472 0.000 -0.720
Cl3 -1.472 0.000 -0.720
C4 0.000 1.276 1.177
C5 0.000 -1.276 1.177
H6 0.000 2.166 0.528
H7 0.000 -2.166 0.528
H8 -0.900 1.295 1.816
H9 0.900 1.295 1.816
H10 0.900 -1.295 1.816
H11 -0.900 -1.295 1.816

Atom - Atom Distances (Å)
  C1 Cl2 Cl3 C4 C5 H6 H7 H8 H9 H10 H11
C11.81311.81311.52661.52662.17382.17382.16082.16082.16082.1608
Cl21.81312.94312.71922.71922.90062.90063.70612.90452.90453.7061
Cl31.81312.94312.71922.71922.90062.90062.90453.70613.70612.9045
C41.52662.71922.71922.55141.10153.50201.10401.10402.79752.7975
C51.52662.71922.71922.55143.50201.10152.79752.79751.10401.1040
H62.17382.90062.90061.10153.50204.33111.79651.79653.80053.8005
H72.17382.90062.90063.50201.10154.33113.80053.80051.79651.7965
H82.16083.70612.90451.10402.79751.79653.80051.80043.15402.5897
H92.16082.90453.70611.10402.79751.79653.80051.80042.58973.1540
H102.16082.90453.70612.79751.10403.80051.79653.15402.58971.8004
H112.16083.70612.90452.79751.10403.80051.79652.58973.15401.8004

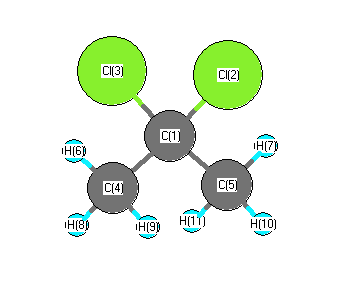
picture of Propane, 2,2-dichloro- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C4 H6 110.571 C1 C4 H8 109.406
C1 C4 H9 109.406 C1 C5 H7 110.571
C1 C5 H10 109.406 C1 C5 H11 109.406
Cl2 C1 Cl3 108.509 Cl2 C1 C4 108.714
Cl2 C1 C5 108.714 Cl3 C1 C4 108.714
Cl3 C1 C5 108.714 C4 C1 C5 113.373
H6 C4 H8 109.089 H6 C4 H9 109.089
H7 C5 H10 109.089 H7 C5 H11 109.089
H8 C4 H9 109.260 H10 C5 H11 109.260
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability