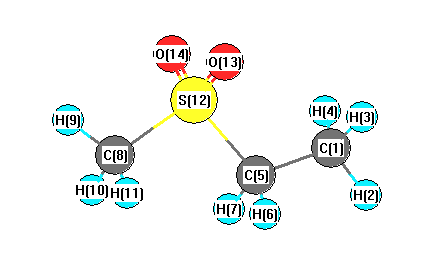
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3121 |
3103 |
0.93 |
|
|
|
| 2 |
A |
3117 |
3099 |
0.95 |
|
|
|
| 3 |
A |
3083 |
3065 |
5.31 |
|
|
|
| 4 |
A |
3065 |
3047 |
7.67 |
|
|
|
| 5 |
A |
3053 |
3036 |
6.33 |
|
|
|
| 6 |
A |
3000 |
2982 |
5.38 |
|
|
|
| 7 |
A |
2995 |
2978 |
1.69 |
|
|
|
| 8 |
A |
2982 |
2965 |
17.00 |
|
|
|
| 9 |
A |
1425 |
1417 |
6.31 |
|
|
|
| 10 |
A |
1416 |
1407 |
9.45 |
|
|
|
| 11 |
A |
1382 |
1374 |
5.48 |
|
|
|
| 12 |
A |
1372 |
1364 |
3.46 |
|
|
|
| 13 |
A |
1369 |
1361 |
5.42 |
|
|
|
| 14 |
A |
1338 |
1330 |
1.26 |
|
|
|
| 15 |
A |
1254 |
1246 |
101.25 |
|
|
|
| 16 |
A |
1241 |
1234 |
46.73 |
|
|
|
| 17 |
A |
1213 |
1206 |
30.99 |
|
|
|
| 18 |
A |
1197 |
1191 |
10.45 |
|
|
|
| 19 |
A |
1063 |
1057 |
107.07 |
|
|
|
| 20 |
A |
1051 |
1045 |
15.72 |
|
|
|
| 21 |
A |
997 |
991 |
8.34 |
|
|
|
| 22 |
A |
949 |
944 |
3.49 |
|
|
|
| 23 |
A |
902 |
897 |
32.88 |
|
|
|
| 24 |
A |
888 |
883 |
1.19 |
|
|
|
| 25 |
A |
755 |
751 |
38.36 |
|
|
|
| 26 |
A |
666 |
662 |
45.94 |
|
|
|
| 27 |
A |
591 |
587 |
14.22 |
|
|
|
| 28 |
A |
447 |
444 |
15.01 |
|
|
|
| 29 |
A |
404 |
401 |
22.38 |
|
|
|
| 30 |
A |
364 |
362 |
7.82 |
|
|
|
| 31 |
A |
285 |
284 |
1.08 |
|
|
|
| 32 |
A |
266 |
264 |
1.02 |
|
|
|
| 33 |
A |
213 |
212 |
2.08 |
|
|
|
| 34 |
A |
195 |
193 |
0.90 |
|
|
|
| 35 |
A |
168 |
167 |
0.30 |
|
|
|
| 36 |
A |
83 |
82 |
2.03 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23954.9 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 23816.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.064 |
|
|
|
| 2 |
H |
0.067 |
|
|
|
| 3 |
H |
0.049 |
|
|
|
| 4 |
H |
0.044 |
|
|
|
| 5 |
C |
-0.184 |
|
|
|
| 6 |
H |
0.084 |
|
|
|
| 7 |
H |
0.063 |
|
|
|
| 8 |
C |
-0.161 |
|
|
|
| 9 |
H |
0.075 |
|
|
|
| 10 |
H |
0.095 |
|
|
|
| 11 |
H |
0.075 |
|
|
|
| 12 |
S |
0.695 |
|
|
|
| 13 |
O |
-0.417 |
|
|
|
| 14 |
O |
-0.421 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.655 |
2.584 |
-2.091 |
4.254 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-43.733 |
-4.752 |
-1.232 |
| y |
-4.752 |
-42.101 |
-0.650 |
| z |
-1.232 |
-0.650 |
-47.520 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.077 |
-4.752 |
-1.232 |
| y |
-4.752 |
3.525 |
-0.650 |
| z |
-1.232 |
-0.650 |
-4.602 |
|
| Polar |
|---|
| 3z2-r2 | -9.205 |
|---|
| x2-y2 | -1.632 |
|---|
| xy | -4.752 |
|---|
| xz | -1.232 |
|---|
| yz | -0.650 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.802 |
-0.298 |
0.152 |
| y |
-0.298 |
8.622 |
0.093 |
| z |
0.152 |
0.093 |
8.125 |
<r2> (average value of r
2) Å
2
| <r2> |
196.767 |
| (<r2>)1/2 |
14.027 |