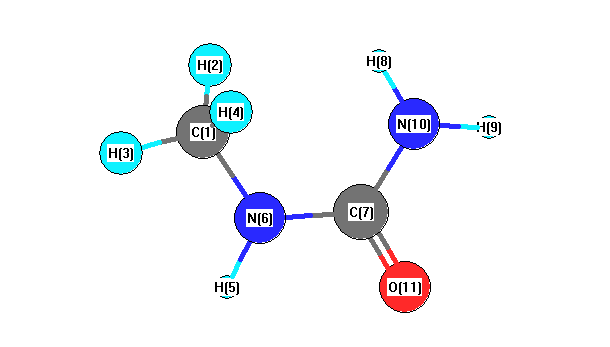
Vibrational Frequencies calculated at BLYP/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3529 |
3535 |
16.16 |
|
|
|
| 2 |
A |
3491 |
3496 |
19.88 |
|
|
|
| 3 |
A |
3412 |
3418 |
5.15 |
|
|
|
| 4 |
A |
3038 |
3042 |
19.72 |
|
|
|
| 5 |
A |
2959 |
2963 |
46.53 |
|
|
|
| 6 |
A |
2914 |
2919 |
57.77 |
|
|
|
| 7 |
A |
1732 |
1735 |
344.83 |
|
|
|
| 8 |
A |
1555 |
1557 |
93.92 |
|
|
|
| 9 |
A |
1451 |
1453 |
27.08 |
|
|
|
| 10 |
A |
1427 |
1429 |
47.88 |
|
|
|
| 11 |
A |
1417 |
1419 |
8.29 |
|
|
|
| 12 |
A |
1386 |
1388 |
4.87 |
|
|
|
| 13 |
A |
1343 |
1345 |
150.35 |
|
|
|
| 14 |
A |
1152 |
1153 |
1.20 |
|
|
|
| 15 |
A |
1102 |
1104 |
28.85 |
|
|
|
| 16 |
A |
1096 |
1098 |
14.20 |
|
|
|
| 17 |
A |
1031 |
1033 |
34.01 |
|
|
|
| 18 |
A |
852 |
853 |
3.31 |
|
|
|
| 19 |
A |
742 |
743 |
48.25 |
|
|
|
| 20 |
A |
601 |
602 |
30.91 |
|
|
|
| 21 |
A |
577 |
578 |
150.33 |
|
|
|
| 22 |
A |
524 |
525 |
12.29 |
|
|
|
| 23 |
A |
477 |
478 |
40.09 |
|
|
|
| 24 |
A |
383 |
383 |
63.70 |
|
|
|
| 25 |
A |
265 |
265 |
2.28 |
|
|
|
| 26 |
A |
148 |
148 |
5.49 |
|
|
|
| 27 |
A |
102 |
102 |
2.97 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19351.1 cm
-1
Scaled (by 1.0016) Zero Point Vibrational Energy (zpe) 19382.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.063 |
|
|
|
| 2 |
H |
0.016 |
|
|
|
| 3 |
H |
0.018 |
|
|
|
| 4 |
H |
0.022 |
|
|
|
| 5 |
H |
0.082 |
|
|
|
| 6 |
N |
-0.113 |
|
|
|
| 7 |
C |
0.081 |
|
|
|
| 8 |
H |
0.075 |
|
|
|
| 9 |
H |
0.088 |
|
|
|
| 10 |
N |
-0.078 |
|
|
|
| 11 |
O |
-0.254 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.022 |
1.781 |
-0.641 |
3.566 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.171 |
4.973 |
-0.900 |
| y |
4.973 |
-26.622 |
-2.041 |
| z |
-0.900 |
-2.041 |
-31.207 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.256 |
4.973 |
-0.900 |
| y |
4.973 |
4.567 |
-2.041 |
| z |
-0.900 |
-2.041 |
-2.310 |
|
| Polar |
|---|
| 3z2-r2 | -4.621 |
|---|
| x2-y2 | -4.549 |
|---|
| xy | 4.973 |
|---|
| xz | -0.900 |
|---|
| yz | -2.041 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.688 |
-0.002 |
0.121 |
| y |
-0.002 |
6.540 |
-0.141 |
| z |
0.121 |
-0.141 |
3.832 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |