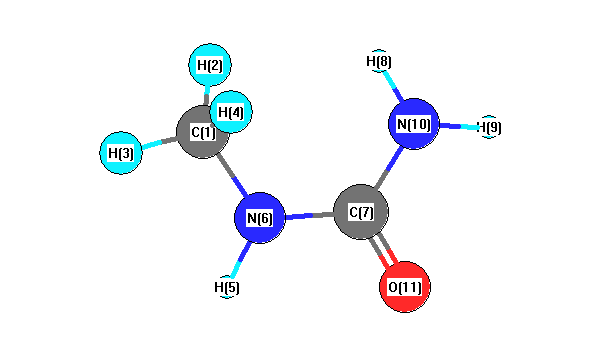
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3572 |
3551 |
23.09 |
|
|
|
| 2 |
A |
3531 |
3510 |
27.36 |
|
|
|
| 3 |
A |
3448 |
3428 |
7.16 |
|
|
|
| 4 |
A |
3067 |
3049 |
13.51 |
|
|
|
| 5 |
A |
2981 |
2964 |
40.21 |
|
|
|
| 6 |
A |
2929 |
2912 |
54.52 |
|
|
|
| 7 |
A |
1766 |
1756 |
364.41 |
|
|
|
| 8 |
A |
1547 |
1538 |
127.63 |
|
|
|
| 9 |
A |
1446 |
1437 |
67.29 |
|
|
|
| 10 |
A |
1430 |
1421 |
55.12 |
|
|
|
| 11 |
A |
1407 |
1399 |
8.16 |
|
|
|
| 12 |
A |
1382 |
1374 |
8.49 |
|
|
|
| 13 |
A |
1350 |
1343 |
81.50 |
|
|
|
| 14 |
A |
1164 |
1158 |
6.68 |
|
|
|
| 15 |
A |
1108 |
1101 |
27.51 |
|
|
|
| 16 |
A |
1095 |
1088 |
0.67 |
|
|
|
| 17 |
A |
1036 |
1030 |
29.14 |
|
|
|
| 18 |
A |
869 |
864 |
3.93 |
|
|
|
| 19 |
A |
748 |
744 |
44.32 |
|
|
|
| 20 |
A |
597 |
593 |
33.09 |
|
|
|
| 21 |
A |
569 |
565 |
141.79 |
|
|
|
| 22 |
A |
525 |
522 |
13.44 |
|
|
|
| 23 |
A |
473 |
471 |
41.17 |
|
|
|
| 24 |
A |
383 |
381 |
73.59 |
|
|
|
| 25 |
A |
264 |
262 |
2.90 |
|
|
|
| 26 |
A |
146 |
145 |
5.96 |
|
|
|
| 27 |
A |
104 |
103 |
2.75 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19467.3 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 19354.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.002 |
|
|
|
| 2 |
H |
0.040 |
|
|
|
| 3 |
H |
0.045 |
|
|
|
| 4 |
H |
0.046 |
|
|
|
| 5 |
H |
0.098 |
|
|
|
| 6 |
N |
-0.156 |
|
|
|
| 7 |
C |
0.125 |
|
|
|
| 8 |
H |
0.089 |
|
|
|
| 9 |
H |
0.102 |
|
|
|
| 10 |
N |
-0.119 |
|
|
|
| 11 |
O |
-0.268 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.092 |
1.778 |
-0.640 |
3.623 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.015 |
4.962 |
-0.866 |
| y |
4.962 |
-26.283 |
-2.076 |
| z |
-0.866 |
-2.076 |
-31.175 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.286 |
4.962 |
-0.866 |
| y |
4.962 |
4.812 |
-2.076 |
| z |
-0.866 |
-2.076 |
-2.526 |
|
| Polar |
|---|
| 3z2-r2 | -5.052 |
|---|
| x2-y2 | -4.733 |
|---|
| xy | 4.962 |
|---|
| xz | -0.866 |
|---|
| yz | -2.076 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.598 |
-0.029 |
0.110 |
| y |
-0.029 |
6.458 |
-0.134 |
| z |
0.110 |
-0.134 |
3.826 |
<r2> (average value of r
2) Å
2
| <r2> |
119.824 |
| (<r2>)1/2 |
10.946 |