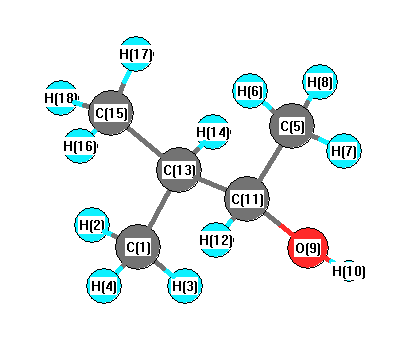
Vibrational Frequencies calculated at PBEPBE/cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3666 |
3645 |
9.89 |
|
|
|
| 2 |
A |
3081 |
3063 |
12.11 |
|
|
|
| 3 |
A |
3065 |
3047 |
21.25 |
|
|
|
| 4 |
A |
3056 |
3038 |
28.28 |
|
|
|
| 5 |
A |
3041 |
3023 |
74.28 |
|
|
|
| 6 |
A |
3039 |
3021 |
10.39 |
|
|
|
| 7 |
A |
3036 |
3019 |
3.54 |
|
|
|
| 8 |
A |
2966 |
2948 |
20.13 |
|
|
|
| 9 |
A |
2963 |
2946 |
41.53 |
|
|
|
| 10 |
A |
2961 |
2944 |
21.69 |
|
|
|
| 11 |
A |
2944 |
2927 |
18.26 |
|
|
|
| 12 |
A |
2851 |
2834 |
57.44 |
|
|
|
| 13 |
A |
1442 |
1434 |
6.47 |
|
|
|
| 14 |
A |
1428 |
1419 |
2.07 |
|
|
|
| 15 |
A |
1421 |
1413 |
10.25 |
|
|
|
| 16 |
A |
1416 |
1408 |
7.60 |
|
|
|
| 17 |
A |
1410 |
1401 |
3.65 |
|
|
|
| 18 |
A |
1407 |
1399 |
2.61 |
|
|
|
| 19 |
A |
1362 |
1354 |
10.26 |
|
|
|
| 20 |
A |
1346 |
1338 |
4.61 |
|
|
|
| 21 |
A |
1338 |
1330 |
8.20 |
|
|
|
| 22 |
A |
1331 |
1323 |
5.60 |
|
|
|
| 23 |
A |
1305 |
1297 |
4.65 |
|
|
|
| 24 |
A |
1286 |
1278 |
1.36 |
|
|
|
| 25 |
A |
1278 |
1271 |
17.91 |
|
|
|
| 26 |
A |
1223 |
1216 |
22.92 |
|
|
|
| 27 |
A |
1151 |
1144 |
6.10 |
|
|
|
| 28 |
A |
1138 |
1132 |
6.91 |
|
|
|
| 29 |
A |
1110 |
1104 |
1.56 |
|
|
|
| 30 |
A |
1062 |
1056 |
78.44 |
|
|
|
| 31 |
A |
1054 |
1048 |
13.48 |
|
|
|
| 32 |
A |
950 |
945 |
5.11 |
|
|
|
| 33 |
A |
934 |
929 |
5.61 |
|
|
|
| 34 |
A |
919 |
914 |
8.13 |
|
|
|
| 35 |
A |
895 |
889 |
0.23 |
|
|
|
| 36 |
A |
877 |
872 |
9.25 |
|
|
|
| 37 |
A |
768 |
763 |
1.67 |
|
|
|
| 38 |
A |
512 |
509 |
3.12 |
|
|
|
| 39 |
A |
447 |
445 |
7.72 |
|
|
|
| 40 |
A |
398 |
396 |
1.35 |
|
|
|
| 41 |
A |
357 |
355 |
3.26 |
|
|
|
| 42 |
A |
350 |
348 |
1.94 |
|
|
|
| 43 |
A |
277 |
276 |
5.94 |
|
|
|
| 44 |
A |
268 |
266 |
85.63 |
|
|
|
| 45 |
A |
238 |
236 |
0.31 |
|
|
|
| 46 |
A |
223 |
221 |
0.01 |
|
|
|
| 47 |
A |
209 |
208 |
0.30 |
|
|
|
| 48 |
A |
76 |
76 |
1.35 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 34936.3 cm
-1
Scaled (by 0.9942) Zero Point Vibrational Energy (zpe) 34733.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.004 |
|
|
|
| 2 |
H |
0.020 |
|
|
|
| 3 |
H |
0.036 |
|
|
|
| 4 |
H |
0.018 |
|
|
|
| 5 |
C |
-0.012 |
|
|
|
| 6 |
H |
0.030 |
|
|
|
| 7 |
H |
0.015 |
|
|
|
| 8 |
H |
0.034 |
|
|
|
| 9 |
O |
-0.229 |
|
|
|
| 10 |
H |
0.132 |
|
|
|
| 11 |
C |
0.032 |
|
|
|
| 12 |
H |
-0.010 |
|
|
|
| 13 |
C |
-0.129 |
|
|
|
| 14 |
H |
0.004 |
|
|
|
| 15 |
C |
-0.017 |
|
|
|
| 16 |
H |
0.025 |
|
|
|
| 17 |
H |
0.028 |
|
|
|
| 18 |
H |
0.027 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.863 |
0.750 |
0.703 |
1.342 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.547 |
-1.063 |
1.809 |
| y |
-1.063 |
-41.612 |
-1.095 |
| z |
1.809 |
-1.095 |
-40.010 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.264 |
-1.063 |
1.809 |
| y |
-1.063 |
-3.333 |
-1.095 |
| z |
1.809 |
-1.095 |
-0.931 |
|
| Polar |
|---|
| 3z2-r2 | -1.862 |
|---|
| x2-y2 | 5.064 |
|---|
| xy | -1.063 |
|---|
| xz | 1.809 |
|---|
| yz | -1.095 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.473 |
0.138 |
0.119 |
| y |
0.138 |
9.653 |
-0.014 |
| z |
0.119 |
-0.014 |
8.335 |
<r2> (average value of r
2) Å
2
| <r2> |
197.184 |
| (<r2>)1/2 |
14.042 |